Jump to
S1C2
Energy calculated at PBE1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -133.133864 |
| Energy at 298.15K | -133.138044 |
| HF Energy | -133.133864 |
| Nuclear repulsion energy | 64.900446 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBE1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3480 |
3317 |
1.44 |
|
|
|
| 2 |
A' |
3314 |
3159 |
4.08 |
|
|
|
| 3 |
A' |
3205 |
3055 |
3.23 |
|
|
|
| 4 |
A' |
3090 |
2945 |
49.03 |
|
|
|
| 5 |
A' |
1539 |
1467 |
8.98 |
|
|
|
| 6 |
A' |
1481 |
1411 |
2.38 |
|
|
|
| 7 |
A' |
1382 |
1317 |
35.52 |
|
|
|
| 8 |
A' |
1274 |
1215 |
8.29 |
|
|
|
| 9 |
A' |
1134 |
1081 |
22.47 |
|
|
|
| 10 |
A' |
1003 |
956 |
9.76 |
|
|
|
| 11 |
A' |
488 |
465 |
11.79 |
|
|
|
| 12 |
A" |
1055 |
1006 |
0.25 |
|
|
|
| 13 |
A" |
808 |
770 |
10.89 |
|
|
|
| 14 |
A" |
685 |
653 |
116.81 |
|
|
|
| 15 |
A" |
534 |
509 |
5.79 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12235.1 cm
-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 11661.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBE1PBE/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.418 |
0.000 |
| C2 |
1.151 |
-0.376 |
0.000 |
| N3 |
-1.190 |
-0.137 |
0.000 |
| H4 |
0.142 |
1.506 |
0.000 |
| H5 |
2.143 |
0.061 |
0.000 |
| H6 |
1.050 |
-1.457 |
0.000 |
| H7 |
-1.907 |
0.594 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
| C1 | | 1.3981 | 1.3132 | 1.0977 | 2.1728 | 2.1488 | 1.9148 |
C2 | 1.3981 | | 2.3531 | 2.1357 | 1.0846 | 1.0852 | 3.2075 | N3 | 1.3132 | 2.3531 | | 2.1155 | 3.3394 | 2.6004 | 1.0234 | H4 | 1.0977 | 2.1357 | 2.1155 | | 2.4687 | 3.0992 | 2.2427 | H5 | 2.1728 | 1.0846 | 3.3394 | 2.4687 | | 1.8703 | 4.0849 | H6 | 2.1488 | 1.0852 | 2.6004 | 3.0992 | 1.8703 | | 3.5984 | H7 | 1.9148 | 3.2075 | 1.0234 | 2.2427 | 4.0849 | 3.5984 | |
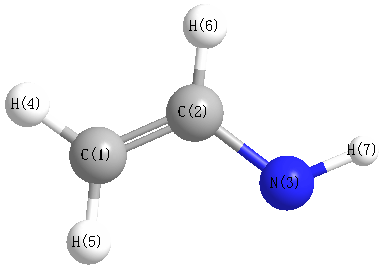 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H5 |
121.623 |
|
C1 |
C2 |
H6 |
119.301 |
| C1 |
N3 |
H7 |
109.442 |
|
C2 |
C1 |
N3 |
120.399 |
| C2 |
C1 |
H4 |
117.175 |
|
N3 |
C1 |
H4 |
122.426 |
| H5 |
C2 |
H6 |
119.076 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.023 |
|
|
|
| 2 |
C |
-0.337 |
|
|
|
| 3 |
N |
-0.531 |
|
|
|
| 4 |
H |
0.154 |
|
|
|
| 5 |
H |
0.182 |
|
|
|
| 6 |
H |
0.199 |
|
|
|
| 7 |
H |
0.309 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.040 |
1.618 |
0.000 |
1.923 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-16.491 |
-2.850 |
0.000 |
| y |
-2.850 |
-17.399 |
0.000 |
| z |
0.000 |
0.000 |
-19.874 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.145 |
-2.850 |
0.000 |
| y |
-2.850 |
0.783 |
0.000 |
| z |
0.000 |
0.000 |
-2.929 |
|
| Polar |
|---|
| 3z2-r2 | -5.858 |
|---|
| x2-y2 | 0.908 |
|---|
| xy | -2.850 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.286 |
-0.413 |
0.000 |
| y |
-0.413 |
4.000 |
0.000 |
| z |
0.000 |
0.000 |
1.750 |
<r2> (average value of r
2) Å
2
| <r2> |
45.182 |
| (<r2>)1/2 |
6.722 |
Jump to
S1C1
Energy calculated at PBE1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -133.133544 |
| Energy at 298.15K | -133.137708 |
| HF Energy | -133.133544 |
| Nuclear repulsion energy | 64.888169 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBE1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3427 |
3266 |
10.51 |
|
|
|
| 2 |
A' |
3293 |
3139 |
7.94 |
|
|
|
| 3 |
A' |
3188 |
3038 |
3.78 |
|
|
|
| 4 |
A' |
3157 |
3009 |
23.42 |
|
|
|
| 5 |
A' |
1522 |
1450 |
4.91 |
|
|
|
| 6 |
A' |
1476 |
1407 |
3.70 |
|
|
|
| 7 |
A' |
1404 |
1338 |
14.09 |
|
|
|
| 8 |
A' |
1263 |
1204 |
32.52 |
|
|
|
| 9 |
A' |
1161 |
1107 |
33.02 |
|
|
|
| 10 |
A' |
996 |
949 |
0.92 |
|
|
|
| 11 |
A' |
494 |
471 |
8.30 |
|
|
|
| 12 |
A" |
1087 |
1036 |
49.97 |
|
|
|
| 13 |
A" |
817 |
779 |
73.90 |
|
|
|
| 14 |
A" |
686 |
654 |
6.27 |
|
|
|
| 15 |
A" |
492 |
469 |
0.15 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12231.5 cm
-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 11657.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBE1PBE/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.447 |
0.000 |
| C2 |
1.128 |
-0.382 |
0.000 |
| N3 |
-1.257 |
0.068 |
0.000 |
| H4 |
0.166 |
1.527 |
0.000 |
| H5 |
2.133 |
0.028 |
0.000 |
| H6 |
1.023 |
-1.464 |
0.000 |
| H7 |
-1.294 |
-0.959 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
| C1 | | 1.3997 | 1.3125 | 1.0932 | 2.1736 | 2.1678 | 1.9105 |
C2 | 1.3997 | | 2.4267 | 2.1379 | 1.0852 | 1.0876 | 2.4899 | N3 | 1.3125 | 2.4267 | | 2.0380 | 3.3897 | 2.7468 | 1.0272 | H4 | 1.0932 | 2.1379 | 2.0380 | | 2.4733 | 3.1121 | 2.8830 | H5 | 2.1736 | 1.0852 | 3.3897 | 2.4733 | | 1.8598 | 3.5662 | H6 | 2.1678 | 1.0876 | 2.7468 | 3.1121 | 1.8598 | | 2.3717 | H7 | 1.9105 | 2.4899 | 1.0272 | 2.8830 | 3.5662 | 2.3717 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H5 |
121.510 |
|
C1 |
C2 |
H6 |
120.764 |
| C1 |
N3 |
H7 |
108.871 |
|
C2 |
C1 |
N3 |
126.917 |
| C2 |
C1 |
H4 |
117.572 |
|
N3 |
C1 |
H4 |
115.511 |
| H5 |
C2 |
H6 |
117.726 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.037 |
|
|
|
| 2 |
C |
-0.366 |
|
|
|
| 3 |
N |
-0.521 |
|
|
|
| 4 |
H |
0.178 |
|
|
|
| 5 |
H |
0.189 |
|
|
|
| 6 |
H |
0.183 |
|
|
|
| 7 |
H |
0.300 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.150 |
-1.118 |
0.000 |
2.423 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-20.502 |
2.138 |
0.000 |
| y |
2.138 |
-15.999 |
0.000 |
| z |
0.000 |
0.000 |
-19.887 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.559 |
2.138 |
0.000 |
| y |
2.138 |
4.195 |
0.000 |
| z |
0.000 |
0.000 |
-1.636 |
|
| Polar |
|---|
| 3z2-r2 | -3.272 |
|---|
| x2-y2 | -4.503 |
|---|
| xy | 2.138 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.966 |
-0.078 |
0.000 |
| y |
-0.078 |
4.188 |
0.000 |
| z |
0.000 |
0.000 |
1.753 |
<r2> (average value of r
2) Å
2
| <r2> |
45.228 |
| (<r2>)1/2 |
6.725 |
