Jump to
S1C2
Energy calculated at B2PLYP/6-31G**
| | hartrees |
|---|
| Energy at 0K | -133.155495 |
| Energy at 298.15K | -133.159685 |
| HF Energy | -133.021085 |
| Nuclear repulsion energy | 64.914618 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3489 |
3489 |
1.63 |
|
|
|
| 2 |
A' |
3328 |
3328 |
4.55 |
|
|
|
| 3 |
A' |
3214 |
3214 |
3.43 |
|
|
|
| 4 |
A' |
3103 |
3103 |
47.38 |
|
|
|
| 5 |
A' |
1538 |
1538 |
9.10 |
|
|
|
| 6 |
A' |
1490 |
1490 |
1.74 |
|
|
|
| 7 |
A' |
1384 |
1384 |
35.32 |
|
|
|
| 8 |
A' |
1266 |
1266 |
10.40 |
|
|
|
| 9 |
A' |
1115 |
1115 |
21.60 |
|
|
|
| 10 |
A' |
1011 |
1011 |
10.77 |
|
|
|
| 11 |
A' |
493 |
493 |
11.73 |
|
|
|
| 12 |
A" |
1074 |
1074 |
0.45 |
|
|
|
| 13 |
A" |
804 |
804 |
6.89 |
|
|
|
| 14 |
A" |
696 |
696 |
109.28 |
|
|
|
| 15 |
A" |
533 |
533 |
3.56 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12269.7 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 12269.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.419 |
0.000 |
| C2 |
1.151 |
-0.380 |
0.000 |
| N3 |
-1.191 |
-0.134 |
0.000 |
| H4 |
0.145 |
1.503 |
0.000 |
| H5 |
2.141 |
0.055 |
0.000 |
| H6 |
1.047 |
-1.456 |
0.000 |
| H7 |
-1.905 |
0.597 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
| C1 | | 1.4010 | 1.3128 | 1.0940 | 2.1716 | 2.1477 | 1.9132 |
C2 | 1.4010 | | 2.3545 | 2.1351 | 1.0809 | 1.0814 | 3.2083 | N3 | 1.3128 | 2.3545 | | 2.1128 | 3.3367 | 2.5992 | 1.0223 | H4 | 1.0940 | 2.1351 | 2.1128 | | 2.4664 | 3.0941 | 2.2408 | H5 | 2.1716 | 1.0809 | 3.3367 | 2.4664 | | 1.8651 | 4.0819 | H6 | 2.1477 | 1.0814 | 2.5992 | 3.0941 | 1.8651 | | 3.5960 | H7 | 1.9132 | 3.2083 | 1.0223 | 2.2408 | 4.0819 | 3.5960 | |
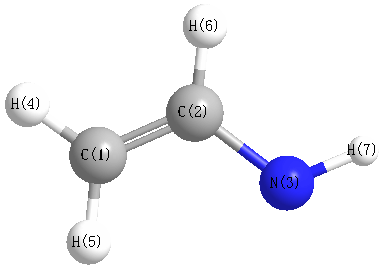 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H5 |
121.548 |
|
C1 |
C2 |
H6 |
119.246 |
| C1 |
N3 |
H7 |
109.410 |
|
C2 |
C1 |
N3 |
120.332 |
| C2 |
C1 |
H4 |
117.157 |
|
N3 |
C1 |
H4 |
122.511 |
| H5 |
C2 |
H6 |
119.206 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.090 |
|
|
|
| 2 |
C |
-0.202 |
|
|
|
| 3 |
N |
-0.469 |
|
|
|
| 4 |
H |
0.091 |
|
|
|
| 5 |
H |
0.119 |
|
|
|
| 6 |
H |
0.135 |
|
|
|
| 7 |
H |
0.236 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.075 |
1.604 |
0.000 |
1.931 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-16.817 |
-2.777 |
0.000 |
| y |
-2.777 |
-17.599 |
0.000 |
| z |
0.000 |
0.000 |
-19.838 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.901 |
-2.777 |
0.000 |
| y |
-2.777 |
0.728 |
0.000 |
| z |
0.000 |
0.000 |
-2.629 |
|
| Polar |
|---|
| 3z2-r2 | -5.259 |
|---|
| x2-y2 | 0.782 |
|---|
| xy | -2.777 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.315 |
-0.412 |
0.000 |
| y |
-0.412 |
4.002 |
0.000 |
| z |
0.000 |
0.000 |
1.766 |
<r2> (average value of r
2) Å
2
| <r2> |
45.281 |
| (<r2>)1/2 |
6.729 |
Jump to
S1C1
Energy calculated at B2PLYP/6-31G**
| | hartrees |
|---|
| Energy at 0K | -133.155170 |
| Energy at 298.15K | -133.159345 |
| HF Energy | -133.020614 |
| Nuclear repulsion energy | 64.906936 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3438 |
3438 |
10.94 |
|
|
|
| 2 |
A' |
3308 |
3308 |
8.32 |
|
|
|
| 3 |
A' |
3197 |
3197 |
3.49 |
|
|
|
| 4 |
A' |
3169 |
3169 |
23.73 |
|
|
|
| 5 |
A' |
1531 |
1531 |
4.74 |
|
|
|
| 6 |
A' |
1479 |
1479 |
3.00 |
|
|
|
| 7 |
A' |
1405 |
1405 |
13.61 |
|
|
|
| 8 |
A' |
1254 |
1254 |
41.36 |
|
|
|
| 9 |
A' |
1144 |
1144 |
26.09 |
|
|
|
| 10 |
A' |
1003 |
1003 |
0.25 |
|
|
|
| 11 |
A' |
499 |
499 |
8.73 |
|
|
|
| 12 |
A" |
1107 |
1107 |
46.19 |
|
|
|
| 13 |
A" |
814 |
814 |
67.61 |
|
|
|
| 14 |
A" |
698 |
698 |
4.75 |
|
|
|
| 15 |
A" |
494 |
494 |
0.18 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12269.4 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 12269.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.448 |
0.000 |
| C2 |
1.128 |
-0.385 |
0.000 |
| N3 |
-1.256 |
0.070 |
0.000 |
| H4 |
0.169 |
1.525 |
0.000 |
| H5 |
2.130 |
0.023 |
0.000 |
| H6 |
1.019 |
-1.463 |
0.000 |
| H7 |
-1.293 |
-0.955 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
| C1 | | 1.4025 | 1.3122 | 1.0896 | 2.1722 | 2.1659 | 1.9086 |
C2 | 1.4025 | | 2.4277 | 2.1367 | 1.0814 | 1.0837 | 2.4878 | N3 | 1.3122 | 2.4277 | | 2.0370 | 3.3868 | 2.7437 | 1.0261 | H4 | 1.0896 | 2.1367 | 2.0370 | | 2.4699 | 3.1061 | 2.8793 | H5 | 2.1722 | 1.0814 | 3.3868 | 2.4699 | | 1.8550 | 3.5601 | H6 | 2.1659 | 1.0837 | 2.7437 | 3.1061 | 1.8550 | | 2.3673 | H7 | 1.9086 | 2.4878 | 1.0261 | 2.8793 | 3.5601 | 2.3673 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H5 |
121.436 |
|
C1 |
C2 |
H6 |
120.654 |
| C1 |
N3 |
H7 |
108.804 |
|
C2 |
C1 |
N3 |
126.805 |
| C2 |
C1 |
H4 |
117.495 |
|
N3 |
C1 |
H4 |
115.700 |
| H5 |
C2 |
H6 |
117.910 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.105 |
|
|
|
| 2 |
C |
-0.229 |
|
|
|
| 3 |
N |
-0.461 |
|
|
|
| 4 |
H |
0.115 |
|
|
|
| 5 |
H |
0.125 |
|
|
|
| 6 |
H |
0.119 |
|
|
|
| 7 |
H |
0.226 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.163 |
-1.076 |
0.000 |
2.416 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-20.719 |
2.099 |
0.000 |
| y |
2.099 |
-16.255 |
0.000 |
| z |
0.000 |
0.000 |
-19.849 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.667 |
2.099 |
0.000 |
| y |
2.099 |
4.029 |
0.000 |
| z |
0.000 |
0.000 |
-1.362 |
|
| Polar |
|---|
| 3z2-r2 | -2.724 |
|---|
| x2-y2 | -4.464 |
|---|
| xy | 2.099 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.974 |
-0.049 |
0.000 |
| y |
-0.049 |
4.196 |
0.000 |
| z |
0.000 |
0.000 |
1.771 |
<r2> (average value of r
2) Å
2
| <r2> |
45.303 |
| (<r2>)1/2 |
6.731 |
