Jump to
S1C2
Energy calculated at B2PLYP=FULL/6-31G**
| | hartrees |
|---|
| Energy at 0K | -133.159415 |
| Energy at 298.15K | -133.163606 |
| HF Energy | -133.021097 |
| Nuclear repulsion energy | 64.932863 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULL/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3490 |
3490 |
1.60 |
|
|
|
| 2 |
A' |
3329 |
3329 |
4.51 |
|
|
|
| 3 |
A' |
3215 |
3215 |
3.40 |
|
|
|
| 4 |
A' |
3104 |
3104 |
47.36 |
|
|
|
| 5 |
A' |
1539 |
1539 |
9.06 |
|
|
|
| 6 |
A' |
1491 |
1491 |
1.75 |
|
|
|
| 7 |
A' |
1385 |
1385 |
35.42 |
|
|
|
| 8 |
A' |
1267 |
1267 |
10.44 |
|
|
|
| 9 |
A' |
1115 |
1115 |
21.68 |
|
|
|
| 10 |
A' |
1011 |
1011 |
10.75 |
|
|
|
| 11 |
A' |
493 |
493 |
11.73 |
|
|
|
| 12 |
A" |
1074 |
1074 |
0.45 |
|
|
|
| 13 |
A" |
806 |
806 |
6.97 |
|
|
|
| 14 |
A" |
697 |
697 |
109.27 |
|
|
|
| 15 |
A" |
534 |
534 |
3.58 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12274.9 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 12274.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP=FULL/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.419 |
0.000 |
| C2 |
1.151 |
-0.379 |
0.000 |
| N3 |
-1.190 |
-0.134 |
0.000 |
| H4 |
0.145 |
1.503 |
0.000 |
| H5 |
2.140 |
0.055 |
0.000 |
| H6 |
1.047 |
-1.455 |
0.000 |
| H7 |
-1.905 |
0.597 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
| C1 | | 1.4005 | 1.3124 | 1.0937 | 2.1709 | 2.1471 | 1.9129 |
C2 | 1.4005 | | 2.3539 | 2.1344 | 1.0806 | 1.0811 | 3.2076 | N3 | 1.3124 | 2.3539 | | 2.1121 | 3.3358 | 2.5986 | 1.0220 | H4 | 1.0937 | 2.1344 | 2.1121 | | 2.4657 | 3.0931 | 2.2404 | H5 | 2.1709 | 1.0806 | 3.3358 | 2.4657 | | 1.8645 | 4.0809 | H6 | 2.1471 | 1.0811 | 2.5986 | 3.0931 | 1.8645 | | 3.5952 | H7 | 1.9129 | 3.2076 | 1.0220 | 2.2404 | 4.0809 | 3.5952 | |
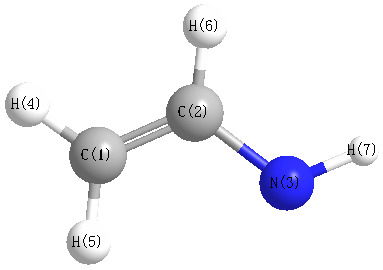 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H5 |
121.552 |
|
C1 |
C2 |
H6 |
119.246 |
| C1 |
N3 |
H7 |
109.427 |
|
C2 |
C1 |
N3 |
120.342 |
| C2 |
C1 |
H4 |
117.155 |
|
N3 |
C1 |
H4 |
122.503 |
| H5 |
C2 |
H6 |
119.203 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at B2PLYP=FULL/6-31G**
| | hartrees |
|---|
| Energy at 0K | -133.159093 |
| Energy at 298.15K | -133.163270 |
| HF Energy | -133.020626 |
| Nuclear repulsion energy | 64.925638 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULL/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3439 |
3439 |
10.87 |
|
|
|
| 2 |
A' |
3309 |
3309 |
8.27 |
|
|
|
| 3 |
A' |
3198 |
3198 |
3.48 |
|
|
|
| 4 |
A' |
3169 |
3169 |
23.72 |
|
|
|
| 5 |
A' |
1532 |
1532 |
4.76 |
|
|
|
| 6 |
A' |
1479 |
1479 |
2.99 |
|
|
|
| 7 |
A' |
1406 |
1406 |
13.52 |
|
|
|
| 8 |
A' |
1254 |
1254 |
41.49 |
|
|
|
| 9 |
A' |
1145 |
1145 |
26.19 |
|
|
|
| 10 |
A' |
1003 |
1003 |
0.26 |
|
|
|
| 11 |
A' |
499 |
499 |
8.73 |
|
|
|
| 12 |
A" |
1108 |
1108 |
46.10 |
|
|
|
| 13 |
A" |
815 |
815 |
67.73 |
|
|
|
| 14 |
A" |
699 |
699 |
4.81 |
|
|
|
| 15 |
A" |
495 |
495 |
0.18 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12274.7 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 12274.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP=FULL/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.448 |
0.000 |
| C2 |
1.128 |
-0.385 |
0.000 |
| N3 |
-1.256 |
0.070 |
0.000 |
| H4 |
0.169 |
1.524 |
0.000 |
| H5 |
2.129 |
0.023 |
0.000 |
| H6 |
1.019 |
-1.462 |
0.000 |
| H7 |
-1.293 |
-0.955 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
| C1 | | 1.4021 | 1.3118 | 1.0894 | 2.1715 | 2.1653 | 1.9082 |
C2 | 1.4021 | | 2.4270 | 2.1360 | 1.0811 | 1.0834 | 2.4874 | N3 | 1.3118 | 2.4270 | | 2.0364 | 3.3859 | 2.7430 | 1.0258 | H4 | 1.0894 | 2.1360 | 2.0364 | | 2.4691 | 3.1052 | 2.8786 | H5 | 2.1715 | 1.0811 | 3.3859 | 2.4691 | | 1.8544 | 3.5595 | H6 | 2.1653 | 1.0834 | 2.7430 | 3.1052 | 1.8544 | | 2.3670 | H7 | 1.9082 | 2.4874 | 1.0258 | 2.8786 | 3.5595 | 2.3670 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H5 |
121.436 |
|
C1 |
C2 |
H6 |
120.658 |
| C1 |
N3 |
H7 |
108.820 |
|
C2 |
C1 |
N3 |
126.810 |
| C2 |
C1 |
H4 |
117.489 |
|
N3 |
C1 |
H4 |
115.701 |
| H5 |
C2 |
H6 |
117.906 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
