Jump to
S1C2
Energy calculated at CCD/6-31G**
| | hartrees |
|---|
| Energy at 0K | -132.902936 |
| Energy at 298.15K | -132.907165 |
| HF Energy | -132.473960 |
| Nuclear repulsion energy | 65.079981 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3543 |
3322 |
2.40 |
|
|
|
| 2 |
A' |
3370 |
3160 |
4.62 |
|
|
|
| 3 |
A' |
3255 |
3052 |
4.86 |
|
|
|
| 4 |
A' |
3162 |
2965 |
39.48 |
|
|
|
| 5 |
A' |
1565 |
1468 |
13.35 |
|
|
|
| 6 |
A' |
1504 |
1410 |
11.92 |
|
|
|
| 7 |
A' |
1402 |
1315 |
16.09 |
|
|
|
| 8 |
A' |
1250 |
1172 |
28.68 |
|
|
|
| 9 |
A' |
1102 |
1033 |
6.14 |
|
|
|
| 10 |
A' |
1031 |
966 |
9.31 |
|
|
|
| 11 |
A' |
501 |
470 |
12.59 |
|
|
|
| 12 |
A" |
1109 |
1040 |
0.45 |
|
|
|
| 13 |
A" |
810 |
759 |
3.79 |
|
|
|
| 14 |
A" |
725 |
680 |
115.61 |
|
|
|
| 15 |
A" |
550 |
516 |
3.12 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12439.1 cm
-1
Scaled (by 0.9376) Zero Point Vibrational Energy (zpe) 11662.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCD/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.419 |
0.000 |
| C2 |
1.147 |
-0.381 |
0.000 |
| N3 |
-1.186 |
-0.132 |
0.000 |
| H4 |
0.141 |
1.501 |
0.000 |
| H5 |
2.137 |
0.049 |
0.000 |
| H6 |
1.042 |
-1.455 |
0.000 |
| H7 |
-1.900 |
0.597 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
| C1 | | 1.3989 | 1.3080 | 1.0909 | 2.1685 | 2.1450 | 1.9084 |
C2 | 1.3989 | | 2.3467 | 2.1346 | 1.0790 | 1.0793 | 3.2006 | N3 | 1.3080 | 2.3467 | | 2.1041 | 3.3278 | 2.5919 | 1.0205 | H4 | 1.0909 | 2.1346 | 2.1041 | | 2.4683 | 3.0909 | 2.2319 | H5 | 2.1685 | 1.0790 | 3.3278 | 2.4683 | | 1.8604 | 4.0739 | H6 | 2.1450 | 1.0793 | 2.5919 | 3.0909 | 1.8604 | | 3.5877 | H7 | 1.9084 | 3.2006 | 1.0205 | 2.2319 | 4.0739 | 3.5877 | |
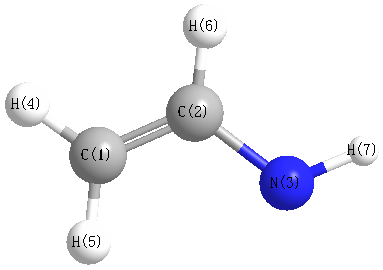 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H5 |
121.588 |
|
C1 |
C2 |
H6 |
119.331 |
| C1 |
N3 |
H7 |
109.467 |
|
C2 |
C1 |
N3 |
120.170 |
| C2 |
C1 |
H4 |
117.502 |
|
N3 |
C1 |
H4 |
122.328 |
| H5 |
C2 |
H6 |
119.081 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CCD/6-31G**
| | hartrees |
|---|
| Energy at 0K | -132.902210 |
| Energy at 298.15K | -132.906434 |
| HF Energy | -132.473351 |
| Nuclear repulsion energy | 65.070267 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3498 |
3280 |
10.13 |
|
|
|
| 2 |
A' |
3352 |
3143 |
7.84 |
|
|
|
| 3 |
A' |
3241 |
3039 |
3.50 |
|
|
|
| 4 |
A' |
3219 |
3018 |
22.87 |
|
|
|
| 5 |
A' |
1560 |
1463 |
4.49 |
|
|
|
| 6 |
A' |
1492 |
1399 |
1.54 |
|
|
|
| 7 |
A' |
1403 |
1316 |
32.35 |
|
|
|
| 8 |
A' |
1243 |
1165 |
30.80 |
|
|
|
| 9 |
A' |
1125 |
1055 |
26.56 |
|
|
|
| 10 |
A' |
1022 |
958 |
0.37 |
|
|
|
| 11 |
A' |
507 |
476 |
9.61 |
|
|
|
| 12 |
A" |
1137 |
1066 |
49.68 |
|
|
|
| 13 |
A" |
820 |
769 |
70.91 |
|
|
|
| 14 |
A" |
724 |
679 |
2.34 |
|
|
|
| 15 |
A" |
526 |
493 |
0.19 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12434.3 cm
-1
Scaled (by 0.9376) Zero Point Vibrational Energy (zpe) 11658.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCD/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.450 |
0.000 |
| C2 |
1.124 |
-0.385 |
0.000 |
| N3 |
-1.252 |
0.068 |
0.000 |
| H4 |
0.169 |
1.524 |
0.000 |
| H5 |
2.124 |
0.020 |
0.000 |
| H6 |
1.014 |
-1.461 |
0.000 |
| H7 |
-1.286 |
-0.955 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
| C1 | | 1.3999 | 1.3088 | 1.0869 | 2.1673 | 2.1631 | 1.9045 |
C2 | 1.3999 | | 2.4184 | 2.1342 | 1.0792 | 1.0814 | 2.4761 | N3 | 1.3088 | 2.4184 | | 2.0337 | 3.3764 | 2.7336 | 1.0238 | H4 | 1.0869 | 2.1342 | 2.0337 | | 2.4669 | 3.1019 | 2.8739 | H5 | 2.1673 | 1.0792 | 3.3764 | 2.4669 | | 1.8506 | 3.5466 | H6 | 2.1631 | 1.0814 | 2.7336 | 3.1019 | 1.8506 | | 2.3549 | H7 | 1.9045 | 2.4761 | 1.0238 | 2.8739 | 3.5466 | 2.3549 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H5 |
121.361 |
|
C1 |
C2 |
H6 |
120.790 |
| C1 |
N3 |
H7 |
108.854 |
|
C2 |
C1 |
N3 |
126.436 |
| C2 |
C1 |
H4 |
117.684 |
|
N3 |
C1 |
H4 |
115.880 |
| H5 |
C2 |
H6 |
117.849 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
