Jump to
S1C2
Energy calculated at TPSSh/6-31G**
| | hartrees |
|---|
| Energy at 0K | -133.312189 |
| Energy at 298.15K | -133.316343 |
| HF Energy | -133.312189 |
| Nuclear repulsion energy | 64.693510 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at TPSSh/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3421 |
3280 |
4.24 |
|
|
|
| 2 |
A' |
3280 |
3145 |
7.40 |
|
|
|
| 3 |
A' |
3171 |
3040 |
5.32 |
|
|
|
| 4 |
A' |
3053 |
2927 |
56.43 |
|
|
|
| 5 |
A' |
1530 |
1467 |
6.69 |
|
|
|
| 6 |
A' |
1472 |
1411 |
2.22 |
|
|
|
| 7 |
A' |
1360 |
1304 |
35.03 |
|
|
|
| 8 |
A' |
1258 |
1206 |
6.18 |
|
|
|
| 9 |
A' |
1117 |
1071 |
22.10 |
|
|
|
| 10 |
A' |
996 |
955 |
11.16 |
|
|
|
| 11 |
A' |
477 |
457 |
11.03 |
|
|
|
| 12 |
A" |
1046 |
1003 |
0.06 |
|
|
|
| 13 |
A" |
802 |
769 |
10.22 |
|
|
|
| 14 |
A" |
675 |
648 |
106.23 |
|
|
|
| 15 |
A" |
529 |
507 |
5.51 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12094.4 cm
-1
Scaled (by 0.9587) Zero Point Vibrational Energy (zpe) 11594.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at TPSSh/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.421 |
0.000 |
| C2 |
1.155 |
-0.374 |
0.000 |
| N3 |
-1.195 |
-0.142 |
0.000 |
| H4 |
0.140 |
1.509 |
0.000 |
| H5 |
2.146 |
0.067 |
0.000 |
| H6 |
1.059 |
-1.454 |
0.000 |
| H7 |
-1.913 |
0.591 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
| C1 | | 1.4022 | 1.3206 | 1.0971 | 2.1753 | 2.1536 | 1.9208 |
C2 | 1.4022 | | 2.3613 | 2.1394 | 1.0844 | 1.0848 | 3.2167 | N3 | 1.3206 | 2.3613 | | 2.1227 | 3.3474 | 2.6080 | 1.0267 | H4 | 1.0971 | 2.1394 | 2.1227 | | 2.4714 | 3.1029 | 2.2486 | H5 | 2.1753 | 1.0844 | 3.3474 | 2.4714 | | 1.8696 | 4.0932 | H6 | 2.1536 | 1.0848 | 2.6080 | 3.1029 | 1.8696 | | 3.6083 | H7 | 1.9208 | 3.2167 | 1.0267 | 2.2486 | 4.0932 | 3.6083 | |
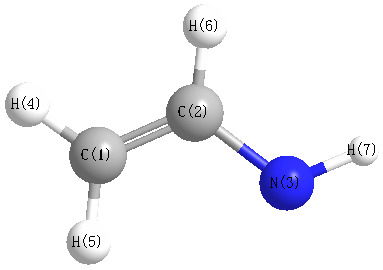 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H5 |
121.514 |
|
C1 |
C2 |
H6 |
119.439 |
| C1 |
N3 |
H7 |
109.191 |
|
C2 |
C1 |
N3 |
120.249 |
| C2 |
C1 |
H4 |
117.219 |
|
N3 |
C1 |
H4 |
122.531 |
| H5 |
C2 |
H6 |
119.047 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at TPSSh/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.076 |
|
|
|
| 2 |
C |
-0.232 |
|
|
|
| 3 |
N |
-0.480 |
|
|
|
| 4 |
H |
0.107 |
|
|
|
| 5 |
H |
0.133 |
|
|
|
| 6 |
H |
0.150 |
|
|
|
| 7 |
H |
0.244 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.043 |
1.571 |
0.000 |
1.886 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.448 |
-0.438 |
0.000 |
| y |
-0.438 |
4.116 |
0.000 |
| z |
0.000 |
0.000 |
1.800 |
<r2> (average value of r
2) Å
2
| <r2> |
45.393 |
| (<r2>)1/2 |
6.737 |
Jump to
S1C1
Energy calculated at TPSSh/6-31G**
| | hartrees |
|---|
| Energy at 0K | -133.311941 |
| Energy at 298.15K | -133.316072 |
| HF Energy | -133.311941 |
| Nuclear repulsion energy | 64.674573 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at TPSSh/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3370 |
3231 |
16.43 |
|
|
|
| 2 |
A' |
3258 |
3123 |
12.93 |
|
|
|
| 3 |
A' |
3153 |
3023 |
6.03 |
|
|
|
| 4 |
A' |
3125 |
2996 |
27.46 |
|
|
|
| 5 |
A' |
1520 |
1457 |
3.81 |
|
|
|
| 6 |
A' |
1464 |
1403 |
3.81 |
|
|
|
| 7 |
A' |
1388 |
1331 |
15.09 |
|
|
|
| 8 |
A' |
1243 |
1192 |
29.30 |
|
|
|
| 9 |
A' |
1143 |
1096 |
31.37 |
|
|
|
| 10 |
A' |
988 |
947 |
0.52 |
|
|
|
| 11 |
A' |
482 |
462 |
8.43 |
|
|
|
| 12 |
A" |
1083 |
1038 |
47.39 |
|
|
|
| 13 |
A" |
808 |
775 |
63.89 |
|
|
|
| 14 |
A" |
678 |
650 |
8.08 |
|
|
|
| 15 |
A" |
480 |
460 |
0.41 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12091.1 cm
-1
Scaled (by 0.9587) Zero Point Vibrational Energy (zpe) 11591.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at TPSSh/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.448 |
0.000 |
| C2 |
1.134 |
-0.380 |
0.000 |
| N3 |
-1.263 |
0.065 |
0.000 |
| H4 |
0.163 |
1.528 |
0.000 |
| H5 |
2.137 |
0.033 |
0.000 |
| H6 |
1.032 |
-1.462 |
0.000 |
| H7 |
-1.295 |
-0.965 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
| C1 | | 1.4040 | 1.3197 | 1.0925 | 2.1773 | 2.1711 | 1.9163 |
C2 | 1.4040 | | 2.4377 | 2.1408 | 1.0851 | 1.0873 | 2.4978 | N3 | 1.3197 | 2.4377 | | 2.0432 | 3.4005 | 2.7565 | 1.0306 | H4 | 1.0925 | 2.1408 | 2.0432 | | 2.4765 | 3.1141 | 2.8880 | H5 | 2.1773 | 1.0851 | 3.4005 | 2.4765 | | 1.8598 | 3.5740 | H6 | 2.1711 | 1.0873 | 2.7565 | 3.1141 | 1.8598 | | 2.3787 | H7 | 1.9163 | 2.4978 | 1.0306 | 2.8880 | 3.5740 | 2.3787 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H5 |
121.501 |
|
C1 |
C2 |
H6 |
120.741 |
| C1 |
N3 |
H7 |
108.621 |
|
C2 |
C1 |
N3 |
126.997 |
| C2 |
C1 |
H4 |
117.543 |
|
N3 |
C1 |
H4 |
115.460 |
| H5 |
C2 |
H6 |
117.758 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at TPSSh/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.093 |
|
|
|
| 2 |
C |
-0.259 |
|
|
|
| 3 |
N |
-0.473 |
|
|
|
| 4 |
H |
0.131 |
|
|
|
| 5 |
H |
0.139 |
|
|
|
| 6 |
H |
0.134 |
|
|
|
| 7 |
H |
0.235 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.121 |
-1.068 |
0.000 |
2.375 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.102 |
-0.070 |
0.000 |
| y |
-0.070 |
4.310 |
0.000 |
| z |
0.000 |
0.000 |
1.804 |
<r2> (average value of r
2) Å
2
| <r2> |
45.448 |
| (<r2>)1/2 |
6.742 |
