Jump to
S1C2
Energy calculated at MP2=FULL/6-31G**
| | hartrees |
|---|
| Energy at 0K | -168.134210 |
| Energy at 298.15K | |
| HF Energy | -167.624855 |
| Nuclear repulsion energy | 59.803883 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Σ |
3590 |
3355 |
264.30 |
34.25 |
0.23 |
0.38 |
| 2 |
Σ |
2291 |
2141 |
419.05 |
2.63 |
0.05 |
0.09 |
| 3 |
Σ |
1332 |
1245 |
53.21 |
31.10 |
0.28 |
0.44 |
| 4 |
Π |
562 |
525 |
0.95 |
0.18 |
0.75 |
0.86 |
| 4 |
Π |
562 |
525 |
0.95 |
0.18 |
0.75 |
0.86 |
| 5 |
Π |
347i |
324i |
87.85 |
1.89 |
0.75 |
0.86 |
| 5 |
Π |
347i |
324i |
87.85 |
1.89 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 3821.6 cm
-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 3570.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31G**
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
0.000 |
0.000 |
1.193 |
| N2 |
0.000 |
0.000 |
-0.014 |
| C3 |
0.000 |
0.000 |
-1.199 |
| H4 |
0.000 |
0.000 |
-2.256 |
Atom - Atom Distances (Å)
| |
O1 |
N2 |
C3 |
H4 |
| O1 | | 1.2074 | 2.3921 | 3.4497 |
N2 | 1.2074 | | 1.1847 | 2.2423 | C3 | 2.3921 | 1.1847 | | 1.0576 | H4 | 3.4497 | 2.2423 | 1.0576 | |
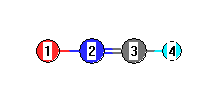 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
N2 |
C3 |
180.000 |
|
N2 |
C3 |
H4 |
180.000 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/6-31G**
| | hartrees |
|---|
| Energy at 0K | -168.135231 |
| Energy at 298.15K | -168.136074 |
| HF Energy | -167.619052 |
| Nuclear repulsion energy | 59.736554 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3536 |
3304 |
165.73 |
|
|
|
| 2 |
A' |
2281 |
2132 |
385.79 |
|
|
|
| 3 |
A' |
1331 |
1244 |
31.34 |
|
|
|
| 4 |
A' |
581 |
543 |
73.94 |
|
|
|
| 5 |
A' |
496 |
463 |
60.11 |
|
|
|
| 6 |
A" |
580 |
542 |
1.57 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 4402.8 cm
-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 4114.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
-1.078 |
-0.501 |
0.000 |
| N2 |
0.000 |
0.036 |
0.000 |
| C3 |
1.132 |
0.422 |
0.000 |
| H4 |
1.833 |
1.219 |
0.000 |
Atom - Atom Distances (Å)
| |
O1 |
N2 |
C3 |
H4 |
| O1 | | 1.2042 | 2.3950 | 3.3814 |
N2 | 1.2042 | | 1.1961 | 2.1821 | C3 | 2.3950 | 1.1961 | | 1.0617 | H4 | 3.3814 | 2.1821 | 1.0617 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
N2 |
C3 |
172.402 |
|
N2 |
C3 |
H4 |
150.187 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
