Jump to
S1C2
Energy calculated at PBE1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -151.746470 |
| Energy at 298.15K | |
| HF Energy | -151.746470 |
| Nuclear repulsion energy | 52.377892 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBE1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Σ |
3518 |
3353 |
113.62 |
46.40 |
0.23 |
0.37 |
| 2 |
Σ |
2154 |
2053 |
191.55 |
2.91 |
0.42 |
0.60 |
| 3 |
Σ |
1334 |
1272 |
22.56 |
30.95 |
0.25 |
0.40 |
| 4 |
Π |
568 |
541 |
0.32 |
1.83 |
0.75 |
0.86 |
| 4 |
Π |
521 |
496 |
10.99 |
1.30 |
0.75 |
0.86 |
| 5 |
Π |
420 |
400 |
26.63 |
0.02 |
0.75 |
0.86 |
| 5 |
Π |
352i |
336i |
144.86 |
4.13 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 4081.2 cm
-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 3889.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBE1PBE/6-31G**
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.019 |
| C2 |
0.000 |
0.000 |
-1.244 |
| O3 |
0.000 |
0.000 |
1.207 |
| H4 |
0.000 |
0.000 |
-2.309 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
H4 |
| C1 | | 1.2628 | 1.1881 | 2.3281 |
C2 | 1.2628 | | 2.4508 | 1.0653 | O3 | 1.1881 | 2.4508 | | 3.5161 | H4 | 2.3281 | 1.0653 | 3.5161 | |
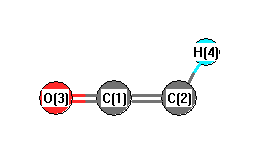 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H4 |
180.000 |
|
C2 |
C1 |
O3 |
180.000 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.300 |
|
|
|
| 2 |
C |
-0.173 |
|
|
|
| 3 |
O |
-0.383 |
|
|
|
| 4 |
H |
0.256 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-2.149 |
2.149 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-17.425 |
0.000 |
0.000 |
| y |
0.000 |
-15.792 |
0.000 |
| z |
0.000 |
0.000 |
-14.302 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.378 |
0.000 |
0.000 |
| y |
0.000 |
0.071 |
0.000 |
| z |
0.000 |
0.000 |
2.307 |
|
| Polar |
|---|
| 3z2-r2 | 4.614 |
|---|
| x2-y2 | -1.633 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
1.428 |
0.000 |
0.000 |
| y |
0.000 |
1.424 |
0.000 |
| z |
0.000 |
0.000 |
5.369 |
<r2> (average value of r
2) Å
2
| <r2> |
36.165 |
| (<r2>)1/2 |
6.014 |
Jump to
S1C1
Energy calculated at PBE1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -151.748484 |
| Energy at 298.15K | |
| HF Energy | -151.748484 |
| Nuclear repulsion energy | 52.262754 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBE1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3406 |
3246 |
42.13 |
73.68 |
0.30 |
0.46 |
| 2 |
A' |
2140 |
2040 |
274.46 |
3.75 |
0.26 |
0.41 |
| 3 |
A' |
1290 |
1230 |
7.13 |
27.79 |
0.31 |
0.47 |
| 4 |
A' |
551 |
525 |
7.23 |
4.45 |
0.75 |
0.85 |
| 5 |
A' |
487 |
464 |
233.34 |
7.72 |
0.19 |
0.32 |
| 6 |
A" |
499 |
476 |
4.21 |
1.11 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 4186.4 cm
-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 3990.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBE1PBE/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.049 |
0.000 |
| C2 |
1.091 |
-0.644 |
0.000 |
| O3 |
-1.087 |
0.504 |
0.000 |
| H4 |
2.149 |
-0.461 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
H4 |
| C1 | | 1.2926 | 1.1781 | 2.2089 |
C2 | 1.2926 | | 2.4618 | 1.0741 | O3 | 1.1781 | 2.4618 | | 3.3768 | H4 | 2.2089 | 1.0741 | 3.3768 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H4 |
137.724 |
|
C2 |
C1 |
O3 |
170.265 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.405 |
|
|
|
| 2 |
C |
-0.285 |
|
|
|
| 3 |
O |
-0.355 |
|
|
|
| 4 |
H |
0.234 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.739 |
0.424 |
0.000 |
1.790 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-14.678 |
0.985 |
0.000 |
| y |
0.985 |
-18.588 |
0.000 |
| z |
0.000 |
0.000 |
-15.895 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.563 |
0.985 |
0.000 |
| y |
0.985 |
-3.301 |
0.000 |
| z |
0.000 |
0.000 |
0.738 |
|
| Polar |
|---|
| 3z2-r2 | 1.477 |
|---|
| x2-y2 | 3.909 |
|---|
| xy | 0.985 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.636 |
-1.603 |
0.000 |
| y |
-1.603 |
2.596 |
0.000 |
| z |
0.000 |
0.000 |
1.897 |
<r2> (average value of r
2) Å
2
| <r2> |
36.192 |
| (<r2>)1/2 |
6.016 |
