Jump to
S1C2
Vibrational Frequencies calculated at PBE1PBE/6-31G**
Geometric Data calculated at PBE1PBE/6-31G**
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at PBE1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -269.031252 |
| Energy at 298.15K | -269.037898 |
| HF Energy | -269.031252 |
| Nuclear repulsion energy | 222.998265 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBE1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3327 |
3171 |
1.18 |
|
|
|
| 2 |
A' |
3301 |
3146 |
1.19 |
|
|
|
| 3 |
A' |
3289 |
3135 |
4.47 |
|
|
|
| 4 |
A' |
3177 |
3028 |
9.07 |
|
|
|
| 5 |
A' |
3071 |
2927 |
27.12 |
|
|
|
| 6 |
A' |
1682 |
1603 |
14.24 |
|
|
|
| 7 |
A' |
1584 |
1510 |
18.20 |
|
|
|
| 8 |
A' |
1519 |
1448 |
7.77 |
|
|
|
| 9 |
A' |
1450 |
1382 |
5.85 |
|
|
|
| 10 |
A' |
1437 |
1369 |
0.57 |
|
|
|
| 11 |
A' |
1292 |
1231 |
18.85 |
|
|
|
| 12 |
A' |
1270 |
1211 |
0.74 |
|
|
|
| 13 |
A' |
1210 |
1153 |
17.31 |
|
|
|
| 14 |
A' |
1141 |
1088 |
3.96 |
|
|
|
| 15 |
A' |
1053 |
1004 |
23.49 |
|
|
|
| 16 |
A' |
1005 |
958 |
2.03 |
|
|
|
| 17 |
A' |
959 |
914 |
15.06 |
|
|
|
| 18 |
A' |
905 |
862 |
3.30 |
|
|
|
| 19 |
A' |
669 |
638 |
0.89 |
|
|
|
| 20 |
A' |
336 |
321 |
2.09 |
|
|
|
| 21 |
A" |
3133 |
2986 |
13.47 |
|
|
|
| 22 |
A" |
1500 |
1429 |
7.86 |
|
|
|
| 23 |
A" |
1077 |
1027 |
3.39 |
|
|
|
| 24 |
A" |
868 |
828 |
0.13 |
|
|
|
| 25 |
A" |
811 |
773 |
23.14 |
|
|
|
| 26 |
A" |
743 |
708 |
44.94 |
|
|
|
| 27 |
A" |
646 |
616 |
2.30 |
|
|
|
| 28 |
A" |
620 |
591 |
5.39 |
|
|
|
| 29 |
A" |
244 |
232 |
5.15 |
|
|
|
| 30 |
A" |
121 |
115 |
0.08 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21719.9 cm
-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 20701.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBE1PBE/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
1.033 |
-0.241 |
0.000 |
| C2 |
0.000 |
0.644 |
0.000 |
| C3 |
0.499 |
-1.488 |
0.000 |
| C4 |
-1.182 |
-0.032 |
0.000 |
| C5 |
-0.856 |
-1.425 |
0.000 |
| C6 |
0.356 |
2.083 |
0.000 |
| H7 |
1.208 |
-2.302 |
0.000 |
| H8 |
-2.169 |
0.411 |
0.000 |
| H9 |
-1.541 |
-2.261 |
0.000 |
| H10 |
-0.553 |
2.690 |
0.000 |
| H11 |
0.947 |
2.351 |
0.884 |
| H12 |
0.947 |
2.351 |
-0.884 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
| O1 | | 1.3602 | 1.3565 | 2.2247 | 2.2292 | 2.4213 | 2.0681 | 3.2673 | 3.2721 | 3.3327 | 2.7407 | 2.7407 |
C2 | 1.3602 | | 2.1896 | 1.3615 | 2.2387 | 1.4830 | 3.1839 | 2.1811 | 3.2885 | 2.1195 | 2.1434 | 2.1434 | C3 | 1.3565 | 2.1896 | | 2.2240 | 1.3566 | 3.5745 | 1.0796 | 3.2745 | 2.1817 | 4.3084 | 3.9656 | 3.9656 | C4 | 2.2247 | 1.3615 | 2.2240 | | 1.4304 | 2.6154 | 3.2965 | 1.0815 | 2.2581 | 2.7934 | 3.3156 | 3.3156 | C5 | 2.2292 | 2.2387 | 1.3566 | 1.4304 | | 3.7117 | 2.2433 | 2.2566 | 1.0812 | 4.1257 | 4.2769 | 4.2769 | C6 | 2.4213 | 1.4830 | 3.5745 | 2.6154 | 3.7117 | | 4.4674 | 3.0284 | 4.7409 | 1.0929 | 1.0965 | 1.0965 | H7 | 2.0681 | 3.1839 | 1.0796 | 3.2965 | 2.2433 | 4.4674 | | 4.3318 | 2.7500 | 5.2934 | 4.7439 | 4.7439 | H8 | 3.2673 | 2.1811 | 3.2745 | 1.0815 | 2.2566 | 3.0284 | 4.3318 | | 2.7448 | 2.7933 | 3.7752 | 3.7752 | H9 | 3.2721 | 3.2885 | 2.1817 | 2.2581 | 1.0812 | 4.7409 | 2.7500 | 2.7448 | | 5.0486 | 5.3150 | 5.3150 | H10 | 3.3327 | 2.1195 | 4.3084 | 2.7934 | 4.1257 | 1.0929 | 5.2934 | 2.7933 | 5.0486 | | 1.7736 | 1.7736 | H11 | 2.7407 | 2.1434 | 3.9656 | 3.3156 | 4.2769 | 1.0965 | 4.7439 | 3.7752 | 5.3150 | 1.7736 | | 1.7681 | H12 | 2.7407 | 2.1434 | 3.9656 | 3.3156 | 4.2769 | 1.0965 | 4.7439 | 3.7752 | 5.3150 | 1.7736 | 1.7681 | |
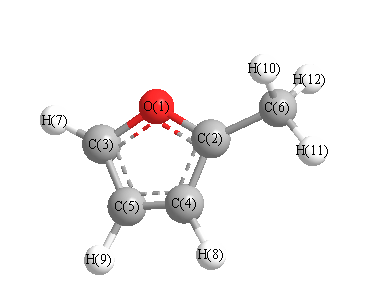 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C4 |
109.653 |
|
O1 |
C2 |
C6 |
116.703 |
| O1 |
C3 |
C5 |
110.502 |
|
O1 |
C3 |
H7 |
115.725 |
| C2 |
O1 |
C3 |
107.409 |
|
C2 |
C4 |
C5 |
106.591 |
| C2 |
C4 |
H8 |
126.072 |
|
C2 |
C6 |
H10 |
109.808 |
| C2 |
C6 |
H11 |
111.511 |
|
C2 |
C6 |
H12 |
111.511 |
| C3 |
C5 |
C6 |
73.619 |
|
C3 |
C5 |
H9 |
126.635 |
| C4 |
C2 |
C6 |
133.645 |
|
C4 |
C5 |
H9 |
127.519 |
| C5 |
C3 |
H7 |
133.773 |
|
C5 |
C4 |
H8 |
127.337 |
| H10 |
C6 |
H11 |
108.210 |
|
H10 |
C6 |
H12 |
108.210 |
| H11 |
C6 |
H12 |
107.462 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.433 |
|
|
|
| 2 |
C |
0.334 |
|
|
|
| 3 |
C |
0.056 |
|
|
|
| 4 |
C |
-0.247 |
|
|
|
| 5 |
C |
-0.232 |
|
|
|
| 6 |
C |
-0.600 |
|
|
|
| 7 |
H |
0.185 |
|
|
|
| 8 |
H |
0.171 |
|
|
|
| 9 |
H |
0.176 |
|
|
|
| 10 |
H |
0.193 |
|
|
|
| 11 |
H |
0.198 |
|
|
|
| 12 |
H |
0.198 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.431 |
0.317 |
0.000 |
0.535 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-33.591 |
-0.489 |
0.000 |
| y |
-0.489 |
-29.557 |
0.000 |
| z |
0.000 |
0.000 |
-37.301 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.162 |
-0.489 |
0.000 |
| y |
-0.489 |
5.889 |
0.000 |
| z |
0.000 |
0.000 |
-5.727 |
|
| Polar |
|---|
| 3z2-r2 | -11.454 |
|---|
| x2-y2 | -4.034 |
|---|
| xy | -0.489 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.079 |
0.350 |
0.000 |
| y |
0.350 |
9.731 |
0.000 |
| z |
0.000 |
0.000 |
4.012 |
<r2> (average value of r
2) Å
2
| <r2> |
140.029 |
| (<r2>)1/2 |
11.833 |
