Jump to
S1C2
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
Geometric Data calculated at HSEh1PBE/6-31G**
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at HSEh1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -269.062352 |
| Energy at 298.15K | -269.068990 |
| HF Energy | -269.062352 |
| Nuclear repulsion energy | 223.028193 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3321 |
3169 |
0.87 |
|
|
|
| 2 |
A' |
3296 |
3146 |
0.97 |
|
|
|
| 3 |
A' |
3285 |
3134 |
4.46 |
|
|
|
| 4 |
A' |
3174 |
3029 |
8.91 |
|
|
|
| 5 |
A' |
3063 |
2923 |
27.38 |
|
|
|
| 6 |
A' |
1679 |
1602 |
14.52 |
|
|
|
| 7 |
A' |
1581 |
1509 |
19.13 |
|
|
|
| 8 |
A' |
1503 |
1434 |
6.11 |
|
|
|
| 9 |
A' |
1442 |
1376 |
5.53 |
|
|
|
| 10 |
A' |
1421 |
1356 |
1.22 |
|
|
|
| 11 |
A' |
1285 |
1226 |
18.60 |
|
|
|
| 12 |
A' |
1265 |
1208 |
0.71 |
|
|
|
| 13 |
A' |
1205 |
1150 |
16.80 |
|
|
|
| 14 |
A' |
1137 |
1085 |
4.13 |
|
|
|
| 15 |
A' |
1049 |
1002 |
23.86 |
|
|
|
| 16 |
A' |
999 |
953 |
2.07 |
|
|
|
| 17 |
A' |
956 |
912 |
14.80 |
|
|
|
| 18 |
A' |
904 |
863 |
3.21 |
|
|
|
| 19 |
A' |
668 |
638 |
0.86 |
|
|
|
| 20 |
A' |
336 |
321 |
2.10 |
|
|
|
| 21 |
A" |
3128 |
2985 |
13.61 |
|
|
|
| 22 |
A" |
1481 |
1414 |
7.27 |
|
|
|
| 23 |
A" |
1066 |
1017 |
3.30 |
|
|
|
| 24 |
A" |
874 |
834 |
0.09 |
|
|
|
| 25 |
A" |
814 |
777 |
20.54 |
|
|
|
| 26 |
A" |
745 |
711 |
44.47 |
|
|
|
| 27 |
A" |
645 |
616 |
2.04 |
|
|
|
| 28 |
A" |
620 |
592 |
5.31 |
|
|
|
| 29 |
A" |
244 |
233 |
5.09 |
|
|
|
| 30 |
A" |
119 |
114 |
0.10 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21652.2 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 20662.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
1.033 |
-0.241 |
0.000 |
| C2 |
0.000 |
0.644 |
0.000 |
| C3 |
0.499 |
-1.489 |
0.000 |
| C4 |
-1.182 |
-0.031 |
0.000 |
| C5 |
-0.856 |
-1.424 |
0.000 |
| C6 |
0.356 |
2.083 |
0.000 |
| H7 |
1.205 |
-2.304 |
0.000 |
| H8 |
-2.167 |
0.412 |
0.000 |
| H9 |
-1.541 |
-2.260 |
0.000 |
| H10 |
-0.553 |
2.688 |
0.000 |
| H11 |
0.946 |
2.350 |
0.883 |
| H12 |
0.946 |
2.350 |
-0.883 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
| O1 | | 1.3606 | 1.3573 | 2.2249 | 2.2291 | 2.4208 | 2.0701 | 3.2664 | 3.2710 | 3.3312 | 2.7391 | 2.7391 |
C2 | 1.3606 | | 2.1905 | 1.3613 | 2.2385 | 1.4820 | 3.1851 | 2.1796 | 3.2875 | 2.1175 | 2.1413 | 2.1413 | C3 | 1.3573 | 2.1905 | | 2.2244 | 1.3564 | 3.5746 | 1.0788 | 3.2741 | 2.1801 | 4.3074 | 3.9646 | 3.9646 | C4 | 2.2249 | 1.3613 | 2.2244 | | 1.4302 | 2.6143 | 3.2959 | 1.0806 | 2.2572 | 2.7916 | 3.3135 | 3.3135 | C5 | 2.2291 | 2.2385 | 1.3564 | 1.4302 | | 3.7105 | 2.2414 | 2.2561 | 1.0803 | 4.1236 | 4.2746 | 4.2746 | C6 | 2.4208 | 1.4820 | 3.5746 | 2.6143 | 3.7105 | | 4.4685 | 3.0261 | 4.7388 | 1.0917 | 1.0954 | 1.0954 | H7 | 2.0701 | 3.1851 | 1.0788 | 3.2959 | 2.2414 | 4.4685 | | 4.3302 | 2.7461 | 5.2929 | 4.7444 | 4.7444 | H8 | 3.2664 | 2.1796 | 3.2741 | 1.0806 | 2.2561 | 3.0261 | 4.3302 | | 2.7444 | 2.7908 | 3.7721 | 3.7721 | H9 | 3.2710 | 3.2875 | 2.1801 | 2.2572 | 1.0803 | 4.7388 | 2.7461 | 2.7444 | | 5.0459 | 5.3119 | 5.3119 | H10 | 3.3312 | 2.1175 | 4.3074 | 2.7916 | 4.1236 | 1.0917 | 5.2929 | 2.7908 | 5.0459 | | 1.7722 | 1.7722 | H11 | 2.7391 | 2.1413 | 3.9646 | 3.3135 | 4.2746 | 1.0954 | 4.7444 | 3.7721 | 5.3119 | 1.7722 | | 1.7665 | H12 | 2.7391 | 2.1413 | 3.9646 | 3.3135 | 4.2746 | 1.0954 | 4.7444 | 3.7721 | 5.3119 | 1.7722 | 1.7665 | |
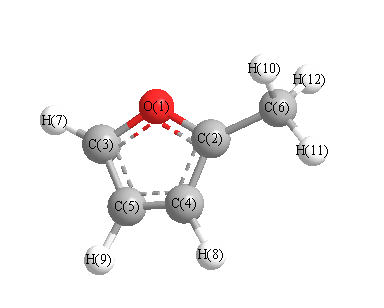 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C4 |
109.650 |
|
O1 |
C2 |
C6 |
116.706 |
| O1 |
C3 |
C5 |
110.454 |
|
O1 |
C3 |
H7 |
115.894 |
| C2 |
O1 |
C3 |
107.403 |
|
C2 |
C4 |
C5 |
106.595 |
| C2 |
C4 |
H8 |
126.020 |
|
C2 |
C6 |
H10 |
109.793 |
| C2 |
C6 |
H11 |
111.479 |
|
C2 |
C6 |
H12 |
111.479 |
| C3 |
C5 |
C6 |
73.673 |
|
C3 |
C5 |
H9 |
126.570 |
| C4 |
C2 |
C6 |
133.645 |
|
C4 |
C5 |
H9 |
127.533 |
| C5 |
C3 |
H7 |
133.652 |
|
C5 |
C4 |
H8 |
127.385 |
| H10 |
C6 |
H11 |
108.247 |
|
H10 |
C6 |
H12 |
108.247 |
| H11 |
C6 |
H12 |
107.473 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.439 |
|
|
|
| 2 |
C |
0.300 |
|
|
|
| 3 |
C |
0.105 |
|
|
|
| 4 |
C |
-0.187 |
|
|
|
| 5 |
C |
-0.182 |
|
|
|
| 6 |
C |
-0.448 |
|
|
|
| 7 |
H |
0.139 |
|
|
|
| 8 |
H |
0.124 |
|
|
|
| 9 |
H |
0.128 |
|
|
|
| 10 |
H |
0.148 |
|
|
|
| 11 |
H |
0.156 |
|
|
|
| 12 |
H |
0.156 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.426 |
0.340 |
0.000 |
0.545 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-33.656 |
-0.464 |
0.000 |
| y |
-0.464 |
-29.539 |
0.000 |
| z |
0.000 |
0.000 |
-37.236 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.268 |
-0.464 |
0.000 |
| y |
-0.464 |
5.906 |
0.000 |
| z |
0.000 |
0.000 |
-5.639 |
|
| Polar |
|---|
| 3z2-r2 | -11.277 |
|---|
| x2-y2 | -4.116 |
|---|
| xy | -0.464 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.158 |
0.347 |
0.000 |
| y |
0.347 |
9.842 |
0.000 |
| z |
0.000 |
0.000 |
4.071 |
<r2> (average value of r
2) Å
2
| <r2> |
139.976 |
| (<r2>)1/2 |
11.831 |
