return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H10O (Methyl propyl ether)

using model chemistry: QCISD/6-31G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2 1A'
Energy calculated at QCISD/6-31G**
 hartrees
Energy at 0K-233.050434
Energy at 298.15K 
HF Energy-232.153936
Nuclear repulsion energy188.670634
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3204 3016 36.22      
2 A' 3189 3002 35.08      
3 A' 3118 2935 22.68      
4 A' 3101 2919 27.33      
5 A' 3056 2876 67.21      
6 A' 3026 2849 37.89      
7 A' 1592 1499 0.92      
8 A' 1568 1476 8.11      
9 A' 1566 1474 2.64      
10 A' 1552 1461 0.77      
11 A' 1530 1441 1.45      
12 A' 1482 1395 29.91      
13 A' 1472 1386 3.13      
14 A' 1377 1296 7.49      
15 A' 1267 1193 49.03      
16 A' 1204 1134 114.67      
17 A' 1160 1092 13.04      
18 A' 1095 1030 2.64      
19 A' 1011 952 20.92      
20 A' 935 880 4.99      
21 A' 448 422 0.68      
22 A' 418 393 3.14      
23 A' 196 185 1.47      
24 A" 3182 2996 66.88      
25 A" 3160 2975 2.46      
26 A" 3112 2930 66.59      
27 A" 3062 2882 59.81      
28 A" 1559 1468 4.95      
29 A" 1547 1456 3.43      
30 A" 1354 1274 0.05      
31 A" 1308 1231 1.25      
32 A" 1239 1167 7.12      
33 A" 1201 1130 0.76      
34 A" 929 874 1.60      
35 A" 783 737 0.52      
36 A" 247 232 1.39      
37 A" 236 222 2.85      
38 A" 116 110 1.68      
39 A" 103 97 1.78      

Unscaled Zero Point Vibrational Energy (zpe) 30851.4 cm-1
Scaled (by 0.9414) Zero Point Vibrational Energy (zpe) 29043.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/6-31G**
ABC
0.66736 0.07076 0.06719

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/6-31G**

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -2.326 0.637 0.000
O2 -1.267 -0.300 0.000
C3 0.000 0.336 0.000
C4 1.074 -0.737 0.000
C5 2.483 -0.146 0.000
H6 -3.255 0.071 0.000
H7 -2.296 1.278 0.889
H8 -2.296 1.278 -0.889
H9 0.104 0.981 0.885
H10 0.104 0.981 -0.885
H11 0.927 -1.371 -0.877
H12 0.927 -1.371 0.877
H13 2.649 0.476 0.882
H14 2.649 0.476 -0.882
H15 3.238 -0.933 0.000

Atom - Atom Distances (Å)
  C1 O2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13 H14 H15
C11.41422.34563.66774.87281.08811.09651.09652.60862.60863.92263.92265.05545.05545.7812
O21.41421.41792.38193.75372.02242.08332.08332.07472.07472.59462.59464.08874.08874.5492
C32.34561.41791.51832.52963.26612.63652.63651.10011.10012.13132.13132.79562.79563.4774
C43.66772.38191.51831.52804.40434.02644.02642.16302.16301.09221.09222.17472.17472.1723
C54.87283.75372.52961.52805.74275.06585.06582.77792.77792.16592.16591.09191.09191.0902
H61.08812.02243.26614.40435.74271.77991.77993.59083.59084.51034.51035.98365.98366.5701
H71.09652.08332.63654.02645.06581.77991.77822.41812.99894.53084.17235.01005.31396.0253
H81.09652.08332.63654.02645.06581.77991.77822.99892.41814.17234.53085.31395.01006.0253
H92.60862.07471.10012.16302.77793.59082.41812.99891.76963.05232.49232.59523.13983.7776
H102.60862.07471.10012.16302.77793.59082.99892.41811.76962.49233.05233.13982.59523.7776
H113.92262.59462.13131.09222.16594.51034.53084.17233.05232.49231.75483.07732.52452.5099
H123.92262.59462.13131.09222.16594.51034.17234.53082.49233.05231.75482.52453.07732.5099
H135.05544.08872.79562.17471.09195.98365.01005.31392.59523.13983.07732.52451.76481.7632
H145.05544.08872.79562.17471.09195.98365.31395.01003.13982.59522.52453.07731.76481.7632
H155.78124.54923.47742.17231.09026.57016.02536.02533.77763.77762.50992.50991.76321.7632

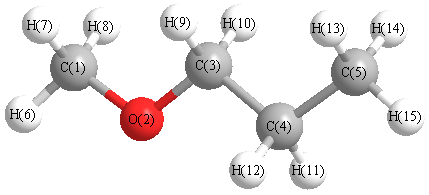
picture of Methyl propyl ether state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 O2 C3 111.835 O2 C1 H6 107.122
O2 C1 H7 111.531 O2 C1 H8 111.531
O2 C3 C4 108.382 O2 C3 H9 110.323
O2 C3 H10 110.323 C3 C4 C5 112.270
C3 C4 H11 108.349 C3 C4 H12 108.349
C4 C3 H9 110.370 C4 C3 H10 110.370
C4 C5 H13 111.112 C4 C5 H14 111.112
C4 C5 H15 111.026 C5 C4 H11 110.398
C5 C4 H12 110.398 H6 C1 H7 109.124
H6 C1 H8 109.124 H7 C1 H8 108.361
H9 C3 H10 107.078 H11 C4 H12 106.898
H13 C5 H14 107.823 H13 C5 H15 107.807
H14 C5 H15 107.807
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability