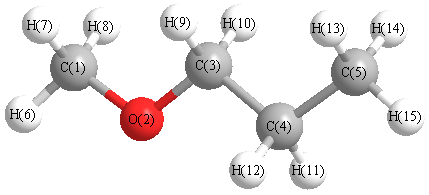
Vibrational Frequencies calculated at PBE1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3160 |
3012 |
32.12 |
|
|
|
| 2 |
A' |
3155 |
3007 |
28.93 |
|
|
|
| 3 |
A' |
3080 |
2936 |
26.40 |
|
|
|
| 4 |
A' |
3068 |
2924 |
23.77 |
|
|
|
| 5 |
A' |
3006 |
2865 |
74.48 |
|
|
|
| 6 |
A' |
2975 |
2836 |
41.27 |
|
|
|
| 7 |
A' |
1560 |
1486 |
1.60 |
|
|
|
| 8 |
A' |
1536 |
1464 |
11.61 |
|
|
|
| 9 |
A' |
1532 |
1460 |
4.71 |
|
|
|
| 10 |
A' |
1519 |
1448 |
1.63 |
|
|
|
| 11 |
A' |
1505 |
1435 |
0.48 |
|
|
|
| 12 |
A' |
1450 |
1382 |
24.82 |
|
|
|
| 13 |
A' |
1439 |
1372 |
3.88 |
|
|
|
| 14 |
A' |
1348 |
1285 |
3.51 |
|
|
|
| 15 |
A' |
1256 |
1197 |
80.98 |
|
|
|
| 16 |
A' |
1213 |
1156 |
107.98 |
|
|
|
| 17 |
A' |
1148 |
1094 |
5.55 |
|
|
|
| 18 |
A' |
1085 |
1034 |
3.56 |
|
|
|
| 19 |
A' |
1023 |
975 |
23.47 |
|
|
|
| 20 |
A' |
924 |
881 |
7.03 |
|
|
|
| 21 |
A' |
445 |
424 |
0.66 |
|
|
|
| 22 |
A' |
411 |
392 |
2.85 |
|
|
|
| 23 |
A' |
194 |
185 |
1.44 |
|
|
|
| 24 |
A" |
3146 |
2999 |
55.33 |
|
|
|
| 25 |
A" |
3120 |
2974 |
3.58 |
|
|
|
| 26 |
A" |
3055 |
2911 |
64.43 |
|
|
|
| 27 |
A" |
3006 |
2865 |
68.72 |
|
|
|
| 28 |
A" |
1529 |
1458 |
8.21 |
|
|
|
| 29 |
A" |
1515 |
1444 |
6.44 |
|
|
|
| 30 |
A" |
1330 |
1268 |
0.14 |
|
|
|
| 31 |
A" |
1285 |
1225 |
2.13 |
|
|
|
| 32 |
A" |
1216 |
1159 |
4.75 |
|
|
|
| 33 |
A" |
1187 |
1131 |
0.06 |
|
|
|
| 34 |
A" |
911 |
868 |
2.14 |
|
|
|
| 35 |
A" |
772 |
736 |
2.13 |
|
|
|
| 36 |
A" |
249 |
237 |
2.58 |
|
|
|
| 37 |
A" |
244 |
232 |
1.13 |
|
|
|
| 38 |
A" |
116 |
111 |
2.61 |
|
|
|
| 39 |
A" |
107 |
102 |
0.75 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30410.3 cm
-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 28984.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.262 |
|
|
|
| 2 |
O |
-0.463 |
|
|
|
| 3 |
C |
-0.049 |
|
|
|
| 4 |
C |
-0.337 |
|
|
|
| 5 |
C |
-0.532 |
|
|
|
| 6 |
H |
0.183 |
|
|
|
| 7 |
H |
0.150 |
|
|
|
| 8 |
H |
0.150 |
|
|
|
| 9 |
H |
0.139 |
|
|
|
| 10 |
H |
0.139 |
|
|
|
| 11 |
H |
0.179 |
|
|
|
| 12 |
H |
0.179 |
|
|
|
| 13 |
H |
0.172 |
|
|
|
| 14 |
H |
0.172 |
|
|
|
| 15 |
H |
0.179 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.237 |
1.064 |
0.000 |
1.090 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-30.746 |
-2.177 |
0.000 |
| y |
-2.177 |
-33.027 |
0.000 |
| z |
0.000 |
0.000 |
-32.779 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.157 |
-2.177 |
0.000 |
| y |
-2.177 |
-1.265 |
0.000 |
| z |
0.000 |
0.000 |
-0.893 |
|
| Polar |
|---|
| 3z2-r2 | -1.785 |
|---|
| x2-y2 | 2.282 |
|---|
| xy | -2.177 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.363 |
-0.386 |
0.000 |
| y |
-0.386 |
6.520 |
0.000 |
| z |
0.000 |
0.000 |
6.366 |
<r2> (average value of r
2) Å
2
| <r2> |
179.723 |
| (<r2>)1/2 |
13.406 |