return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H12 (2,3-dimethylbut-1-ene)

using model chemistry: MP2=FULL/6-31G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2 1A'
Energy calculated at MP2=FULL/6-31G**
 hartrees
Energy at 0K-235.088547
Energy at 298.15K-235.100932
HF Energy-234.195433
Nuclear repulsion energy244.563122
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3330 3111 17.21      
2 A' 3232 3020 19.87      
3 A' 3229 3017 19.71      
4 A' 3225 3013 13.76      
5 A' 3219 3008 61.09      
6 A' 3126 2921 20.51      
7 A' 3122 2917 7.42      
8 A' 3119 2915 22.26      
9 A' 1742 1628 5.11      
10 A' 1572 1469 4.80      
11 A' 1567 1465 7.29      
12 A' 1555 1453 7.34      
13 A' 1503 1405 1.27      
14 A' 1469 1372 2.49      
15 A' 1460 1364 5.51      
16 A' 1385 1294 2.75      
17 A' 1350 1262 0.55      
18 A' 1231 1151 1.79      
19 A' 1142 1067 9.20      
20 A' 1035 967 1.23      
21 A' 1004 939 0.44      
22 A' 935 874 1.36      
23 A' 756 707 1.00      
24 A' 530 495 1.03      
25 A' 447 418 0.84      
26 A' 336 314 0.19      
27 A' 287 268 0.18      
28 A' 273 255 0.25      
29 A" 3225 3013 10.71      
30 A" 3217 3006 5.92      
31 A" 3205 2995 20.12      
32 A" 3122 2917 21.74      
33 A" 1561 1458 0.17      
34 A" 1549 1447 0.81      
35 A" 1537 1437 6.94      
36 A" 1446 1351 3.91      
37 A" 1387 1296 1.81      
38 A" 1171 1094 1.94      
39 A" 1096 1025 0.04      
40 A" 1000 934 0.15      
41 A" 955 892 0.82      
42 A" 915 855 36.73      
43 A" 760 710 0.49      
44 A" 555 519 5.90      
45 A" 278 260 0.01      
46 A" 215 201 0.29      
47 A" 171 160 0.21      
48 A" 22 21 0.09      

Unscaled Zero Point Vibrational Energy (zpe) 37282.9 cm-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 34837.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31G**
ABC
0.14862 0.08396 0.08295

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31G**

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -1.752 -1.061 0.000
C2 -0.472 -0.663 0.000
C3 -0.120 0.806 0.000
C4 0.653 -1.660 0.000
C5 0.653 1.205 1.257
C6 0.653 1.205 -1.257
H7 -2.020 -2.108 0.000
H8 -2.563 -0.346 0.000
H9 -1.062 1.361 0.000
H10 0.267 -2.677 0.000
H11 1.290 -1.542 -0.877
H12 1.290 -1.542 0.877
H13 0.810 2.283 1.282
H14 0.111 0.918 2.156
H15 1.633 0.730 1.284
H16 0.810 2.283 -1.282
H17 0.111 0.918 -2.156
H18 1.633 0.730 -1.284

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16 H17 H18
C11.34012.47972.47883.53553.53551.08101.08092.51782.58663.20203.20204.40293.46874.03854.40293.46874.0385
C21.34011.51071.50362.51742.51742.11772.11472.10782.14562.15562.15563.45912.73622.83163.45912.73622.8316
C32.47971.51072.58441.52891.52893.47902.70141.09333.50452.87582.87582.16542.17132.17372.16542.17132.1737
C42.47881.50362.58443.12883.12882.71103.47433.47371.08771.08961.08964.14913.40402.88404.14913.40402.8840
C53.53552.51741.52893.12882.51454.43903.78582.13244.09873.53642.84571.08941.08871.08902.76323.46832.7642
C63.53552.51741.52893.12882.51454.43903.78582.13244.09872.84573.53642.76323.46832.76421.08941.08871.0890
H71.08102.11773.47902.71104.43904.43901.84333.59852.35773.47073.47075.37894.28304.80055.37894.28304.8005
H81.08092.11472.70143.47433.78583.78581.84332.27313.66674.12844.12844.46453.66004.51784.46453.66004.5178
H92.51782.10781.09333.47372.13242.13243.59852.27314.25093.83753.83752.44902.49413.05082.44902.49413.0508
H102.58662.14563.50451.08774.09874.09872.35773.66674.25091.76091.76095.15154.19453.88805.15154.19453.8880
H113.20202.15562.87581.08963.53642.84573.47074.12843.83751.76091.75304.41854.07903.15363.87653.01312.3335
H123.20202.15562.87581.08962.84573.53643.47074.12843.83751.76091.75303.87653.01312.33354.41854.07903.1536
H134.40293.45912.16544.14911.08942.76325.37894.46452.44905.15154.41853.87651.76541.75782.56453.76503.1101
H143.46872.73622.17133.40401.08873.46834.28303.66002.49414.19454.07903.01311.76541.76463.76504.31243.7661
H154.03852.83162.17372.88401.08902.76424.80054.51783.05083.88803.15362.33351.75781.76463.11013.76612.5670
H164.40293.45912.16544.14912.76321.08945.37894.46452.44905.15153.87654.41852.56453.76503.11011.76541.7578
H173.46872.73622.17133.40403.46831.08874.28303.66002.49414.19453.01314.07903.76504.31243.76611.76541.7646
H184.03852.83162.17372.88402.76421.08904.80054.51783.05083.88802.33353.15363.11013.76612.56701.75781.7646

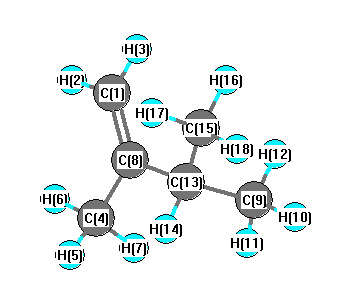
picture of 2,3-dimethylbut-1-ene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 120.758 C1 C2 C4 121.199
C2 C1 H7 121.646 C2 C1 H8 121.358
C2 C3 C5 111.822 C2 C3 C6 111.822
C2 C3 H9 106.991 C2 C4 H10 110.761
C2 C4 H11 111.448 C2 C4 H12 111.448
C3 C2 C4 118.043 C3 C5 H13 110.463
C3 C5 H14 110.979 C3 C5 H15 111.144
C3 C6 H16 110.463 C3 C6 H17 110.979
C3 C6 H18 111.144 C5 C3 C6 110.634
C5 C3 H9 107.662 C6 C3 H9 107.662
H7 C1 H8 116.996 H10 C4 H11 107.948
H10 C4 H12 107.948 H11 C4 H12 107.110
H13 C5 H14 108.296 H13 C5 H15 107.592
H14 C5 H15 108.247 H16 C6 H17 108.296
H16 C6 H18 107.592 H17 C6 H18 108.247
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability