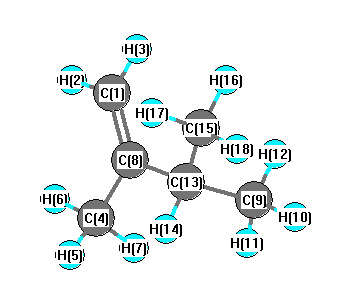
Vibrational Frequencies calculated at PBE1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3257 |
3104 |
19.32 |
|
|
|
| 2 |
A' |
3175 |
3027 |
10.31 |
|
|
|
| 3 |
A' |
3160 |
3012 |
22.27 |
|
|
|
| 4 |
A' |
3154 |
3006 |
23.59 |
|
|
|
| 5 |
A' |
3146 |
2998 |
68.99 |
|
|
|
| 6 |
A' |
3070 |
2926 |
30.55 |
|
|
|
| 7 |
A' |
3064 |
2920 |
1.07 |
|
|
|
| 8 |
A' |
3061 |
2918 |
24.02 |
|
|
|
| 9 |
A' |
1753 |
1670 |
14.96 |
|
|
|
| 10 |
A' |
1540 |
1468 |
9.31 |
|
|
|
| 11 |
A' |
1533 |
1461 |
7.18 |
|
|
|
| 12 |
A' |
1519 |
1448 |
11.77 |
|
|
|
| 13 |
A' |
1474 |
1405 |
1.19 |
|
|
|
| 14 |
A' |
1442 |
1374 |
4.02 |
|
|
|
| 15 |
A' |
1433 |
1366 |
8.72 |
|
|
|
| 16 |
A' |
1360 |
1296 |
3.82 |
|
|
|
| 17 |
A' |
1330 |
1268 |
0.36 |
|
|
|
| 18 |
A' |
1205 |
1149 |
2.52 |
|
|
|
| 19 |
A' |
1124 |
1071 |
13.46 |
|
|
|
| 20 |
A' |
1016 |
969 |
1.77 |
|
|
|
| 21 |
A' |
983 |
937 |
0.61 |
|
|
|
| 22 |
A' |
922 |
879 |
1.86 |
|
|
|
| 23 |
A' |
748 |
712 |
1.34 |
|
|
|
| 24 |
A' |
524 |
499 |
1.44 |
|
|
|
| 25 |
A' |
441 |
420 |
0.82 |
|
|
|
| 26 |
A' |
331 |
315 |
0.25 |
|
|
|
| 27 |
A' |
278 |
265 |
0.47 |
|
|
|
| 28 |
A' |
262 |
250 |
0.09 |
|
|
|
| 29 |
A" |
3152 |
3005 |
17.86 |
|
|
|
| 30 |
A" |
3142 |
2994 |
0.30 |
|
|
|
| 31 |
A" |
3123 |
2976 |
22.46 |
|
|
|
| 32 |
A" |
3065 |
2922 |
23.42 |
|
|
|
| 33 |
A" |
1524 |
1453 |
0.16 |
|
|
|
| 34 |
A" |
1515 |
1444 |
0.90 |
|
|
|
| 35 |
A" |
1501 |
1431 |
10.22 |
|
|
|
| 36 |
A" |
1419 |
1352 |
6.39 |
|
|
|
| 37 |
A" |
1358 |
1295 |
1.67 |
|
|
|
| 38 |
A" |
1148 |
1094 |
1.56 |
|
|
|
| 39 |
A" |
1078 |
1027 |
1.04 |
|
|
|
| 40 |
A" |
979 |
933 |
0.21 |
|
|
|
| 41 |
A" |
939 |
895 |
8.27 |
|
|
|
| 42 |
A" |
926 |
883 |
37.23 |
|
|
|
| 43 |
A" |
749 |
714 |
0.55 |
|
|
|
| 44 |
A" |
556 |
530 |
7.16 |
|
|
|
| 45 |
A" |
263 |
250 |
0.01 |
|
|
|
| 46 |
A" |
215 |
204 |
0.42 |
|
|
|
| 47 |
A" |
165 |
158 |
0.19 |
|
|
|
| 48 |
A" |
39 |
37 |
0.13 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36578.5 cm
-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 34863.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.455 |
|
|
|
| 2 |
C |
0.220 |
|
|
|
| 3 |
C |
-0.203 |
|
|
|
| 4 |
C |
-0.591 |
|
|
|
| 5 |
C |
-0.519 |
|
|
|
| 6 |
C |
-0.519 |
|
|
|
| 7 |
H |
0.158 |
|
|
|
| 8 |
H |
0.159 |
|
|
|
| 9 |
H |
0.165 |
|
|
|
| 10 |
H |
0.181 |
|
|
|
| 11 |
H |
0.183 |
|
|
|
| 12 |
H |
0.183 |
|
|
|
| 13 |
H |
0.171 |
|
|
|
| 14 |
H |
0.179 |
|
|
|
| 15 |
H |
0.168 |
|
|
|
| 16 |
H |
0.171 |
|
|
|
| 17 |
H |
0.179 |
|
|
|
| 18 |
H |
0.168 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.276 |
0.378 |
0.000 |
0.468 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.424 |
-0.674 |
0.000 |
| y |
-0.674 |
-39.072 |
0.000 |
| z |
0.000 |
0.000 |
-40.983 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.603 |
-0.674 |
0.000 |
| y |
-0.674 |
0.631 |
0.000 |
| z |
0.000 |
0.000 |
-2.234 |
|
| Polar |
|---|
| 3z2-r2 | -4.469 |
|---|
| x2-y2 | 0.648 |
|---|
| xy | -0.674 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.598 |
0.802 |
0.000 |
| y |
0.802 |
10.898 |
0.000 |
| z |
0.000 |
0.000 |
7.564 |
<r2> (average value of r
2) Å
2
| <r2> |
190.127 |
| (<r2>)1/2 |
13.789 |