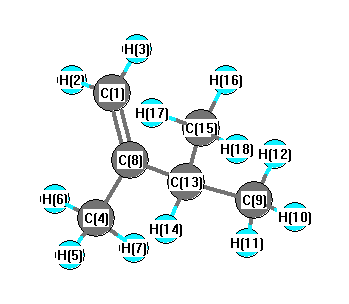
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3253 |
3105 |
18.79 |
|
|
|
| 2 |
A' |
3166 |
3021 |
10.30 |
|
|
|
| 3 |
A' |
3158 |
3013 |
20.92 |
|
|
|
| 4 |
A' |
3150 |
3007 |
24.49 |
|
|
|
| 5 |
A' |
3142 |
2998 |
67.75 |
|
|
|
| 6 |
A' |
3062 |
2922 |
35.31 |
|
|
|
| 7 |
A' |
3060 |
2920 |
3.20 |
|
|
|
| 8 |
A' |
3055 |
2915 |
18.26 |
|
|
|
| 9 |
A' |
1748 |
1668 |
15.64 |
|
|
|
| 10 |
A' |
1523 |
1453 |
9.04 |
|
|
|
| 11 |
A' |
1515 |
1446 |
6.08 |
|
|
|
| 12 |
A' |
1500 |
1432 |
11.50 |
|
|
|
| 13 |
A' |
1459 |
1392 |
1.05 |
|
|
|
| 14 |
A' |
1426 |
1360 |
3.68 |
|
|
|
| 15 |
A' |
1416 |
1351 |
8.96 |
|
|
|
| 16 |
A' |
1348 |
1287 |
3.60 |
|
|
|
| 17 |
A' |
1320 |
1259 |
0.33 |
|
|
|
| 18 |
A' |
1197 |
1142 |
2.39 |
|
|
|
| 19 |
A' |
1115 |
1064 |
13.28 |
|
|
|
| 20 |
A' |
1007 |
961 |
1.82 |
|
|
|
| 21 |
A' |
977 |
932 |
0.62 |
|
|
|
| 22 |
A' |
917 |
876 |
1.90 |
|
|
|
| 23 |
A' |
746 |
711 |
1.40 |
|
|
|
| 24 |
A' |
523 |
500 |
1.48 |
|
|
|
| 25 |
A' |
441 |
421 |
0.85 |
|
|
|
| 26 |
A' |
331 |
316 |
0.27 |
|
|
|
| 27 |
A' |
278 |
265 |
0.50 |
|
|
|
| 28 |
A' |
260 |
249 |
0.07 |
|
|
|
| 29 |
A" |
3149 |
3005 |
17.94 |
|
|
|
| 30 |
A" |
3138 |
2994 |
0.02 |
|
|
|
| 31 |
A" |
3119 |
2976 |
22.26 |
|
|
|
| 32 |
A" |
3058 |
2918 |
23.91 |
|
|
|
| 33 |
A" |
1506 |
1437 |
0.13 |
|
|
|
| 34 |
A" |
1498 |
1429 |
0.85 |
|
|
|
| 35 |
A" |
1481 |
1414 |
9.30 |
|
|
|
| 36 |
A" |
1403 |
1339 |
6.11 |
|
|
|
| 37 |
A" |
1347 |
1286 |
1.65 |
|
|
|
| 38 |
A" |
1141 |
1089 |
1.61 |
|
|
|
| 39 |
A" |
1067 |
1019 |
0.88 |
|
|
|
| 40 |
A" |
973 |
928 |
0.20 |
|
|
|
| 41 |
A" |
933 |
890 |
13.73 |
|
|
|
| 42 |
A" |
924 |
882 |
28.41 |
|
|
|
| 43 |
A" |
747 |
713 |
0.57 |
|
|
|
| 44 |
A" |
555 |
530 |
7.07 |
|
|
|
| 45 |
A" |
260 |
248 |
0.01 |
|
|
|
| 46 |
A" |
214 |
205 |
0.42 |
|
|
|
| 47 |
A" |
163 |
155 |
0.20 |
|
|
|
| 48 |
A" |
40 |
38 |
0.14 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36403.3 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 34739.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.340 |
|
|
|
| 2 |
C |
0.167 |
|
|
|
| 3 |
C |
-0.167 |
|
|
|
| 4 |
C |
-0.436 |
|
|
|
| 5 |
C |
-0.379 |
|
|
|
| 6 |
C |
-0.379 |
|
|
|
| 7 |
H |
0.112 |
|
|
|
| 8 |
H |
0.113 |
|
|
|
| 9 |
H |
0.123 |
|
|
|
| 10 |
H |
0.135 |
|
|
|
| 11 |
H |
0.139 |
|
|
|
| 12 |
H |
0.139 |
|
|
|
| 13 |
H |
0.127 |
|
|
|
| 14 |
H |
0.135 |
|
|
|
| 15 |
H |
0.125 |
|
|
|
| 16 |
H |
0.127 |
|
|
|
| 17 |
H |
0.135 |
|
|
|
| 18 |
H |
0.125 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.295 |
0.390 |
0.000 |
0.489 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.450 |
-0.684 |
0.000 |
| y |
-0.684 |
-39.100 |
0.000 |
| z |
0.000 |
0.000 |
-40.909 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.554 |
-0.684 |
0.000 |
| y |
-0.684 |
0.579 |
0.000 |
| z |
0.000 |
0.000 |
-2.134 |
|
| Polar |
|---|
| 3z2-r2 | -4.267 |
|---|
| x2-y2 | 0.650 |
|---|
| xy | -0.684 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.738 |
0.793 |
0.000 |
| y |
0.793 |
11.059 |
0.000 |
| z |
0.000 |
0.000 |
7.712 |
<r2> (average value of r
2) Å
2
| <r2> |
189.947 |
| (<r2>)1/2 |
13.782 |