return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for HO2CCHNH2CH2OH (Serine)

using model chemistry: MP2=FULL/6-31G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2=FULL/6-31G**
 hartrees
Energy at 0K-397.888563
Energy at 298.15K-397.898706
HF Energy-396.736115
Nuclear repulsion energy326.047643
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3858 3605 62.22      
2 A 3807 3557 80.10      
3 A 3682 3441 4.47      
4 A 3576 3341 1.46      
5 A 3221 3010 19.57      
6 A 3151 2944 9.88      
7 A 3087 2885 47.93      
8 A 1822 1703 196.74      
9 A 1702 1591 34.35      
10 A 1564 1461 1.86      
11 A 1468 1371 34.49      
12 A 1442 1348 12.11      
13 A 1425 1332 62.36      
14 A 1414 1321 11.68      
15 A 1349 1261 19.41      
16 A 1273 1189 9.18      
17 A 1222 1142 3.01      
18 A 1190 1112 169.12      
19 A 1173 1096 73.31      
20 A 1123 1050 89.70      
21 A 1061 991 55.61      
22 A 1025 958 0.93      
23 A 922 862 192.71      
24 A 831 776 19.30      
25 A 750 700 31.33      
26 A 649 607 105.89      
27 A 590 551 6.33      
28 A 566 529 221.22      
29 A 526 491 19.57      
30 A 451 421 7.38      
31 A 310 290 13.41      
32 A 302 282 21.12      
33 A 275 257 20.31      
34 A 225 210 2.60      
35 A 176 165 3.04      
36 A 31 29 0.94      

Unscaled Zero Point Vibrational Energy (zpe) 25618.5 cm-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 23938.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31G**
ABC
0.11938 0.07874 0.05170

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31G**

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.766 -0.550 0.003
O2 2.051 -0.351 -0.364
O3 0.371 -1.567 0.548
C4 -0.081 0.675 -0.310
C5 -1.501 0.459 0.193
O6 -2.105 -0.674 -0.389
N7 0.458 1.913 0.230
H8 2.535 -1.157 -0.127
H9 -0.119 0.765 -1.398
H10 -1.481 0.402 1.287
H11 -2.088 1.330 -0.086
H12 -1.645 -1.437 -0.019
H13 1.363 2.112 -0.174
H14 0.584 1.833 1.232

Atom - Atom Distances (Å)
  C1 O2 O3 C4 C5 O6 N7 H8 H9 H10 H11 H12 H13 H14
C11.35061.22001.52222.48912.90042.49291.87462.11542.75833.41932.56952.73402.6871
O21.35062.26532.36713.68584.16852.83200.96902.65093.97124.47653.86752.56513.0770
O31.22002.26532.44312.78132.79353.49612.30393.07662.80283.85282.09823.87883.4745
C41.52222.36712.44311.52202.43361.45493.19931.09192.14182.12322.64412.04222.0400
C52.48913.68582.78131.52201.40942.44064.35962.12921.09621.08701.91313.32772.7050
O62.90044.16852.79352.43361.40943.69444.67252.65182.08732.02750.96424.45414.0178
N72.49292.83203.49611.45492.44063.69443.72402.07472.67612.63133.96371.01121.0126
H81.87460.96902.30393.19934.35964.67253.72403.51524.53455.25014.19113.47323.8196
H92.11542.65093.07661.09192.12922.65182.07473.51523.03262.43263.01272.34762.9243
H102.75833.97122.80282.14181.09622.08732.67614.53453.03261.76492.26213.62642.5130
H113.41934.47653.85282.12321.08702.02752.63135.25012.43261.76492.80343.54003.0217
H122.56953.86752.09822.64411.91310.96423.96374.19113.01272.26212.80344.65524.1504
H132.73402.56513.87882.04223.32774.45411.01123.47322.34763.62643.54004.65521.6317
H142.68713.07703.47452.04002.70504.01781.01263.81962.92432.51303.02174.15041.6317

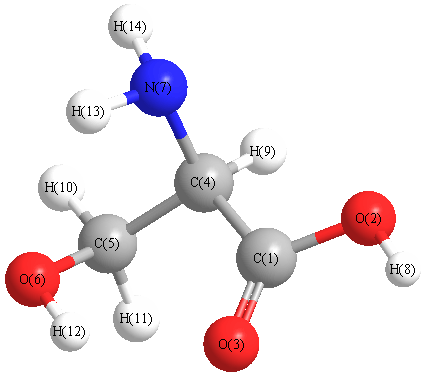
picture of Serine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 O2 H8 106.676 C1 C4 C5 109.704
C1 C4 N7 113.706 C1 C4 H9 106.892
O2 C1 O3 123.504 O2 C1 C4 110.834
O3 C1 C4 125.625 C4 C5 O6 112.178
C4 C5 H10 108.685 C4 C5 H11 107.768
C4 N7 H13 110.510 C4 N7 H14 110.244
C5 C4 N7 110.114 C5 C4 H9 107.954
C5 O6 H12 105.885 O6 C5 H10 112.224
O6 C5 H11 107.922 N7 C4 H9 108.260
H10 C5 H11 107.880 H13 N7 H14 107.463
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability