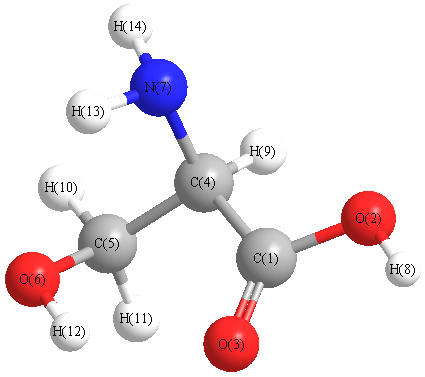
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3799 |
3625 |
42.35 |
|
|
|
| 2 |
A |
3785 |
3612 |
136.67 |
|
|
|
| 3 |
A |
3637 |
3471 |
4.08 |
|
|
|
| 4 |
A |
3545 |
3383 |
1.04 |
|
|
|
| 5 |
A |
3139 |
2996 |
22.25 |
|
|
|
| 6 |
A |
3066 |
2926 |
12.31 |
|
|
|
| 7 |
A |
2984 |
2847 |
67.14 |
|
|
|
| 8 |
A |
1840 |
1756 |
243.85 |
|
|
|
| 9 |
A |
1678 |
1601 |
37.47 |
|
|
|
| 10 |
A |
1521 |
1452 |
1.70 |
|
|
|
| 11 |
A |
1464 |
1397 |
47.07 |
|
|
|
| 12 |
A |
1405 |
1341 |
66.33 |
|
|
|
| 13 |
A |
1402 |
1338 |
44.55 |
|
|
|
| 14 |
A |
1389 |
1326 |
10.05 |
|
|
|
| 15 |
A |
1329 |
1268 |
11.30 |
|
|
|
| 16 |
A |
1247 |
1190 |
7.42 |
|
|
|
| 17 |
A |
1218 |
1163 |
34.02 |
|
|
|
| 18 |
A |
1202 |
1147 |
164.60 |
|
|
|
| 19 |
A |
1168 |
1115 |
46.66 |
|
|
|
| 20 |
A |
1134 |
1082 |
106.41 |
|
|
|
| 21 |
A |
1041 |
993 |
39.15 |
|
|
|
| 22 |
A |
1005 |
959 |
0.82 |
|
|
|
| 23 |
A |
886 |
845 |
186.51 |
|
|
|
| 24 |
A |
820 |
782 |
40.08 |
|
|
|
| 25 |
A |
751 |
716 |
41.54 |
|
|
|
| 26 |
A |
659 |
629 |
89.66 |
|
|
|
| 27 |
A |
597 |
570 |
170.95 |
|
|
|
| 28 |
A |
572 |
546 |
32.49 |
|
|
|
| 29 |
A |
541 |
516 |
35.97 |
|
|
|
| 30 |
A |
426 |
407 |
8.06 |
|
|
|
| 31 |
A |
317 |
303 |
5.07 |
|
|
|
| 32 |
A |
303 |
289 |
8.76 |
|
|
|
| 33 |
A |
252 |
241 |
23.23 |
|
|
|
| 34 |
A |
233 |
222 |
19.55 |
|
|
|
| 35 |
A |
168 |
161 |
1.63 |
|
|
|
| 36 |
A |
32 |
31 |
0.36 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25277.3 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 24122.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.599 |
|
|
|
| 2 |
O |
-0.495 |
|
|
|
| 3 |
O |
-0.490 |
|
|
|
| 4 |
C |
-0.120 |
|
|
|
| 5 |
C |
0.029 |
|
|
|
| 6 |
O |
-0.558 |
|
|
|
| 7 |
N |
-0.613 |
|
|
|
| 8 |
H |
0.341 |
|
|
|
| 9 |
H |
0.185 |
|
|
|
| 10 |
H |
0.099 |
|
|
|
| 11 |
H |
0.151 |
|
|
|
| 12 |
H |
0.336 |
|
|
|
| 13 |
H |
0.274 |
|
|
|
| 14 |
H |
0.261 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
3.621 |
0.326 |
0.715 |
3.705 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.237 |
-2.360 |
-0.732 |
| y |
-2.360 |
-44.114 |
0.977 |
| z |
-0.732 |
0.977 |
-38.760 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.200 |
-2.360 |
-0.732 |
| y |
-2.360 |
-4.115 |
0.977 |
| z |
-0.732 |
0.977 |
3.915 |
|
| Polar |
|---|
| 3z2-r2 | 7.831 |
|---|
| x2-y2 | 2.877 |
|---|
| xy | -2.360 |
|---|
| xz | -0.732 |
|---|
| yz | 0.977 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.795 |
0.208 |
0.016 |
| y |
0.208 |
7.481 |
-0.016 |
| z |
0.016 |
-0.016 |
5.472 |
<r2> (average value of r
2) Å
2
| <r2> |
214.641 |
| (<r2>)1/2 |
14.651 |