return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H6 (Dewar Benzene)

using model chemistry: MP2=FULL/6-31G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2=FULL/6-31G**
 hartrees
Energy at 0K-231.408011
Energy at 298.15K 
HF Energy-230.574506
Nuclear repulsion energy209.257539
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3304 3087 7.53 252.47 0.03 0.06
2 A1 3201 2991 27.39 128.88 0.15 0.25
3 A1 1627 1521 0.10 11.65 0.03 0.05
4 A1 1207 1128 3.45 16.87 0.21 0.34
5 A1 1069 998 2.18 14.06 0.75 0.86
6 A1 987 922 4.85 10.86 0.24 0.39
7 A1 898 839 0.73 6.65 0.74 0.85
8 A1 831 776 65.93 5.54 0.16 0.27
9 A1 398 372 3.20 11.96 0.65 0.79
10 A2 3271 3057 0.00 145.44 0.75 0.86
11 A2 1337 1249 0.00 4.75 0.75 0.86
12 A2 1239 1158 0.00 11.40 0.75 0.86
13 A2 999 934 0.00 0.15 0.75 0.86
14 A2 914 854 0.00 0.00 0.75 0.86
15 A2 783 732 0.00 3.01 0.75 0.86
16 A2 324 303 0.00 2.53 0.75 0.86
17 B1 3302 3085 40.64 54.53 0.75 0.86
18 B1 1602 1497 6.69 1.78 0.75 0.86
19 B1 1256 1174 2.59 3.09 0.75 0.86
20 B1 1139 1065 0.00 0.03 0.75 0.86
21 B1 1048 979 0.07 9.20 0.75 0.86
22 B1 736 688 41.57 5.01 0.75 0.86
23 B2 3273 3058 22.33 41.42 0.75 0.86
24 B2 3194 2985 12.81 101.59 0.75 0.86
25 B2 1340 1252 30.76 0.01 0.75 0.86
26 B2 1192 1113 5.66 0.98 0.75 0.86
27 B2 950 887 8.29 0.01 0.75 0.86
28 B2 923 863 5.88 0.74 0.75 0.86
29 B2 848 793 4.13 1.78 0.75 0.86
30 B2 487 455 5.50 6.39 0.75 0.86

Unscaled Zero Point Vibrational Energy (zpe) 21838.6 cm-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 20406.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31G**
ABC
0.26104 0.15085 0.11724

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31G**

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.785 0.524
C2 0.000 -0.785 0.524
H3 0.000 1.342 1.458
H4 0.000 -1.342 1.458
C5 -1.295 0.673 -0.264
C6 1.295 0.673 -0.264
C7 1.295 -0.673 -0.264
C8 -1.295 -0.673 -0.264
H9 -1.931 1.420 -0.717
H10 1.931 1.420 -0.717
H11 1.931 -1.420 -0.717
H12 -1.931 -1.420 -0.717

Atom - Atom Distances (Å)
  C1 C2 H3 H4 C5 C6 C7 C8 H9 H10 H11 H12
C11.56951.08772.32251.51921.51922.10292.10292.38162.38163.18263.1826
C21.56952.32251.08772.10292.10291.51921.51923.18263.18262.38162.3816
H31.08772.32252.68312.25532.25532.94962.94962.90982.90984.01074.0107
H42.32251.08772.68312.94962.94962.25532.25534.01074.01072.90982.9098
C51.51922.10292.25532.94962.58922.91851.34691.08073.34193.87212.2345
C61.51922.10292.25532.94962.58921.34692.91853.34191.08072.23453.8721
C72.10291.51922.94962.25532.91851.34692.58923.87212.23451.08073.3419
C82.10291.51922.94962.25531.34692.91852.58922.23453.87213.34191.0807
H92.38163.18262.90984.01071.08073.34193.87212.23453.86234.79392.8397
H102.38163.18262.90984.01073.34191.08072.23453.87213.86232.83974.7939
H113.18262.38164.01072.90983.87212.23451.08073.34194.79392.83973.8623
H123.18262.38164.01072.90982.23453.87213.34191.08072.83974.79393.8623

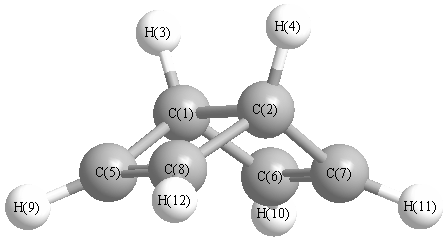
picture of Dewar Benzene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 120.789 C1 C2 H7 85.799
C1 C2 H8 85.799 C1 C5 H8 94.201
C1 C5 H9 131.970 C1 C6 H7 94.201
C1 C6 H10 131.970 C2 C1 C3 120.789
C2 C1 C5 85.799 C2 C1 C6 85.799
C2 H7 C6 94.201 C2 H7 H11 131.970
C2 H8 C5 94.201 C2 H8 H12 131.970
C3 C1 C5 118.859 C3 C1 C6 118.859
C4 C2 H7 118.859 C4 C2 H8 118.859
C5 C1 C6 116.886 C5 H8 H12 133.683
C6 H7 H11 133.683 H7 C6 H10 133.683
H8 C5 H9 133.683
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability