return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H6O (2,5-Cyclohexadienone)

using model chemistry: QCISD/6-31G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at QCISD/6-31G**
 hartrees
Energy at 0K-306.561229
Energy at 298.15K-306.567747
HF Energy-305.549235
Nuclear repulsion energy268.084377
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3270 3079 0.29      
2 A1 3232 3043 7.74      
3 A1 3077 2897 15.56      
4 A1 1798 1693 211.77      
5 A1 1732 1630 20.18      
6 A1 1510 1422 7.17      
7 A1 1450 1365 1.05      
8 A1 1219 1148 4.69      
9 A1 977 920 3.28      
10 A1 903 850 4.87      
11 A1 782 737 2.43      
12 A1 505 475 1.82      
13 A2 1239 1166 0.00      
14 A2 995 937 0.00      
15 A2 753 709 0.00      
16 A2 347 327 0.00      
17 B1 3105 2923 12.67      
18 B1 1021 961 8.39      
19 B1 971 914 8.68      
20 B1 815 768 51.52      
21 B1 572 539 11.09      
22 B1 270 254 2.15      
23 B1 113 106 1.34      
24 B2 3269 3077 18.12      
25 B2 3231 3042 17.41      
26 B2 1714 1613 1.59      
27 B2 1463 1378 33.98      
28 B2 1428 1345 3.19      
29 B2 1310 1233 28.04      
30 B2 1170 1101 7.76      
31 B2 1017 957 4.52      
32 B2 581 547 0.31      
33 B2 453 427 15.28      

Unscaled Zero Point Vibrational Energy (zpe) 23145.1 cm-1
Scaled (by 0.9414) Zero Point Vibrational Energy (zpe) 21788.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/6-31G**
ABC
0.17371 0.08857 0.05929

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/6-31G**

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 1.114
C2 0.000 0.000 -1.830
C3 0.000 1.262 0.332
C4 0.000 -1.262 0.332
C5 0.000 1.259 -1.011
C6 0.000 -1.259 -1.011
O7 0.000 0.000 2.343
H8 0.000 2.179 0.906
H9 0.000 -2.179 0.906
H10 0.000 2.197 -1.556
H11 0.000 -2.197 -1.556
H12 0.869 0.000 -2.499
H13 -0.869 0.000 -2.499

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 O7 H8 H9 H10 H11 H12 H13
C12.94371.48391.48392.46962.46961.22992.18912.18913.45723.45723.71543.7154
C22.94372.50352.50351.50171.50174.17373.49813.49812.21352.21351.09671.0967
C31.48392.50352.52311.34342.85602.37421.08243.48842.10723.94013.21893.2189
C41.48392.50352.52312.85601.34342.37423.48841.08243.94012.10723.21893.2189
C52.46961.50171.34342.85602.51753.58312.12683.93651.08463.49802.13382.1338
C62.46961.50172.85601.34342.51753.58313.93652.12683.49801.08462.13382.1338
O71.22994.17372.37422.37423.58313.58312.61062.61064.47574.47574.91974.9197
H82.18913.49811.08243.48842.12683.93652.61064.35862.46235.02104.13494.1349
H92.18913.49813.48841.08243.93652.12682.61064.35865.02102.46234.13494.1349
H103.45722.21352.10723.94011.08463.49804.47572.46235.02104.39302.54342.5434
H113.45722.21353.94012.10723.49801.08464.47575.02102.46234.39302.54342.5434
H123.71541.09673.21893.21892.13382.13384.91974.13494.13492.54342.54341.7387
H133.71541.09673.21893.21892.13382.13384.91974.13494.13492.54342.54341.7387

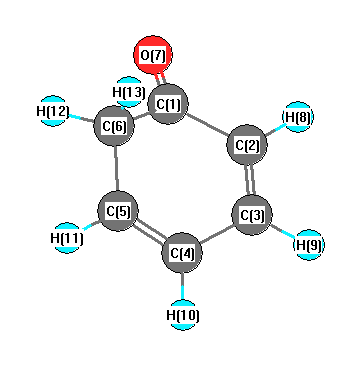
picture of 2,5-Cyclohexadienone state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C3 C5 121.653 C1 C3 H8 116.208
C1 C4 C6 121.653 C1 C4 H9 116.208
C2 C5 C3 123.169 C2 C5 H10 116.787
C2 C6 C4 123.169 C2 C6 H11 116.787
C3 C1 C4 116.454 C3 C1 O7 121.773
C3 C5 H10 120.044 C4 C1 O7 121.773
C4 C6 H11 120.044 C5 C2 C6 113.901
C5 C2 H12 109.418 C5 C2 H13 109.418
C5 C3 H8 122.139 C6 C2 H12 109.418
C6 C2 H13 109.418 C6 C4 H9 122.139
H12 C2 H13 104.879
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability