Jump to
S1C2
S1C3
Energy calculated at CCD/6-31G**
| | hartrees |
|---|
| Energy at 0K | -224.661622 |
| Energy at 298.15K | -224.667926 |
| HF Energy | -223.998406 |
| Nuclear repulsion energy | 123.650833 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3791 |
3554 |
32.81 |
|
|
|
| 2 |
A |
3672 |
3443 |
3.54 |
|
|
|
| 3 |
A |
1906 |
1787 |
387.93 |
|
|
|
| 4 |
A |
1687 |
1582 |
0.04 |
|
|
|
| 5 |
A |
1226 |
1150 |
3.49 |
|
|
|
| 6 |
A |
981 |
920 |
7.01 |
|
|
|
| 7 |
A |
663 |
621 |
129.28 |
|
|
|
| 8 |
A |
481 |
451 |
2.43 |
|
|
|
| 9 |
A |
409 |
384 |
81.88 |
|
|
|
| 10 |
B |
3791 |
3554 |
23.59 |
|
|
|
| 11 |
B |
3669 |
3440 |
46.67 |
|
|
|
| 12 |
B |
1697 |
1591 |
197.35 |
|
|
|
| 13 |
B |
1470 |
1378 |
192.78 |
|
|
|
| 14 |
B |
1104 |
1035 |
20.26 |
|
|
|
| 15 |
B |
810 |
759 |
164.97 |
|
|
|
| 16 |
B |
603 |
565 |
253.25 |
|
|
|
| 17 |
B |
571 |
535 |
73.28 |
|
|
|
| 18 |
B |
455 |
427 |
42.71 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 14492.0 cm
-1
Scaled (by 0.9376) Zero Point Vibrational Energy (zpe) 13587.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCD/6-31G**
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.146 |
| O2 |
0.000 |
0.000 |
1.363 |
| N3 |
0.000 |
1.159 |
-0.619 |
| N4 |
0.000 |
-1.159 |
-0.619 |
| H5 |
0.223 |
1.973 |
-0.070 |
| H6 |
0.505 |
1.111 |
-1.489 |
| H7 |
-0.223 |
-1.973 |
-0.070 |
| H8 |
-0.505 |
-1.111 |
-1.489 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
N4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.2170 | 1.3891 | 1.3891 | 1.9971 | 2.0405 | 1.9971 | 2.0405 |
O2 | 1.2170 | | 2.2965 | 2.2965 | 2.4487 | 3.1024 | 2.4487 | 3.1024 | N3 | 1.3891 | 2.2965 | | 2.3181 | 1.0068 | 1.0071 | 3.1875 | 2.4829 | N4 | 1.3891 | 2.2965 | 2.3181 | | 3.1875 | 2.4829 | 1.0068 | 1.0071 | H5 | 1.9971 | 2.4487 | 1.0068 | 3.1875 | | 1.6840 | 3.9708 | 3.4718 | H6 | 2.0405 | 3.1024 | 1.0071 | 2.4829 | 1.6840 | | 3.4718 | 2.4409 | H7 | 1.9971 | 2.4487 | 3.1875 | 1.0068 | 3.9708 | 3.4718 | | 1.6840 | H8 | 2.0405 | 3.1024 | 2.4829 | 1.0071 | 3.4718 | 2.4409 | 1.6840 | |
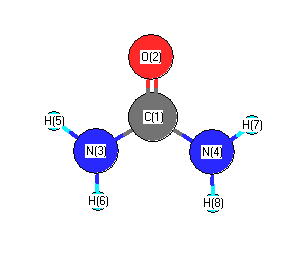 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
111.945 |
|
C1 |
N3 |
H6 |
115.852 |
| C1 |
N4 |
H7 |
111.945 |
|
C1 |
N4 |
H8 |
115.852 |
| O2 |
C1 |
N3 |
123.444 |
|
O2 |
C1 |
N4 |
123.444 |
| N3 |
C1 |
N4 |
113.112 |
|
H5 |
N3 |
H6 |
113.486 |
| H7 |
N4 |
H8 |
113.486 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Energy calculated at CCD/6-31G**
| | hartrees |
|---|
| Energy at 0K | -224.659086 |
| Energy at 298.15K | -224.664982 |
| HF Energy | -223.996579 |
| Nuclear repulsion energy | 123.706512 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3823 |
3584 |
36.01 |
|
|
|
| 2 |
A' |
3699 |
3468 |
11.88 |
|
|
|
| 3 |
A' |
1893 |
1775 |
419.82 |
|
|
|
| 4 |
A' |
1702 |
1595 |
19.04 |
|
|
|
| 5 |
A' |
1222 |
1146 |
4.65 |
|
|
|
| 6 |
A' |
990 |
928 |
11.64 |
|
|
|
| 7 |
A' |
783 |
734 |
65.93 |
|
|
|
| 8 |
A' |
558 |
523 |
101.26 |
|
|
|
| 9 |
A' |
505 |
473 |
113.95 |
|
|
|
| 10 |
A' |
450 |
422 |
248.31 |
|
|
|
| 11 |
A" |
3820 |
3582 |
27.74 |
|
|
|
| 12 |
A" |
3691 |
3461 |
45.75 |
|
|
|
| 13 |
A" |
1690 |
1584 |
209.19 |
|
|
|
| 14 |
A" |
1475 |
1383 |
197.45 |
|
|
|
| 15 |
A" |
1065 |
998 |
17.83 |
|
|
|
| 16 |
A" |
584 |
548 |
35.90 |
|
|
|
| 17 |
A" |
464 |
435 |
112.83 |
|
|
|
| 18 |
A" |
195 |
183 |
32.38 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 14303.3 cm
-1
Scaled (by 0.9376) Zero Point Vibrational Energy (zpe) 13410.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCD/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.008 |
0.141 |
0.000 |
| O2 |
0.042 |
1.360 |
0.000 |
| N3 |
0.042 |
-0.607 |
1.163 |
| N4 |
0.042 |
-0.607 |
-1.163 |
| H5 |
-0.173 |
-0.072 |
1.986 |
| H6 |
-0.318 |
-1.545 |
1.157 |
| H7 |
-0.173 |
-0.072 |
-1.986 |
| H8 |
-0.318 |
-1.545 |
-1.157 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
N4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.2189 | 1.3834 | 1.3834 | 2.0057 | 2.0705 | 2.0057 | 2.0705 |
O2 | 1.2189 | | 2.2849 | 2.2849 | 2.4577 | 3.1470 | 2.4577 | 3.1470 | N3 | 1.3834 | 2.2849 | | 2.3262 | 1.0049 | 1.0045 | 3.2016 | 2.5279 | N4 | 1.3834 | 2.2849 | 2.3262 | | 3.2016 | 2.5279 | 1.0049 | 1.0045 | H5 | 2.0057 | 2.4577 | 1.0049 | 3.2016 | | 1.6966 | 3.9722 | 3.4738 | H6 | 2.0705 | 3.1470 | 1.0045 | 2.5279 | 1.6966 | | 3.4738 | 2.3133 | H7 | 2.0057 | 2.4577 | 3.2016 | 1.0049 | 3.9722 | 3.4738 | | 1.6966 | H8 | 2.0705 | 3.1470 | 2.5279 | 1.0045 | 3.4738 | 2.3133 | 1.6966 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
113.284 |
|
C1 |
N3 |
H6 |
119.386 |
| C1 |
N4 |
H7 |
113.284 |
|
C1 |
N4 |
H8 |
119.386 |
| O2 |
C1 |
N3 |
122.683 |
|
O2 |
C1 |
N4 |
122.683 |
| N3 |
C1 |
N4 |
114.436 |
|
H5 |
N3 |
H6 |
115.192 |
| H7 |
N4 |
H8 |
115.192 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at CCD/6-31G**
| | hartrees |
|---|
| Energy at 0K | -224.657739 |
| Energy at 298.15K | |
| HF Energy | -223.997096 |
| Nuclear repulsion energy | 123.941476 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3883 |
3641 |
61.57 |
|
|
|
| 2 |
A1 |
3746 |
3513 |
11.64 |
|
|
|
| 3 |
A1 |
1883 |
1766 |
486.65 |
|
|
|
| 4 |
A1 |
1686 |
1581 |
3.61 |
|
|
|
| 5 |
A1 |
1183 |
1109 |
0.27 |
|
|
|
| 6 |
A1 |
1005 |
943 |
10.60 |
|
|
|
| 7 |
A1 |
488 |
458 |
2.34 |
|
|
|
| 8 |
A2 |
382 |
358 |
0.00 |
|
|
|
| 9 |
A2 |
472i |
443i |
0.00 |
|
|
|
| 10 |
B1 |
774 |
726 |
14.43 |
|
|
|
| 11 |
B1 |
577 |
541 |
13.15 |
|
|
|
| 12 |
B1 |
367i |
344i |
589.94 |
|
|
|
| 13 |
B2 |
3881 |
3639 |
38.61 |
|
|
|
| 14 |
B2 |
3739 |
3505 |
81.38 |
|
|
|
| 15 |
B2 |
1686 |
1581 |
305.31 |
|
|
|
| 16 |
B2 |
1483 |
1391 |
214.36 |
|
|
|
| 17 |
B2 |
1027 |
962 |
9.15 |
|
|
|
| 18 |
B2 |
573 |
537 |
16.34 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13579.1 cm
-1
Scaled (by 0.9376) Zero Point Vibrational Energy (zpe) 12731.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCD/6-31G**
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.143 |
| O2 |
0.000 |
0.000 |
1.364 |
| N3 |
0.000 |
1.155 |
-0.599 |
| N4 |
0.000 |
-1.155 |
-0.599 |
| H5 |
0.000 |
2.019 |
-0.093 |
| H6 |
0.000 |
1.174 |
-1.599 |
| H7 |
0.000 |
-2.019 |
-0.093 |
| H8 |
0.000 |
-1.174 |
-1.599 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
N4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.2211 | 1.3727 | 1.3727 | 2.0324 | 2.1011 | 2.0324 | 2.1011 |
O2 | 1.2211 | | 2.2776 | 2.2776 | 2.4898 | 3.1876 | 2.4898 | 3.1876 | N3 | 1.3727 | 2.2776 | | 2.3095 | 1.0010 | 1.0004 | 3.2134 | 2.5347 | N4 | 1.3727 | 2.2776 | 2.3095 | | 3.2134 | 2.5347 | 1.0010 | 1.0004 | H5 | 2.0324 | 2.4898 | 1.0010 | 3.2134 | | 1.7265 | 4.0372 | 3.5302 | H6 | 2.1011 | 3.1876 | 1.0004 | 2.5347 | 1.7265 | | 3.5302 | 2.3486 | H7 | 2.0324 | 2.4898 | 3.2134 | 1.0010 | 4.0372 | 3.5302 | | 1.7265 | H8 | 2.1011 | 3.1876 | 2.5347 | 1.0004 | 3.5302 | 2.3486 | 1.7265 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
116.926 |
|
C1 |
N3 |
H6 |
123.846 |
| C1 |
N4 |
H7 |
116.926 |
|
C1 |
N4 |
H8 |
123.846 |
| O2 |
C1 |
N3 |
122.728 |
|
O2 |
C1 |
N4 |
122.728 |
| N3 |
C1 |
N4 |
114.545 |
|
H5 |
N3 |
H6 |
119.228 |
| H7 |
N4 |
H8 |
119.228 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
