Jump to
S1C2
S1C3
Energy calculated at TPSSh/6-31G**
| | hartrees |
|---|
| Energy at 0K | -225.272891 |
| Energy at 298.15K | -225.279053 |
| HF Energy | -225.272891 |
| Nuclear repulsion energy | 123.201368 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at TPSSh/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3683 |
3530 |
21.61 |
|
|
|
| 2 |
A |
3560 |
3413 |
0.73 |
|
|
|
| 3 |
A |
1825 |
1750 |
345.57 |
|
|
|
| 4 |
A |
1639 |
1572 |
0.08 |
|
|
|
| 5 |
A |
1181 |
1132 |
3.07 |
|
|
|
| 6 |
A |
950 |
911 |
5.70 |
|
|
|
| 7 |
A |
608 |
583 |
91.64 |
|
|
|
| 8 |
A |
458 |
439 |
3.50 |
|
|
|
| 9 |
A |
390 |
374 |
66.84 |
|
|
|
| 10 |
B |
3682 |
3530 |
25.43 |
|
|
|
| 11 |
B |
3557 |
3410 |
29.73 |
|
|
|
| 12 |
B |
1644 |
1576 |
178.93 |
|
|
|
| 13 |
B |
1418 |
1360 |
199.00 |
|
|
|
| 14 |
B |
1047 |
1004 |
23.21 |
|
|
|
| 15 |
B |
774 |
742 |
108.10 |
|
|
|
| 16 |
B |
576 |
552 |
207.41 |
|
|
|
| 17 |
B |
541 |
519 |
107.14 |
|
|
|
| 18 |
B |
452 |
433 |
90.50 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13992.3 cm
-1
Scaled (by 0.9587) Zero Point Vibrational Energy (zpe) 13414.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at TPSSh/6-31G**
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.144 |
| O2 |
0.000 |
0.000 |
1.368 |
| N3 |
0.000 |
1.164 |
-0.618 |
| N4 |
0.000 |
-1.164 |
-0.618 |
| H5 |
0.229 |
1.980 |
-0.066 |
| H6 |
0.472 |
1.131 |
-1.513 |
| H7 |
-0.229 |
-1.980 |
-0.066 |
| H8 |
-0.472 |
-1.131 |
-1.513 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
N4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.2240 | 1.3907 | 1.3907 | 2.0042 | 2.0600 | 2.0042 | 2.0600 |
O2 | 1.2240 | | 2.3012 | 2.3012 | 2.4551 | 3.1299 | 2.4551 | 3.1299 | N3 | 1.3907 | 2.3012 | | 2.3279 | 1.0113 | 1.0124 | 3.2003 | 2.5077 | N4 | 1.3907 | 2.3012 | 2.3279 | | 3.2003 | 2.5077 | 1.0113 | 1.0124 | H5 | 2.0042 | 2.4551 | 1.0113 | 3.2003 | | 1.6954 | 3.9865 | 3.5014 | H6 | 2.0600 | 3.1299 | 1.0124 | 2.5077 | 1.6954 | | 3.5014 | 2.4501 | H7 | 2.0042 | 2.4551 | 3.2003 | 1.0113 | 3.9865 | 3.5014 | | 1.6954 | H8 | 2.0600 | 3.1299 | 2.5077 | 1.0124 | 3.5014 | 2.4501 | 1.6954 | |
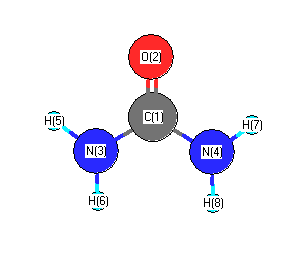 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
112.138 |
|
C1 |
N3 |
H6 |
117.147 |
| C1 |
N4 |
H7 |
112.138 |
|
C1 |
N4 |
H8 |
117.147 |
| O2 |
C1 |
N3 |
123.182 |
|
O2 |
C1 |
N4 |
123.182 |
| N3 |
C1 |
N4 |
113.635 |
|
H5 |
N3 |
H6 |
113.808 |
| H7 |
N4 |
H8 |
113.808 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at TPSSh/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.672 |
|
|
|
| 2 |
O |
-0.511 |
|
|
|
| 3 |
N |
-0.650 |
|
|
|
| 4 |
N |
-0.650 |
|
|
|
| 5 |
H |
0.293 |
|
|
|
| 6 |
H |
0.277 |
|
|
|
| 7 |
H |
0.293 |
|
|
|
| 8 |
H |
0.277 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-3.508 |
3.508 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.135 |
0.210 |
0.000 |
| y |
0.210 |
4.772 |
0.000 |
| z |
0.000 |
0.000 |
4.910 |
<r2> (average value of r
2) Å
2
| <r2> |
68.722 |
| (<r2>)1/2 |
8.290 |
Jump to
S1C1
S1C3
Energy calculated at TPSSh/6-31G**
| | hartrees |
|---|
| Energy at 0K | -225.270897 |
| Energy at 298.15K | -225.276611 |
| HF Energy | -225.270897 |
| Nuclear repulsion energy | 123.246597 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at TPSSh/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3713 |
3560 |
22.68 |
|
|
|
| 2 |
A' |
3585 |
3437 |
7.01 |
|
|
|
| 3 |
A' |
1815 |
1740 |
364.95 |
|
|
|
| 4 |
A' |
1649 |
1581 |
14.34 |
|
|
|
| 5 |
A' |
1176 |
1127 |
3.58 |
|
|
|
| 6 |
A' |
958 |
919 |
7.60 |
|
|
|
| 7 |
A' |
756 |
725 |
36.45 |
|
|
|
| 8 |
A' |
549 |
526 |
53.68 |
|
|
|
| 9 |
A' |
471 |
451 |
60.96 |
|
|
|
| 10 |
A' |
418 |
401 |
327.81 |
|
|
|
| 11 |
A" |
3711 |
3558 |
33.40 |
|
|
|
| 12 |
A" |
3576 |
3428 |
26.77 |
|
|
|
| 13 |
A" |
1636 |
1569 |
196.45 |
|
|
|
| 14 |
A" |
1423 |
1364 |
202.07 |
|
|
|
| 15 |
A" |
1013 |
972 |
21.03 |
|
|
|
| 16 |
A" |
558 |
535 |
28.98 |
|
|
|
| 17 |
A" |
423 |
406 |
80.46 |
|
|
|
| 18 |
A" |
172 |
165 |
43.19 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13800.9 cm
-1
Scaled (by 0.9587) Zero Point Vibrational Energy (zpe) 13230.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at TPSSh/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.006 |
0.139 |
0.000 |
| O2 |
0.040 |
1.364 |
0.000 |
| N3 |
0.040 |
-0.607 |
1.167 |
| N4 |
0.040 |
-0.607 |
-1.167 |
| H5 |
-0.162 |
-0.067 |
1.996 |
| H6 |
-0.293 |
-1.560 |
1.176 |
| H7 |
-0.162 |
-0.067 |
-1.996 |
| H8 |
-0.293 |
-1.560 |
-1.176 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
N4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.2258 | 1.3855 | 1.3855 | 2.0136 | 2.0884 | 2.0136 | 2.0884 |
O2 | 1.2258 | | 2.2909 | 2.2909 | 2.4646 | 3.1700 | 2.4646 | 3.1700 | N3 | 1.3855 | 2.2909 | | 2.3341 | 1.0094 | 1.0100 | 3.2150 | 2.5516 | N4 | 1.3855 | 2.2909 | 2.3341 | | 3.2150 | 2.5516 | 1.0094 | 1.0100 | H5 | 2.0136 | 2.4646 | 1.0094 | 3.2150 | | 1.7084 | 3.9918 | 3.5083 | H6 | 2.0884 | 3.1700 | 1.0100 | 2.5516 | 1.7084 | | 3.5083 | 2.3523 | H7 | 2.0136 | 2.4646 | 3.2150 | 1.0094 | 3.9918 | 3.5083 | | 1.7084 | H8 | 2.0884 | 3.1700 | 2.5516 | 1.0100 | 3.5083 | 2.3523 | 1.7084 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
113.518 |
|
C1 |
N3 |
H6 |
120.528 |
| C1 |
N4 |
H7 |
113.518 |
|
C1 |
N4 |
H8 |
120.528 |
| O2 |
C1 |
N3 |
122.520 |
|
O2 |
C1 |
N4 |
122.520 |
| N3 |
C1 |
N4 |
114.771 |
|
H5 |
N3 |
H6 |
115.559 |
| H7 |
N4 |
H8 |
115.559 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at TPSSh/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.691 |
|
|
|
| 2 |
O |
-0.522 |
|
|
|
| 3 |
N |
-0.647 |
|
|
|
| 4 |
N |
-0.647 |
|
|
|
| 5 |
H |
0.295 |
|
|
|
| 6 |
H |
0.268 |
|
|
|
| 7 |
H |
0.295 |
|
|
|
| 8 |
H |
0.268 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.809 |
-3.837 |
0.000 |
4.242 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.013 |
0.184 |
0.000 |
| y |
0.184 |
4.942 |
0.000 |
| z |
0.000 |
0.000 |
4.814 |
<r2> (average value of r
2) Å
2
| <r2> |
68.772 |
| (<r2>)1/2 |
8.293 |
Jump to
S1C1
S1C2
Energy calculated at TPSSh/6-31G**
| | hartrees |
|---|
| Energy at 0K | -225.269992 |
| Energy at 298.15K | |
| HF Energy | -225.269992 |
| Nuclear repulsion energy | 123.412981 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at TPSSh/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3763 |
3608 |
35.97 |
|
|
|
| 2 |
A1 |
3623 |
3474 |
3.51 |
|
|
|
| 3 |
A1 |
1812 |
1737 |
404.46 |
|
|
|
| 4 |
A1 |
1635 |
1567 |
0.92 |
|
|
|
| 5 |
A1 |
1142 |
1095 |
0.17 |
|
|
|
| 6 |
A1 |
969 |
929 |
6.49 |
|
|
|
| 7 |
A1 |
465 |
446 |
3.36 |
|
|
|
| 8 |
A2 |
370 |
355 |
0.00 |
|
|
|
| 9 |
A2 |
430i |
412i |
0.00 |
|
|
|
| 10 |
B1 |
757 |
726 |
9.55 |
|
|
|
| 11 |
B1 |
571 |
548 |
11.72 |
|
|
|
| 12 |
B1 |
323i |
309i |
533.17 |
|
|
|
| 13 |
B2 |
3761 |
3606 |
45.42 |
|
|
|
| 14 |
B2 |
3614 |
3465 |
46.22 |
|
|
|
| 15 |
B2 |
1631 |
1564 |
271.91 |
|
|
|
| 16 |
B2 |
1428 |
1369 |
213.79 |
|
|
|
| 17 |
B2 |
979 |
939 |
12.33 |
|
|
|
| 18 |
B2 |
548 |
525 |
14.81 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13157.5 cm
-1
Scaled (by 0.9587) Zero Point Vibrational Energy (zpe) 12614.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at TPSSh/6-31G**
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.141 |
| O2 |
0.000 |
0.000 |
1.368 |
| N3 |
0.000 |
1.161 |
-0.600 |
| N4 |
0.000 |
-1.161 |
-0.600 |
| H5 |
0.000 |
2.025 |
-0.086 |
| H6 |
0.000 |
1.193 |
-1.607 |
| H7 |
0.000 |
-2.025 |
-0.086 |
| H8 |
0.000 |
-1.193 |
-1.607 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
N4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.2269 | 1.3773 | 1.3773 | 2.0380 | 2.1157 | 2.0380 | 2.1157 |
O2 | 1.2269 | | 2.2851 | 2.2851 | 2.4931 | 3.2047 | 2.4931 | 3.2047 | N3 | 1.3773 | 2.2851 | | 2.3213 | 1.0061 | 1.0066 | 3.2272 | 2.5593 | N4 | 1.3773 | 2.2851 | 2.3213 | | 3.2272 | 2.5593 | 1.0061 | 1.0066 | H5 | 2.0380 | 2.4931 | 1.0061 | 3.2272 | | 1.7337 | 4.0506 | 3.5590 | H6 | 2.1157 | 3.2047 | 1.0066 | 2.5593 | 1.7337 | | 3.5590 | 2.3851 | H7 | 2.0380 | 2.4931 | 3.2272 | 1.0061 | 4.0506 | 3.5590 | | 1.7337 | H8 | 2.1157 | 3.2047 | 2.5593 | 1.0066 | 3.5590 | 2.3851 | 1.7337 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
116.674 |
|
C1 |
N3 |
H6 |
124.388 |
| C1 |
N4 |
H7 |
116.674 |
|
C1 |
N4 |
H8 |
124.388 |
| O2 |
C1 |
N3 |
122.572 |
|
O2 |
C1 |
N4 |
122.572 |
| N3 |
C1 |
N4 |
114.857 |
|
H5 |
N3 |
H6 |
118.938 |
| H7 |
N4 |
H8 |
118.938 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at TPSSh/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.735 |
|
|
|
| 2 |
O |
-0.538 |
|
|
|
| 3 |
N |
-0.671 |
|
|
|
| 4 |
N |
-0.671 |
|
|
|
| 5 |
H |
0.300 |
|
|
|
| 6 |
H |
0.273 |
|
|
|
| 7 |
H |
0.300 |
|
|
|
| 8 |
H |
0.273 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-4.245 |
4.245 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
1.868 |
0.000 |
0.000 |
| y |
0.000 |
4.786 |
0.000 |
| z |
0.000 |
0.000 |
4.907 |
<r2> (average value of r
2) Å
2
| <r2> |
68.654 |
| (<r2>)1/2 |
8.286 |
