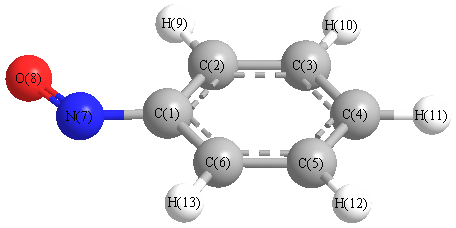
Vibrational Frequencies calculated at TPSSh/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3219 |
3086 |
9.71 |
|
|
|
| 2 |
A' |
3215 |
3082 |
15.42 |
|
|
|
| 3 |
A' |
3204 |
3072 |
13.38 |
|
|
|
| 4 |
A' |
3197 |
3065 |
6.59 |
|
|
|
| 5 |
A' |
3186 |
3054 |
1.14 |
|
|
|
| 6 |
A' |
1651 |
1583 |
1.60 |
|
|
|
| 7 |
A' |
1644 |
1576 |
1.71 |
|
|
|
| 8 |
A' |
1578 |
1513 |
162.85 |
|
|
|
| 9 |
A' |
1505 |
1443 |
9.30 |
|
|
|
| 10 |
A' |
1489 |
1428 |
43.86 |
|
|
|
| 11 |
A' |
1388 |
1331 |
13.05 |
|
|
|
| 12 |
A' |
1338 |
1282 |
7.23 |
|
|
|
| 13 |
A' |
1201 |
1151 |
12.85 |
|
|
|
| 14 |
A' |
1188 |
1139 |
0.71 |
|
|
|
| 15 |
A' |
1128 |
1082 |
140.36 |
|
|
|
| 16 |
A' |
1097 |
1051 |
6.63 |
|
|
|
| 17 |
A' |
1037 |
995 |
3.17 |
|
|
|
| 18 |
A' |
1012 |
970 |
0.98 |
|
|
|
| 19 |
A' |
830 |
795 |
31.72 |
|
|
|
| 20 |
A' |
673 |
645 |
8.36 |
|
|
|
| 21 |
A' |
612 |
587 |
0.05 |
|
|
|
| 22 |
A' |
440 |
422 |
0.71 |
|
|
|
| 23 |
A' |
249 |
239 |
2.51 |
|
|
|
| 24 |
A" |
1012 |
971 |
0.14 |
|
|
|
| 25 |
A" |
996 |
955 |
0.06 |
|
|
|
| 26 |
A" |
960 |
921 |
3.51 |
|
|
|
| 27 |
A" |
868 |
833 |
0.00 |
|
|
|
| 28 |
A" |
776 |
744 |
53.38 |
|
|
|
| 29 |
A" |
689 |
661 |
16.67 |
|
|
|
| 30 |
A" |
466 |
446 |
1.03 |
|
|
|
| 31 |
A" |
414 |
397 |
0.00 |
|
|
|
| 32 |
A" |
246 |
236 |
0.34 |
|
|
|
| 33 |
A" |
120 |
115 |
0.17 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21313.2 cm
-1
Scaled (by 0.9587) Zero Point Vibrational Energy (zpe) 20432.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at TPSSh/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.169 |
|
|
|
| 2 |
C |
-0.083 |
|
|
|
| 3 |
C |
-0.133 |
|
|
|
| 4 |
C |
-0.097 |
|
|
|
| 5 |
C |
-0.130 |
|
|
|
| 6 |
C |
-0.091 |
|
|
|
| 7 |
N |
-0.052 |
|
|
|
| 8 |
O |
-0.286 |
|
|
|
| 9 |
H |
0.152 |
|
|
|
| 10 |
H |
0.135 |
|
|
|
| 11 |
H |
0.134 |
|
|
|
| 12 |
H |
0.134 |
|
|
|
| 13 |
H |
0.147 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.202 |
-3.595 |
0.000 |
3.790 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
12.301 |
-1.776 |
0.000 |
| y |
-1.776 |
15.009 |
0.000 |
| z |
0.000 |
0.000 |
3.837 |
<r2> (average value of r
2) Å
2
| <r2> |
249.596 |
| (<r2>)1/2 |
15.799 |