Jump to
S1C2
Energy calculated at MP2=FULL/6-31G**
| | hartrees |
|---|
| Energy at 0K | -615.738742 |
| Energy at 298.15K | -615.745867 |
| HF Energy | -615.019888 |
| Nuclear repulsion energy | 206.194149 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3269 |
3054 |
6.73 |
|
|
|
| 2 |
A' |
3227 |
3015 |
24.30 |
|
|
|
| 3 |
A' |
3224 |
3013 |
3.45 |
|
|
|
| 4 |
A' |
3155 |
2948 |
19.62 |
|
|
|
| 5 |
A' |
3122 |
2917 |
23.40 |
|
|
|
| 6 |
A' |
1779 |
1662 |
0.39 |
|
|
|
| 7 |
A' |
1556 |
1454 |
8.73 |
|
|
|
| 8 |
A' |
1535 |
1434 |
2.66 |
|
|
|
| 9 |
A' |
1469 |
1373 |
3.09 |
|
|
|
| 10 |
A' |
1386 |
1295 |
33.35 |
|
|
|
| 11 |
A' |
1359 |
1270 |
3.56 |
|
|
|
| 12 |
A' |
1339 |
1251 |
0.56 |
|
|
|
| 13 |
A' |
1161 |
1085 |
0.03 |
|
|
|
| 14 |
A' |
1087 |
1016 |
8.21 |
|
|
|
| 15 |
A' |
948 |
885 |
7.84 |
|
|
|
| 16 |
A' |
758 |
708 |
16.04 |
|
|
|
| 17 |
A' |
601 |
561 |
2.46 |
|
|
|
| 18 |
A' |
343 |
320 |
0.86 |
|
|
|
| 19 |
A' |
180 |
168 |
0.69 |
|
|
|
| 20 |
A" |
3214 |
3003 |
7.09 |
|
|
|
| 21 |
A" |
3205 |
2995 |
18.34 |
|
|
|
| 22 |
A" |
1541 |
1440 |
5.20 |
|
|
|
| 23 |
A" |
1245 |
1163 |
2.22 |
|
|
|
| 24 |
A" |
1096 |
1024 |
0.42 |
|
|
|
| 25 |
A" |
1014 |
948 |
31.72 |
|
|
|
| 26 |
A" |
983 |
919 |
0.36 |
|
|
|
| 27 |
A" |
726 |
678 |
0.18 |
|
|
|
| 28 |
A" |
239 |
224 |
4.23 |
|
|
|
| 29 |
A" |
199 |
186 |
1.10 |
|
|
|
| 30 |
A" |
104 |
98 |
1.49 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 22531.6 cm
-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 21053.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.313 |
0.295 |
0.000 |
| C2 |
0.000 |
1.012 |
0.000 |
| C3 |
1.219 |
0.467 |
0.000 |
| C4 |
2.486 |
1.261 |
0.000 |
| Cl5 |
-1.191 |
-1.483 |
0.000 |
| H6 |
-1.894 |
0.568 |
0.879 |
| H7 |
-1.894 |
0.568 |
-0.879 |
| H8 |
-0.100 |
2.093 |
0.000 |
| H9 |
1.312 |
-0.612 |
0.000 |
| H10 |
2.283 |
2.330 |
0.000 |
| H11 |
3.091 |
1.030 |
0.876 |
| H12 |
3.091 |
1.030 |
-0.876 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
Cl5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
| C1 | | 1.4958 | 2.5377 | 3.9196 | 1.7822 | 1.0884 | 1.0884 | 2.1688 | 2.7766 | 4.1315 | 4.5496 | 4.5496 |
C2 | 1.4958 | | 1.3354 | 2.4983 | 2.7645 | 2.1345 | 2.1345 | 1.0858 | 2.0871 | 2.6359 | 3.2126 | 3.2126 | C3 | 2.5377 | 1.3354 | | 1.4954 | 3.0997 | 3.2362 | 3.2362 | 2.0941 | 1.0822 | 2.1455 | 2.1419 | 2.1419 | C4 | 3.9196 | 2.4983 | 1.4954 | | 4.5878 | 4.5205 | 4.5205 | 2.7165 | 2.2104 | 1.0880 | 1.0897 | 1.0897 | Cl5 | 1.7822 | 2.7645 | 3.0997 | 4.5878 | | 2.3397 | 2.3397 | 3.7387 | 2.6498 | 5.1579 | 5.0410 | 5.0410 | H6 | 1.0884 | 2.1345 | 3.2362 | 4.5205 | 2.3397 | | 1.7576 | 2.5130 | 3.5270 | 4.6173 | 5.0059 | 5.3046 | H7 | 1.0884 | 2.1345 | 3.2362 | 4.5205 | 2.3397 | 1.7576 | | 2.5130 | 3.5270 | 4.6173 | 5.3046 | 5.0059 | H8 | 2.1688 | 1.0858 | 2.0941 | 2.7165 | 3.7387 | 2.5130 | 2.5130 | | 3.0509 | 2.3945 | 3.4757 | 3.4757 | H9 | 2.7766 | 2.0871 | 1.0822 | 2.2104 | 2.6498 | 3.5270 | 3.5270 | 3.0509 | | 3.0977 | 2.5741 | 2.5741 | H10 | 4.1315 | 2.6359 | 2.1455 | 1.0880 | 5.1579 | 4.6173 | 4.6173 | 2.3945 | 3.0977 | | 1.7641 | 1.7641 | H11 | 4.5496 | 3.2126 | 2.1419 | 1.0897 | 5.0410 | 5.0059 | 5.3046 | 3.4757 | 2.5741 | 1.7641 | | 1.7525 | H12 | 4.5496 | 3.2126 | 2.1419 | 1.0897 | 5.0410 | 5.3046 | 5.0059 | 3.4757 | 2.5741 | 1.7641 | 1.7525 | |
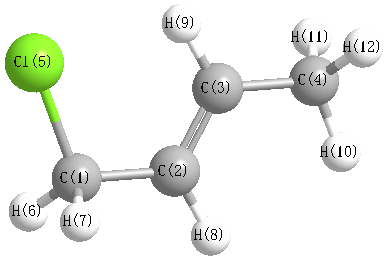 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
127.267 |
|
C1 |
C2 |
H8 |
113.347 |
| C2 |
C1 |
Cl5 |
114.710 |
|
C2 |
C1 |
H6 |
110.379 |
| C2 |
C1 |
H7 |
110.379 |
|
C2 |
C3 |
C4 |
123.812 |
| C2 |
C3 |
H9 |
119.004 |
|
C3 |
C2 |
H8 |
119.386 |
| C3 |
C4 |
H10 |
111.329 |
|
C3 |
C4 |
H11 |
110.935 |
| C3 |
C4 |
H12 |
110.935 |
|
C4 |
C3 |
H9 |
117.184 |
| Cl5 |
C1 |
H6 |
106.682 |
|
Cl5 |
C1 |
H7 |
106.682 |
| H6 |
C1 |
H7 |
107.691 |
|
H10 |
C4 |
H11 |
108.213 |
| H10 |
C4 |
H12 |
108.213 |
|
H11 |
C4 |
H12 |
107.058 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/6-31G**
| | hartrees |
|---|
| Energy at 0K | -615.740871 |
| Energy at 298.15K | -615.747977 |
| HF Energy | -615.022433 |
| Nuclear repulsion energy | 201.926333 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3254 |
3040 |
17.16 |
|
|
|
| 2 |
A |
3239 |
3027 |
2.33 |
|
|
|
| 3 |
A |
3237 |
3025 |
23.68 |
|
|
|
| 4 |
A |
3224 |
3012 |
3.41 |
|
|
|
| 5 |
A |
3206 |
2996 |
16.15 |
|
|
|
| 6 |
A |
3170 |
2962 |
17.73 |
|
|
|
| 7 |
A |
3124 |
2919 |
20.47 |
|
|
|
| 8 |
A |
1767 |
1651 |
6.44 |
|
|
|
| 9 |
A |
1554 |
1452 |
9.43 |
|
|
|
| 10 |
A |
1540 |
1439 |
6.01 |
|
|
|
| 11 |
A |
1537 |
1436 |
2.74 |
|
|
|
| 12 |
A |
1468 |
1372 |
2.31 |
|
|
|
| 13 |
A |
1390 |
1299 |
14.18 |
|
|
|
| 14 |
A |
1362 |
1273 |
3.82 |
|
|
|
| 15 |
A |
1338 |
1250 |
32.65 |
|
|
|
| 16 |
A |
1249 |
1167 |
1.89 |
|
|
|
| 17 |
A |
1165 |
1088 |
0.51 |
|
|
|
| 18 |
A |
1137 |
1062 |
2.69 |
|
|
|
| 19 |
A |
1096 |
1024 |
0.88 |
|
|
|
| 20 |
A |
1020 |
953 |
30.00 |
|
|
|
| 21 |
A |
982 |
918 |
8.13 |
|
|
|
| 22 |
A |
914 |
854 |
2.09 |
|
|
|
| 23 |
A |
815 |
762 |
24.68 |
|
|
|
| 24 |
A |
736 |
687 |
29.82 |
|
|
|
| 25 |
A |
501 |
468 |
1.52 |
|
|
|
| 26 |
A |
365 |
341 |
2.08 |
|
|
|
| 27 |
A |
281 |
263 |
1.84 |
|
|
|
| 28 |
A |
204 |
191 |
0.95 |
|
|
|
| 29 |
A |
161 |
150 |
1.84 |
|
|
|
| 30 |
A |
86 |
80 |
0.42 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 22560.6 cm
-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 21080.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31G**
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.780 |
0.817 |
0.116 |
| C2 |
0.513 |
0.167 |
0.454 |
| C3 |
1.595 |
0.241 |
-0.330 |
| C4 |
2.916 |
-0.380 |
-0.011 |
| Cl5 |
-2.089 |
-0.395 |
-0.106 |
| H6 |
-1.118 |
1.480 |
0.908 |
| H7 |
-0.704 |
1.381 |
-0.809 |
| H8 |
0.554 |
-0.382 |
1.388 |
| H9 |
1.523 |
0.786 |
-1.266 |
| H10 |
2.881 |
-0.906 |
0.940 |
| H11 |
3.207 |
-1.091 |
-0.784 |
| H12 |
3.701 |
0.374 |
0.044 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
Cl5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
| C1 | | 1.4862 | 2.4841 | 3.8869 | 1.7973 | 1.0869 | 1.0864 | 2.1983 | 2.6858 | 4.1289 | 4.5105 | 4.5028 |
C2 | 1.4862 | | 1.3382 | 2.5077 | 2.7205 | 2.1429 | 2.1337 | 1.0838 | 2.0885 | 2.6444 | 3.2205 | 3.2203 | C3 | 2.4841 | 1.3382 | | 1.4941 | 3.7455 | 3.2293 | 2.6108 | 2.1033 | 1.0854 | 2.1407 | 2.1398 | 2.1425 | C4 | 3.8869 | 2.5077 | 1.4941 | | 5.0061 | 4.5364 | 4.1043 | 2.7454 | 2.2081 | 1.0875 | 1.0896 | 1.0897 | Cl5 | 1.7973 | 2.7205 | 3.7455 | 5.0061 | | 2.3418 | 2.3589 | 3.0358 | 3.9734 | 5.1045 | 5.3848 | 5.8424 | H6 | 1.0869 | 2.1429 | 3.2293 | 4.5364 | 2.3418 | | 1.7689 | 2.5481 | 3.4901 | 4.6569 | 5.3084 | 5.0189 | H7 | 1.0864 | 2.1337 | 2.6108 | 4.1043 | 2.3589 | 1.7689 | | 3.0851 | 2.3501 | 4.5983 | 4.6270 | 4.5984 | H8 | 2.1983 | 1.0838 | 2.1033 | 2.7454 | 3.0358 | 2.5481 | 3.0851 | | 3.0572 | 2.4270 | 3.5015 | 3.5043 | H9 | 2.6858 | 2.0885 | 1.0854 | 2.2081 | 3.9734 | 3.4901 | 2.3501 | 3.0572 | | 3.0943 | 2.5674 | 2.5744 | H10 | 4.1289 | 2.6444 | 2.1407 | 1.0875 | 5.1045 | 4.6569 | 4.5983 | 2.4270 | 3.0943 | | 1.7645 | 1.7647 | H11 | 4.5105 | 3.2205 | 2.1398 | 1.0896 | 5.3848 | 5.3084 | 4.6270 | 3.5015 | 2.5674 | 1.7645 | | 1.7534 | H12 | 4.5028 | 3.2203 | 2.1425 | 1.0897 | 5.8424 | 5.0189 | 4.5984 | 3.5043 | 2.5744 | 1.7647 | 1.7534 | |
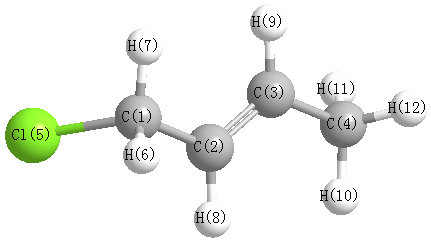 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
123.080 |
|
C1 |
C2 |
H8 |
116.735 |
| C2 |
C1 |
Cl5 |
111.548 |
|
C2 |
C1 |
H6 |
111.838 |
| C2 |
C1 |
H7 |
111.124 |
|
C2 |
C3 |
C4 |
124.503 |
| C2 |
C3 |
H9 |
118.645 |
|
C3 |
C2 |
H8 |
120.184 |
| C3 |
C4 |
H10 |
111.055 |
|
C3 |
C4 |
H11 |
110.854 |
| C3 |
C4 |
H12 |
111.067 |
|
C4 |
C3 |
H9 |
116.852 |
| Cl5 |
C1 |
H6 |
105.933 |
|
Cl5 |
C1 |
H7 |
107.187 |
| H6 |
C1 |
H7 |
108.965 |
|
H10 |
C4 |
H11 |
108.288 |
| H10 |
C4 |
H12 |
108.297 |
|
H11 |
C4 |
H12 |
107.138 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
