Jump to
S1C2
Energy calculated at HSEh1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -616.495382 |
| Energy at 298.15K | -616.502488 |
| HF Energy | -616.495382 |
| Nuclear repulsion energy | 205.742042 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3195 |
3049 |
8.11 |
|
|
|
| 2 |
A' |
3159 |
3015 |
13.15 |
|
|
|
| 3 |
A' |
3150 |
3006 |
14.59 |
|
|
|
| 4 |
A' |
3087 |
2946 |
19.74 |
|
|
|
| 5 |
A' |
3053 |
2913 |
26.90 |
|
|
|
| 6 |
A' |
1789 |
1707 |
0.85 |
|
|
|
| 7 |
A' |
1501 |
1432 |
11.62 |
|
|
|
| 8 |
A' |
1481 |
1413 |
2.91 |
|
|
|
| 9 |
A' |
1424 |
1359 |
3.20 |
|
|
|
| 10 |
A' |
1340 |
1278 |
14.98 |
|
|
|
| 11 |
A' |
1330 |
1270 |
22.39 |
|
|
|
| 12 |
A' |
1307 |
1247 |
2.06 |
|
|
|
| 13 |
A' |
1139 |
1087 |
0.17 |
|
|
|
| 14 |
A' |
1063 |
1014 |
12.88 |
|
|
|
| 15 |
A' |
925 |
883 |
10.79 |
|
|
|
| 16 |
A' |
731 |
698 |
21.70 |
|
|
|
| 17 |
A' |
594 |
566 |
4.52 |
|
|
|
| 18 |
A' |
338 |
323 |
1.22 |
|
|
|
| 19 |
A' |
173 |
165 |
0.85 |
|
|
|
| 20 |
A" |
3138 |
2994 |
6.29 |
|
|
|
| 21 |
A" |
3117 |
2975 |
17.68 |
|
|
|
| 22 |
A" |
1488 |
1420 |
7.29 |
|
|
|
| 23 |
A" |
1208 |
1153 |
1.98 |
|
|
|
| 24 |
A" |
1071 |
1022 |
0.16 |
|
|
|
| 25 |
A" |
1001 |
955 |
34.63 |
|
|
|
| 26 |
A" |
956 |
912 |
|
|
|
|
| 27 |
A" |
733 |
699 |
0.42 |
|
|
|
| 28 |
A" |
244 |
233 |
4.36 |
|
|
|
| 29 |
A" |
209 |
199 |
2.36 |
|
|
|
| 30 |
A" |
120 |
114 |
1.50 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 22030.9 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 21024.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.311 |
0.298 |
0.000 |
| C2 |
0.000 |
1.014 |
0.000 |
| C3 |
1.218 |
0.478 |
0.000 |
| C4 |
2.488 |
1.263 |
0.000 |
| Cl5 |
-1.192 |
-1.490 |
0.000 |
| H6 |
-1.900 |
0.566 |
0.883 |
| H7 |
-1.900 |
0.566 |
-0.883 |
| H8 |
-0.109 |
2.100 |
0.000 |
| H9 |
1.308 |
-0.607 |
0.000 |
| H10 |
2.301 |
2.340 |
0.000 |
| H11 |
3.098 |
1.023 |
0.879 |
| H12 |
3.098 |
1.023 |
-0.879 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
Cl5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
| C1 | | 1.4936 | 2.5349 | 3.9196 | 1.7924 | 1.0943 | 1.0943 | 2.1658 | 2.7711 | 4.1493 | 4.5540 | 4.5540 |
C2 | 1.4936 | | 1.3302 | 2.5005 | 2.7734 | 2.1424 | 2.1424 | 1.0917 | 2.0829 | 2.6560 | 3.2206 | 3.2206 | C3 | 2.5349 | 1.3302 | | 1.4933 | 3.1114 | 3.2414 | 3.2414 | 2.0955 | 1.0889 | 2.1544 | 2.1462 | 2.1462 | C4 | 3.9196 | 2.5005 | 1.4933 | | 4.5959 | 4.5300 | 4.5300 | 2.7290 | 2.2109 | 1.0937 | 1.0964 | 1.0964 | Cl5 | 1.7924 | 2.7734 | 3.1114 | 4.5959 | | 2.3467 | 2.3467 | 3.7500 | 2.6517 | 5.1841 | 5.0493 | 5.0493 | H6 | 1.0943 | 2.1424 | 3.2414 | 4.5300 | 2.3467 | | 1.7651 | 2.5179 | 3.5281 | 4.6452 | 5.0193 | 5.3193 | H7 | 1.0943 | 2.1424 | 3.2414 | 4.5300 | 2.3467 | 1.7651 | | 2.5179 | 3.5281 | 4.6452 | 5.3193 | 5.0193 | H8 | 2.1658 | 1.0917 | 2.0955 | 2.7290 | 3.7500 | 2.5179 | 2.5179 | | 3.0557 | 2.4223 | 3.4960 | 3.4960 | H9 | 2.7711 | 2.0829 | 1.0889 | 2.2109 | 2.6517 | 3.5281 | 3.5281 | 3.0557 | | 3.1100 | 2.5754 | 2.5754 | H10 | 4.1493 | 2.6560 | 2.1544 | 1.0937 | 5.1841 | 4.6452 | 4.6452 | 2.4223 | 3.1100 | | 1.7731 | 1.7731 | H11 | 4.5540 | 3.2206 | 2.1462 | 1.0964 | 5.0493 | 5.0193 | 5.3193 | 3.4960 | 2.5754 | 1.7731 | | 1.7573 | H12 | 4.5540 | 3.2206 | 2.1462 | 1.0964 | 5.0493 | 5.3193 | 5.0193 | 3.4960 | 2.5754 | 1.7731 | 1.7573 | |
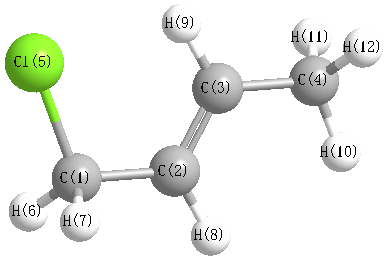 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
127.619 |
|
C1 |
C2 |
H8 |
112.885 |
| C2 |
C1 |
Cl5 |
114.830 |
|
C2 |
C1 |
H6 |
110.817 |
| C2 |
C1 |
H7 |
110.817 |
|
C2 |
C3 |
C4 |
124.553 |
| C2 |
C3 |
H9 |
118.523 |
|
C3 |
C2 |
H8 |
119.496 |
| C3 |
C4 |
H10 |
111.850 |
|
C3 |
C4 |
H11 |
111.013 |
| C3 |
C4 |
H12 |
111.013 |
|
C4 |
C3 |
H9 |
116.925 |
| Cl5 |
C1 |
H6 |
106.240 |
|
Cl5 |
C1 |
H7 |
106.240 |
| H6 |
C1 |
H7 |
107.501 |
|
H10 |
C4 |
H11 |
108.114 |
| H10 |
C4 |
H12 |
108.114 |
|
H11 |
C4 |
H12 |
106.525 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.388 |
|
|
|
| 2 |
C |
-0.101 |
|
|
|
| 3 |
C |
-0.074 |
|
|
|
| 4 |
C |
-0.432 |
|
|
|
| 5 |
Cl |
-0.070 |
|
|
|
| 6 |
H |
0.188 |
|
|
|
| 7 |
H |
0.188 |
|
|
|
| 8 |
H |
0.123 |
|
|
|
| 9 |
H |
0.140 |
|
|
|
| 10 |
H |
0.134 |
|
|
|
| 11 |
H |
0.147 |
|
|
|
| 12 |
H |
0.147 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.340 |
2.000 |
0.000 |
2.028 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.041 |
-2.560 |
0.000 |
| y |
-2.560 |
-38.164 |
0.000 |
| z |
0.000 |
0.000 |
-39.013 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.547 |
-2.560 |
0.000 |
| y |
-2.560 |
-1.137 |
0.000 |
| z |
0.000 |
0.000 |
-2.410 |
|
| Polar |
|---|
| 3z2-r2 | -4.821 |
|---|
| x2-y2 | 3.123 |
|---|
| xy | -2.560 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.863 |
0.415 |
0.000 |
| y |
0.415 |
8.387 |
0.000 |
| z |
0.000 |
0.000 |
4.931 |
<r2> (average value of r
2) Å
2
| <r2> |
208.793 |
| (<r2>)1/2 |
14.450 |
Jump to
S1C1
Energy calculated at HSEh1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -616.497221 |
| Energy at 298.15K | -616.504274 |
| HF Energy | -616.497221 |
| Nuclear repulsion energy | 201.278616 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3192 |
3046 |
16.90 |
|
|
|
| 2 |
A |
3175 |
3030 |
1.69 |
|
|
|
| 3 |
A |
3168 |
3023 |
27.09 |
|
|
|
| 4 |
A |
3152 |
3008 |
1.82 |
|
|
|
| 5 |
A |
3119 |
2976 |
15.17 |
|
|
|
| 6 |
A |
3110 |
2968 |
16.67 |
|
|
|
| 7 |
A |
3055 |
2915 |
22.57 |
|
|
|
| 8 |
A |
1771 |
1690 |
13.86 |
|
|
|
| 9 |
A |
1499 |
1430 |
14.28 |
|
|
|
| 10 |
A |
1489 |
1421 |
2.91 |
|
|
|
| 11 |
A |
1487 |
1419 |
8.69 |
|
|
|
| 12 |
A |
1422 |
1357 |
2.37 |
|
|
|
| 13 |
A |
1347 |
1286 |
6.57 |
|
|
|
| 14 |
A |
1335 |
1274 |
1.39 |
|
|
|
| 15 |
A |
1286 |
1228 |
41.27 |
|
|
|
| 16 |
A |
1213 |
1158 |
3.21 |
|
|
|
| 17 |
A |
1133 |
1081 |
0.72 |
|
|
|
| 18 |
A |
1118 |
1067 |
2.36 |
|
|
|
| 19 |
A |
1069 |
1020 |
0.42 |
|
|
|
| 20 |
A |
1009 |
963 |
32.27 |
|
|
|
| 21 |
A |
956 |
913 |
12.67 |
|
|
|
| 22 |
A |
899 |
858 |
2.93 |
|
|
|
| 23 |
A |
806 |
770 |
15.00 |
|
|
|
| 24 |
A |
701 |
669 |
62.94 |
|
|
|
| 25 |
A |
495 |
472 |
1.75 |
|
|
|
| 26 |
A |
357 |
341 |
4.52 |
|
|
|
| 27 |
A |
278 |
266 |
2.23 |
|
|
|
| 28 |
A |
215 |
205 |
1.18 |
|
|
|
| 29 |
A |
160 |
153 |
2.60 |
|
|
|
| 30 |
A |
89 |
85 |
0.33 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 22052.9 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 21045.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G**
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.772 |
0.821 |
0.112 |
| C2 |
0.519 |
0.172 |
0.452 |
| C3 |
1.600 |
0.233 |
-0.327 |
| C4 |
2.922 |
-0.380 |
-0.010 |
| Cl5 |
-2.100 |
-0.395 |
-0.103 |
| H6 |
-1.122 |
1.485 |
0.905 |
| H7 |
-0.709 |
1.382 |
-0.822 |
| H8 |
0.556 |
-0.370 |
1.397 |
| H9 |
1.529 |
0.771 |
-1.274 |
| H10 |
2.904 |
-0.902 |
0.950 |
| H11 |
3.216 |
-1.098 |
-0.785 |
| H12 |
3.712 |
0.379 |
0.028 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
Cl5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
| C1 | | 1.4846 | 2.4823 | 3.8861 | 1.8139 | 1.0923 | 1.0913 | 2.1982 | 2.6861 | 4.1455 | 4.5152 | 4.5059 |
C2 | 1.4846 | | 1.3338 | 2.5084 | 2.7367 | 2.1502 | 2.1444 | 1.0892 | 2.0877 | 2.6626 | 3.2270 | 3.2274 | C3 | 2.4823 | 1.3338 | | 1.4919 | 3.7592 | 3.2393 | 2.6261 | 2.1034 | 1.0913 | 2.1502 | 2.1432 | 2.1467 | C4 | 3.8861 | 2.5084 | 1.4919 | | 5.0229 | 4.5464 | 4.1173 | 2.7525 | 2.2057 | 1.0931 | 1.0964 | 1.0964 | Cl5 | 1.8139 | 2.7367 | 3.7592 | 5.0229 | | 2.3475 | 2.3688 | 3.0504 | 3.9873 | 5.1388 | 5.4049 | 5.8645 | H6 | 1.0923 | 2.1502 | 3.2393 | 4.5464 | 2.3475 | | 1.7790 | 2.5489 | 3.5050 | 4.6808 | 5.3238 | 5.0354 | H7 | 1.0913 | 2.1444 | 2.6261 | 4.1173 | 2.3688 | 1.7790 | | 3.0973 | 2.3635 | 4.6279 | 4.6432 | 4.6122 | H8 | 2.1982 | 1.0892 | 2.1034 | 2.7525 | 3.0504 | 2.5489 | 3.0973 | | 3.0627 | 2.4487 | 3.5159 | 3.5204 | H9 | 2.6861 | 2.0877 | 1.0913 | 2.2057 | 3.9873 | 3.5050 | 2.3635 | 3.0627 | | 3.1049 | 2.5652 | 2.5717 | H10 | 4.1455 | 2.6626 | 2.1502 | 1.0931 | 5.1388 | 4.6808 | 4.6279 | 2.4487 | 3.1049 | | 1.7736 | 1.7738 | H11 | 4.5152 | 3.2270 | 2.1432 | 1.0964 | 5.4049 | 5.3238 | 4.6432 | 3.5159 | 2.5652 | 1.7736 | | 1.7578 | H12 | 4.5059 | 3.2274 | 2.1467 | 1.0964 | 5.8645 | 5.0354 | 4.6122 | 3.5204 | 2.5717 | 1.7738 | 1.7578 | |
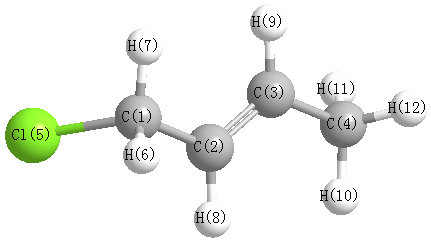 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
123.381 |
|
C1 |
C2 |
H8 |
116.472 |
| C2 |
C1 |
Cl5 |
111.740 |
|
C2 |
C1 |
H6 |
112.210 |
| C2 |
C1 |
H7 |
111.801 |
|
C2 |
C3 |
C4 |
125.083 |
| C2 |
C3 |
H9 |
118.500 |
|
C3 |
C2 |
H8 |
120.146 |
| C3 |
C4 |
H10 |
111.637 |
|
C3 |
C4 |
H11 |
110.869 |
| C3 |
C4 |
H12 |
111.151 |
|
C4 |
C3 |
H9 |
116.417 |
| Cl5 |
C1 |
H6 |
105.022 |
|
Cl5 |
C1 |
H7 |
106.584 |
| H6 |
C1 |
H7 |
109.112 |
|
H10 |
C4 |
H11 |
108.199 |
| H10 |
C4 |
H12 |
108.219 |
|
H11 |
C4 |
H12 |
106.570 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.393 |
|
|
|
| 2 |
C |
-0.068 |
|
|
|
| 3 |
C |
-0.081 |
|
|
|
| 4 |
C |
-0.430 |
|
|
|
| 5 |
Cl |
-0.086 |
|
|
|
| 6 |
H |
0.189 |
|
|
|
| 7 |
H |
0.180 |
|
|
|
| 8 |
H |
0.130 |
|
|
|
| 9 |
H |
0.120 |
|
|
|
| 10 |
H |
0.142 |
|
|
|
| 11 |
H |
0.151 |
|
|
|
| 12 |
H |
0.147 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.316 |
1.248 |
0.155 |
2.636 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-39.274 |
-2.052 |
-1.022 |
| y |
-2.052 |
-37.424 |
-0.838 |
| z |
-1.022 |
-0.838 |
-36.821 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.152 |
-2.052 |
-1.022 |
| y |
-2.052 |
0.623 |
-0.838 |
| z |
-1.022 |
-0.838 |
1.528 |
|
| Polar |
|---|
| 3z2-r2 | 3.057 |
|---|
| x2-y2 | -1.850 |
|---|
| xy | -2.052 |
|---|
| xz | -1.022 |
|---|
| yz | -0.838 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.554 |
0.223 |
-0.838 |
| y |
0.223 |
6.110 |
-0.367 |
| z |
-0.838 |
-0.367 |
6.406 |
<r2> (average value of r
2) Å
2
| <r2> |
231.131 |
| (<r2>)1/2 |
15.203 |
