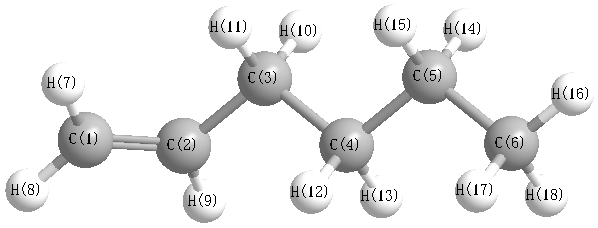
Vibrational Frequencies calculated at PBE1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3264 |
3111 |
18.37 |
|
|
|
| 2 |
A |
3180 |
3031 |
5.12 |
|
|
|
| 3 |
A |
3163 |
3015 |
27.30 |
|
|
|
| 4 |
A |
3148 |
3000 |
35.30 |
|
|
|
| 5 |
A |
3143 |
2996 |
51.27 |
|
|
|
| 6 |
A |
3111 |
2965 |
48.67 |
|
|
|
| 7 |
A |
3093 |
2948 |
2.18 |
|
|
|
| 8 |
A |
3080 |
2936 |
6.08 |
|
|
|
| 9 |
A |
3066 |
2922 |
36.21 |
|
|
|
| 10 |
A |
3057 |
2913 |
57.93 |
|
|
|
| 11 |
A |
3048 |
2905 |
1.54 |
|
|
|
| 12 |
A |
3038 |
2895 |
14.73 |
|
|
|
| 13 |
A |
1753 |
1671 |
11.14 |
|
|
|
| 14 |
A |
1536 |
1464 |
6.19 |
|
|
|
| 15 |
A |
1526 |
1455 |
7.62 |
|
|
|
| 16 |
A |
1523 |
1452 |
1.72 |
|
|
|
| 17 |
A |
1512 |
1441 |
0.43 |
|
|
|
| 18 |
A |
1504 |
1434 |
2.27 |
|
|
|
| 19 |
A |
1471 |
1402 |
1.03 |
|
|
|
| 20 |
A |
1437 |
1370 |
3.24 |
|
|
|
| 21 |
A |
1421 |
1354 |
0.47 |
|
|
|
| 22 |
A |
1370 |
1306 |
0.66 |
|
|
|
| 23 |
A |
1347 |
1283 |
0.67 |
|
|
|
| 24 |
A |
1334 |
1272 |
0.20 |
|
|
|
| 25 |
A |
1326 |
1264 |
0.51 |
|
|
|
| 26 |
A |
1279 |
1219 |
2.59 |
|
|
|
| 27 |
A |
1261 |
1202 |
0.15 |
|
|
|
| 28 |
A |
1211 |
1155 |
0.37 |
|
|
|
| 29 |
A |
1142 |
1088 |
2.99 |
|
|
|
| 30 |
A |
1094 |
1042 |
0.20 |
|
|
|
| 31 |
A |
1065 |
1015 |
6.20 |
|
|
|
| 32 |
A |
1051 |
1002 |
3.23 |
|
|
|
| 33 |
A |
1039 |
990 |
8.79 |
|
|
|
| 34 |
A |
956 |
912 |
2.25 |
|
|
|
| 35 |
A |
942 |
898 |
40.37 |
|
|
|
| 36 |
A |
932 |
889 |
3.40 |
|
|
|
| 37 |
A |
919 |
876 |
1.99 |
|
|
|
| 38 |
A |
802 |
764 |
1.08 |
|
|
|
| 39 |
A |
741 |
706 |
4.05 |
|
|
|
| 40 |
A |
655 |
624 |
10.13 |
|
|
|
| 41 |
A |
458 |
437 |
1.12 |
|
|
|
| 42 |
A |
361 |
344 |
0.07 |
|
|
|
| 43 |
A |
354 |
337 |
0.47 |
|
|
|
| 44 |
A |
249 |
237 |
0.00 |
|
|
|
| 45 |
A |
175 |
167 |
0.02 |
|
|
|
| 46 |
A |
125 |
119 |
0.07 |
|
|
|
| 47 |
A |
90 |
86 |
0.02 |
|
|
|
| 48 |
A |
73 |
70 |
0.02 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36711.5 cm
-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 34989.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.397 |
|
|
|
| 2 |
C |
-0.077 |
|
|
|
| 3 |
C |
-0.370 |
|
|
|
| 4 |
C |
-0.314 |
|
|
|
| 5 |
C |
-0.317 |
|
|
|
| 6 |
C |
-0.520 |
|
|
|
| 7 |
H |
0.162 |
|
|
|
| 8 |
H |
0.166 |
|
|
|
| 9 |
H |
0.158 |
|
|
|
| 10 |
H |
0.172 |
|
|
|
| 11 |
H |
0.168 |
|
|
|
| 12 |
H |
0.170 |
|
|
|
| 13 |
H |
0.160 |
|
|
|
| 14 |
H |
0.162 |
|
|
|
| 15 |
H |
0.164 |
|
|
|
| 16 |
H |
0.172 |
|
|
|
| 17 |
H |
0.170 |
|
|
|
| 18 |
H |
0.169 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.369 |
0.053 |
0.107 |
0.388 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-40.363 |
-0.883 |
-0.472 |
| y |
-0.883 |
-40.014 |
-1.142 |
| z |
-0.472 |
-1.142 |
-38.708 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.002 |
-0.883 |
-0.472 |
| y |
-0.883 |
-0.479 |
-1.142 |
| z |
-0.472 |
-1.142 |
1.481 |
|
| Polar |
|---|
| 3z2-r2 | 2.962 |
|---|
| x2-y2 | -0.348 |
|---|
| xy | -0.883 |
|---|
| xz | -0.472 |
|---|
| yz | -1.142 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
12.279 |
-0.485 |
-1.132 |
| y |
-0.485 |
7.816 |
-0.548 |
| z |
-1.132 |
-0.548 |
8.430 |
<r2> (average value of r
2) Å
2
| <r2> |
284.254 |
| (<r2>)1/2 |
16.860 |