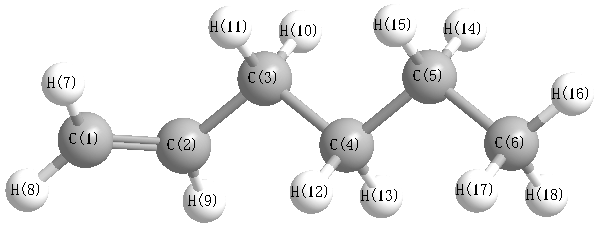
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3261 |
3112 |
17.74 |
|
|
|
| 2 |
A |
3172 |
3027 |
4.56 |
|
|
|
| 3 |
A |
3157 |
3013 |
26.81 |
|
|
|
| 4 |
A |
3145 |
3001 |
34.57 |
|
|
|
| 5 |
A |
3139 |
2996 |
49.81 |
|
|
|
| 6 |
A |
3107 |
2965 |
49.45 |
|
|
|
| 7 |
A |
3089 |
2948 |
1.90 |
|
|
|
| 8 |
A |
3076 |
2936 |
5.05 |
|
|
|
| 9 |
A |
3059 |
2919 |
38.26 |
|
|
|
| 10 |
A |
3051 |
2911 |
57.74 |
|
|
|
| 11 |
A |
3042 |
2903 |
1.64 |
|
|
|
| 12 |
A |
3032 |
2893 |
14.54 |
|
|
|
| 13 |
A |
1748 |
1668 |
11.47 |
|
|
|
| 14 |
A |
1519 |
1449 |
5.51 |
|
|
|
| 15 |
A |
1509 |
1440 |
6.94 |
|
|
|
| 16 |
A |
1506 |
1437 |
1.57 |
|
|
|
| 17 |
A |
1495 |
1427 |
0.42 |
|
|
|
| 18 |
A |
1487 |
1419 |
2.37 |
|
|
|
| 19 |
A |
1459 |
1392 |
1.01 |
|
|
|
| 20 |
A |
1422 |
1357 |
2.89 |
|
|
|
| 21 |
A |
1410 |
1345 |
0.43 |
|
|
|
| 22 |
A |
1359 |
1297 |
0.74 |
|
|
|
| 23 |
A |
1337 |
1276 |
0.70 |
|
|
|
| 24 |
A |
1325 |
1265 |
0.18 |
|
|
|
| 25 |
A |
1316 |
1256 |
0.47 |
|
|
|
| 26 |
A |
1268 |
1210 |
2.70 |
|
|
|
| 27 |
A |
1251 |
1194 |
0.15 |
|
|
|
| 28 |
A |
1203 |
1148 |
0.41 |
|
|
|
| 29 |
A |
1135 |
1083 |
2.83 |
|
|
|
| 30 |
A |
1088 |
1038 |
0.20 |
|
|
|
| 31 |
A |
1059 |
1011 |
6.50 |
|
|
|
| 32 |
A |
1046 |
998 |
4.22 |
|
|
|
| 33 |
A |
1037 |
989 |
6.69 |
|
|
|
| 34 |
A |
952 |
909 |
4.49 |
|
|
|
| 35 |
A |
942 |
899 |
36.44 |
|
|
|
| 36 |
A |
927 |
884 |
2.63 |
|
|
|
| 37 |
A |
914 |
872 |
1.80 |
|
|
|
| 38 |
A |
797 |
761 |
1.07 |
|
|
|
| 39 |
A |
737 |
704 |
3.96 |
|
|
|
| 40 |
A |
654 |
624 |
9.63 |
|
|
|
| 41 |
A |
458 |
437 |
1.13 |
|
|
|
| 42 |
A |
361 |
344 |
0.06 |
|
|
|
| 43 |
A |
354 |
338 |
0.46 |
|
|
|
| 44 |
A |
248 |
237 |
0.00 |
|
|
|
| 45 |
A |
175 |
167 |
0.02 |
|
|
|
| 46 |
A |
125 |
119 |
0.06 |
|
|
|
| 47 |
A |
90 |
86 |
0.02 |
|
|
|
| 48 |
A |
73 |
69 |
0.02 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36557.0 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 34886.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.297 |
|
|
|
| 2 |
C |
-0.054 |
|
|
|
| 3 |
C |
-0.275 |
|
|
|
| 4 |
C |
-0.227 |
|
|
|
| 5 |
C |
-0.238 |
|
|
|
| 6 |
C |
-0.391 |
|
|
|
| 7 |
H |
0.117 |
|
|
|
| 8 |
H |
0.122 |
|
|
|
| 9 |
H |
0.113 |
|
|
|
| 10 |
H |
0.131 |
|
|
|
| 11 |
H |
0.124 |
|
|
|
| 12 |
H |
0.128 |
|
|
|
| 13 |
H |
0.119 |
|
|
|
| 14 |
H |
0.121 |
|
|
|
| 15 |
H |
0.123 |
|
|
|
| 16 |
H |
0.128 |
|
|
|
| 17 |
H |
0.129 |
|
|
|
| 18 |
H |
0.128 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.380 |
0.053 |
0.107 |
0.398 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-40.314 |
-0.828 |
-0.441 |
| y |
-0.828 |
-39.979 |
-1.108 |
| z |
-0.441 |
-1.108 |
-38.782 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.934 |
-0.828 |
-0.441 |
| y |
-0.828 |
-0.430 |
-1.108 |
| z |
-0.441 |
-1.108 |
1.364 |
|
| Polar |
|---|
| 3z2-r2 | 2.728 |
|---|
| x2-y2 | -0.335 |
|---|
| xy | -0.828 |
|---|
| xz | -0.441 |
|---|
| yz | -1.108 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
12.490 |
-0.474 |
-1.127 |
| y |
-0.474 |
7.951 |
-0.551 |
| z |
-1.127 |
-0.551 |
8.542 |
<r2> (average value of r
2) Å
2
| <r2> |
284.187 |
| (<r2>)1/2 |
16.858 |