Jump to
S1C2
Energy calculated at PBE1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -190.431378 |
| Energy at 298.15K | |
| HF Energy | -190.431378 |
| Nuclear repulsion energy | 88.274161 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBE1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3120 |
2974 |
28.05 |
191.17 |
0.11 |
0.20 |
| 2 |
A1 |
2374 |
2262 |
886.21 |
30.54 |
0.35 |
0.51 |
| 3 |
A1 |
1837 |
1750 |
25.08 |
0.62 |
0.52 |
0.69 |
| 4 |
A1 |
1501 |
1431 |
4.12 |
17.56 |
0.52 |
0.69 |
| 5 |
A1 |
948 |
904 |
6.37 |
39.68 |
0.28 |
0.44 |
| 6 |
B1 |
1019 |
971 |
25.91 |
0.13 |
0.75 |
0.86 |
| 7 |
B1 |
657 |
626 |
22.71 |
2.29 |
0.75 |
0.86 |
| 8 |
B1 |
234 |
223 |
0.03 |
4.71 |
0.75 |
0.86 |
| 9 |
B2 |
3190 |
3041 |
11.64 |
126.28 |
0.75 |
0.86 |
| 10 |
B2 |
1074 |
1024 |
1.03 |
0.34 |
0.75 |
0.86 |
| 11 |
B2 |
455 |
433 |
16.23 |
1.90 |
0.75 |
0.86 |
| 12 |
B2 |
77i |
74i |
15.85 |
0.08 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 8165.8 cm
-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 7782.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBE1PBE/6-31G**
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-1.874 |
| C2 |
0.000 |
0.000 |
-0.558 |
| C3 |
0.000 |
0.000 |
0.720 |
| O4 |
0.000 |
0.000 |
1.898 |
| H5 |
0.000 |
0.924 |
-2.457 |
| H6 |
0.000 |
-0.924 |
-2.457 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
| C1 | | 1.3165 | 2.5944 | 3.7728 | 1.0926 | 1.0926 |
C2 | 1.3165 | | 1.2779 | 2.4564 | 2.1119 | 2.1119 | C3 | 2.5944 | 1.2779 | | 1.1784 | 3.3085 | 3.3085 | O4 | 3.7728 | 2.4564 | 1.1784 | | 4.4522 | 4.4522 | H5 | 1.0926 | 2.1119 | 3.3085 | 4.4522 | | 1.8490 | H6 | 1.0926 | 2.1119 | 3.3085 | 4.4522 | 1.8490 | |
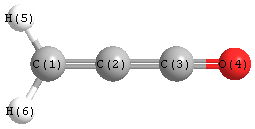 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
180.000 |
|
C2 |
C1 |
H5 |
122.207 |
| C2 |
C1 |
H6 |
122.207 |
|
C2 |
C3 |
O4 |
180.000 |
| H5 |
C1 |
H6 |
115.587 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.466 |
|
|
|
| 2 |
C |
0.266 |
|
|
|
| 3 |
C |
0.176 |
|
|
|
| 4 |
O |
-0.389 |
|
|
|
| 5 |
H |
0.207 |
|
|
|
| 6 |
H |
0.207 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-2.707 |
2.707 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-22.182 |
0.000 |
0.000 |
| y |
0.000 |
-21.954 |
0.000 |
| z |
0.000 |
0.000 |
-21.282 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.564 |
0.000 |
0.000 |
| y |
0.000 |
-0.222 |
0.000 |
| z |
0.000 |
0.000 |
0.786 |
|
| Polar |
|---|
| 3z2-r2 | 1.572 |
|---|
| x2-y2 | -0.228 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
1.862 |
0.000 |
0.000 |
| y |
0.000 |
2.724 |
0.000 |
| z |
0.000 |
0.000 |
10.110 |
<r2> (average value of r
2) Å
2
| <r2> |
82.292 |
| (<r2>)1/2 |
9.071 |
Jump to
S1C1
Energy calculated at PBE1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -190.431749 |
| Energy at 298.15K | |
| HF Energy | -190.431749 |
| Nuclear repulsion energy | 88.421005 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBE1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3204 |
3054 |
11.44 |
119.37 |
0.69 |
0.82 |
| 2 |
A' |
3127 |
2981 |
26.98 |
157.75 |
0.12 |
0.21 |
| 3 |
A' |
2300 |
2192 |
891.33 |
34.82 |
0.36 |
0.53 |
| 4 |
A' |
1815 |
1730 |
10.88 |
1.08 |
0.65 |
0.79 |
| 5 |
A' |
1505 |
1435 |
1.75 |
18.53 |
0.55 |
0.71 |
| 6 |
A' |
1076 |
1026 |
11.75 |
4.20 |
0.16 |
0.28 |
| 7 |
A' |
966 |
921 |
4.10 |
33.69 |
0.23 |
0.37 |
| 8 |
A' |
486 |
463 |
15.70 |
3.95 |
0.60 |
0.75 |
| 9 |
A' |
132 |
126 |
20.21 |
5.07 |
0.74 |
0.85 |
| 10 |
A" |
1024 |
976 |
25.82 |
0.23 |
0.75 |
0.86 |
| 11 |
A" |
693 |
660 |
17.86 |
2.78 |
0.75 |
0.86 |
| 12 |
A" |
261 |
249 |
1.51 |
6.28 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 8294.6 cm
-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 7905.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBE1PBE/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.881 |
-1.614 |
0.000 |
| C2 |
0.000 |
-0.626 |
0.000 |
| C3 |
-0.296 |
0.635 |
0.000 |
| O4 |
-0.754 |
1.716 |
0.000 |
| H5 |
1.961 |
-1.448 |
0.000 |
| H6 |
0.565 |
-2.658 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
| C1 | | 1.3233 | 2.5382 | 3.7096 | 1.0928 | 1.0909 |
C2 | 1.3233 | | 1.2955 | 2.4607 | 2.1263 | 2.1089 | C3 | 2.5382 | 1.2955 | | 1.1741 | 3.0716 | 3.4038 | O4 | 3.7096 | 2.4607 | 1.1741 | | 4.1695 | 4.5687 | H5 | 1.0928 | 2.1263 | 3.0716 | 4.1695 | | 1.8471 | H6 | 1.0909 | 2.1089 | 3.4038 | 4.5687 | 1.8471 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
151.490 |
|
C2 |
C1 |
H5 |
123.013 |
| C2 |
C1 |
H6 |
121.452 |
|
C2 |
C3 |
O4 |
170.260 |
| H5 |
C1 |
H6 |
115.535 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.397 |
|
|
|
| 2 |
C |
0.018 |
|
|
|
| 3 |
C |
0.343 |
|
|
|
| 4 |
O |
-0.373 |
|
|
|
| 5 |
H |
0.205 |
|
|
|
| 6 |
H |
0.203 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.673 |
-1.888 |
0.000 |
2.523 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-21.489 |
-0.839 |
0.000 |
| y |
-0.839 |
-22.020 |
0.000 |
| z |
0.000 |
0.000 |
-22.268 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.655 |
-0.839 |
0.000 |
| y |
-0.839 |
-0.141 |
0.000 |
| z |
0.000 |
0.000 |
-0.514 |
|
| Polar |
|---|
| 3z2-r2 | -1.028 |
|---|
| x2-y2 | 0.531 |
|---|
| xy | -0.839 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.287 |
-2.596 |
0.000 |
| y |
-2.596 |
8.575 |
0.000 |
| z |
0.000 |
0.000 |
1.909 |
<r2> (average value of r
2) Å
2
| <r2> |
80.710 |
| (<r2>)1/2 |
8.984 |
