Jump to
S1C2
Energy calculated at HSEh1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -190.451667 |
| Energy at 298.15K | |
| HF Energy | -190.451667 |
| Nuclear repulsion energy | 88.291960 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3108 |
2966 |
27.12 |
195.71 |
0.11 |
0.20 |
| 2 |
A1 |
2371 |
2262 |
880.07 |
30.42 |
0.35 |
0.51 |
| 3 |
A1 |
1834 |
1750 |
25.89 |
0.60 |
0.54 |
0.70 |
| 4 |
A1 |
1487 |
1419 |
3.61 |
17.56 |
0.52 |
0.69 |
| 5 |
A1 |
947 |
904 |
6.50 |
40.07 |
0.28 |
0.44 |
| 6 |
B1 |
1018 |
972 |
24.34 |
0.09 |
0.75 |
0.86 |
| 7 |
B1 |
658 |
628 |
22.77 |
2.22 |
0.75 |
0.86 |
| 8 |
B1 |
235 |
224 |
0.03 |
4.66 |
0.75 |
0.86 |
| 9 |
B2 |
3182 |
3036 |
11.11 |
128.23 |
0.75 |
0.86 |
| 10 |
B2 |
1065 |
1016 |
1.15 |
0.34 |
0.75 |
0.86 |
| 11 |
B2 |
454 |
434 |
16.11 |
1.92 |
0.75 |
0.86 |
| 12 |
B2 |
73i |
70i |
15.54 |
0.09 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 8141.5 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 7769.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G**
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-1.874 |
| C2 |
0.000 |
0.000 |
-0.558 |
| C3 |
0.000 |
0.000 |
0.720 |
| O4 |
0.000 |
0.000 |
1.898 |
| H5 |
0.000 |
0.925 |
-2.455 |
| H6 |
0.000 |
-0.925 |
-2.455 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
| C1 | | 1.3161 | 2.5937 | 3.7722 | 1.0920 | 1.0920 |
C2 | 1.3161 | | 1.2776 | 2.4562 | 2.1104 | 2.1104 | C3 | 2.5937 | 1.2776 | | 1.1786 | 3.3065 | 3.3065 | O4 | 3.7722 | 2.4562 | 1.1786 | | 4.4503 | 4.4503 | H5 | 1.0920 | 2.1104 | 3.3065 | 4.4503 | | 1.8492 | H6 | 1.0920 | 2.1104 | 3.3065 | 4.4503 | 1.8492 | |
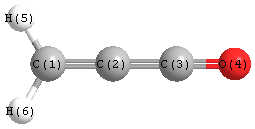 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
180.000 |
|
C2 |
C1 |
H5 |
122.143 |
| C2 |
C1 |
H6 |
122.143 |
|
C2 |
C3 |
O4 |
180.000 |
| H5 |
C1 |
H6 |
115.714 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.343 |
|
|
|
| 2 |
C |
0.236 |
|
|
|
| 3 |
C |
0.170 |
|
|
|
| 4 |
O |
-0.391 |
|
|
|
| 5 |
H |
0.164 |
|
|
|
| 6 |
H |
0.164 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-2.706 |
2.706 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-22.145 |
0.000 |
0.000 |
| y |
0.000 |
-21.943 |
0.000 |
| z |
0.000 |
0.000 |
-21.320 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.513 |
0.000 |
0.000 |
| y |
0.000 |
-0.210 |
0.000 |
| z |
0.000 |
0.000 |
0.724 |
|
| Polar |
|---|
| 3z2-r2 | 1.447 |
|---|
| x2-y2 | -0.202 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
1.889 |
0.000 |
0.000 |
| y |
0.000 |
2.749 |
0.000 |
| z |
0.000 |
0.000 |
10.144 |
<r2> (average value of r
2) Å
2
| <r2> |
82.254 |
| (<r2>)1/2 |
9.069 |
Jump to
S1C1
Energy calculated at HSEh1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -190.451984 |
| Energy at 298.15K | |
| HF Energy | -190.451984 |
| Nuclear repulsion energy | 88.426298 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3195 |
3049 |
10.81 |
121.14 |
0.70 |
0.82 |
| 2 |
A' |
3115 |
2972 |
25.97 |
164.06 |
0.11 |
0.20 |
| 3 |
A' |
2300 |
2195 |
885.76 |
34.59 |
0.36 |
0.53 |
| 4 |
A' |
1813 |
1730 |
11.78 |
1.02 |
0.66 |
0.79 |
| 5 |
A' |
1491 |
1423 |
1.62 |
18.60 |
0.55 |
0.71 |
| 6 |
A' |
1067 |
1018 |
11.08 |
4.17 |
0.16 |
0.27 |
| 7 |
A' |
964 |
920 |
4.31 |
34.22 |
0.23 |
0.38 |
| 8 |
A' |
483 |
461 |
15.67 |
3.82 |
0.60 |
0.75 |
| 9 |
A' |
128 |
122 |
19.95 |
4.89 |
0.74 |
0.85 |
| 10 |
A" |
1022 |
976 |
24.01 |
0.19 |
0.75 |
0.86 |
| 11 |
A" |
691 |
659 |
18.30 |
2.70 |
0.75 |
0.86 |
| 12 |
A" |
260 |
248 |
1.33 |
6.16 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 8264.0 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 7886.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.859 |
-1.628 |
0.000 |
| C2 |
0.000 |
-0.622 |
0.000 |
| C3 |
-0.289 |
0.640 |
0.000 |
| O4 |
-0.735 |
1.726 |
0.000 |
| H5 |
1.941 |
-1.483 |
0.000 |
| H6 |
0.521 |
-2.664 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
| C1 | | 1.3226 | 2.5411 | 3.7132 | 1.0921 | 1.0903 |
C2 | 1.3226 | | 1.2942 | 2.4602 | 2.1237 | 2.1076 | C3 | 2.5411 | 1.2942 | | 1.1745 | 3.0787 | 3.4015 | O4 | 3.7132 | 2.4602 | 1.1745 | | 4.1787 | 4.5661 | H5 | 1.0921 | 2.1237 | 3.0787 | 4.1787 | | 1.8475 | H6 | 1.0903 | 2.1076 | 3.4015 | 4.5661 | 1.8475 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
152.390 |
|
C2 |
C1 |
H5 |
122.883 |
| C2 |
C1 |
H6 |
121.433 |
|
C2 |
C3 |
O4 |
170.518 |
| H5 |
C1 |
H6 |
115.684 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.288 |
|
|
|
| 2 |
C |
0.004 |
|
|
|
| 3 |
C |
0.338 |
|
|
|
| 4 |
O |
-0.375 |
|
|
|
| 5 |
H |
0.161 |
|
|
|
| 6 |
H |
0.160 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.633 |
-1.935 |
0.000 |
2.532 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-21.519 |
-0.804 |
0.000 |
| y |
-0.804 |
-21.997 |
0.000 |
| z |
0.000 |
0.000 |
-22.227 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.593 |
-0.804 |
0.000 |
| y |
-0.804 |
-0.123 |
0.000 |
| z |
0.000 |
0.000 |
-0.469 |
|
| Polar |
|---|
| 3z2-r2 | -0.939 |
|---|
| x2-y2 | 0.478 |
|---|
| xy | -0.804 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.236 |
-2.561 |
0.000 |
| y |
-2.561 |
8.681 |
0.000 |
| z |
0.000 |
0.000 |
1.933 |
<r2> (average value of r
2) Å
2
| <r2> |
80.775 |
| (<r2>)1/2 |
8.987 |
