Jump to
S1C2
Energy calculated at HSEh1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -278.021606 |
| Energy at 298.15K | |
| HF Energy | -278.021606 |
| Nuclear repulsion energy | 125.991469 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3070 |
2930 |
0.00 |
204.16 |
0.09 |
0.16 |
| 2 |
Ag |
1533 |
1463 |
0.00 |
22.68 |
0.72 |
0.83 |
| 3 |
Ag |
1476 |
1409 |
0.00 |
3.55 |
0.70 |
0.83 |
| 4 |
Ag |
1118 |
1067 |
0.00 |
9.43 |
0.67 |
0.80 |
| 5 |
Ag |
1104 |
1054 |
0.00 |
6.17 |
0.29 |
0.45 |
| 6 |
Ag |
466 |
445 |
0.00 |
2.74 |
0.52 |
0.68 |
| 7 |
Au |
3144 |
3000 |
68.69 |
0.00 |
0.00 |
0.00 |
| 8 |
Au |
1244 |
1187 |
2.65 |
0.00 |
0.00 |
0.00 |
| 9 |
Au |
819 |
782 |
0.01 |
0.00 |
0.00 |
0.00 |
| 10 |
Au |
131 |
125 |
11.17 |
0.00 |
0.00 |
0.00 |
| 11 |
Bg |
3119 |
2976 |
0.00 |
127.76 |
0.75 |
0.86 |
| 12 |
Bg |
1307 |
1247 |
0.00 |
18.57 |
0.75 |
0.86 |
| 13 |
Bg |
1193 |
1139 |
0.00 |
5.37 |
0.75 |
0.86 |
| 14 |
Bu |
3076 |
2936 |
66.23 |
0.00 |
0.00 |
0.00 |
| 15 |
Bu |
1543 |
1473 |
2.48 |
0.00 |
0.00 |
0.00 |
| 16 |
Bu |
1371 |
1308 |
15.22 |
0.00 |
0.00 |
0.00 |
| 17 |
Bu |
1131 |
1080 |
185.70 |
0.00 |
0.00 |
0.00 |
| 18 |
Bu |
274 |
261 |
17.12 |
0.00 |
0.00 |
0.00 |
Unscaled Zero Point Vibrational Energy (zpe) 13559.5 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 12939.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G**
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.422 |
0.628 |
0.000 |
| C2 |
-0.422 |
-0.628 |
0.000 |
| F3 |
-0.422 |
1.719 |
0.000 |
| F4 |
0.422 |
-1.719 |
0.000 |
| H5 |
1.059 |
0.660 |
0.891 |
| H6 |
1.059 |
0.660 |
-0.891 |
| H7 |
-1.059 |
-0.660 |
0.891 |
| H8 |
-1.059 |
-0.660 |
-0.891 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5132 | 1.3797 | 2.3468 | 1.0956 | 1.0956 | 2.1556 | 2.1556 |
C2 | 1.5132 | | 2.3468 | 1.3797 | 2.1556 | 2.1556 | 1.0956 | 1.0956 | F3 | 1.3797 | 2.3468 | | 3.5401 | 2.0272 | 2.0272 | 2.6188 | 2.6188 | F4 | 2.3468 | 1.3797 | 3.5401 | | 2.6188 | 2.6188 | 2.0272 | 2.0272 | H5 | 1.0956 | 2.1556 | 2.0272 | 2.6188 | | 1.7816 | 2.4959 | 3.0665 | H6 | 1.0956 | 2.1556 | 2.0272 | 2.6188 | 1.7816 | | 3.0665 | 2.4959 | H7 | 2.1556 | 1.0956 | 2.6188 | 2.0272 | 2.4959 | 3.0665 | | 1.7816 | H8 | 2.1556 | 1.0956 | 2.6188 | 2.0272 | 3.0665 | 2.4959 | 1.7816 | |
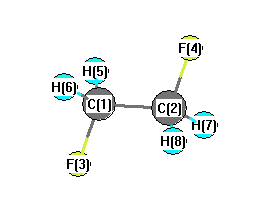 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
108.342 |
|
C1 |
C2 |
H7 |
110.411 |
| C1 |
C2 |
H8 |
110.411 |
|
C2 |
C1 |
F3 |
108.342 |
| C2 |
C1 |
H5 |
110.411 |
|
C2 |
C1 |
H6 |
110.411 |
| F3 |
C1 |
H5 |
109.432 |
|
F3 |
C1 |
H6 |
109.432 |
| F4 |
C2 |
H7 |
109.432 |
|
F4 |
C2 |
H8 |
109.432 |
| H5 |
C1 |
H6 |
108.797 |
|
H7 |
C2 |
H8 |
108.797 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.054 |
|
|
|
| 2 |
C |
0.054 |
|
|
|
| 3 |
F |
-0.315 |
|
|
|
| 4 |
F |
-0.315 |
|
|
|
| 5 |
H |
0.131 |
|
|
|
| 6 |
H |
0.131 |
|
|
|
| 7 |
H |
0.131 |
|
|
|
| 8 |
H |
0.131 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-21.554 |
2.499 |
0.000 |
| y |
2.499 |
-27.949 |
0.000 |
| z |
0.000 |
0.000 |
-21.265 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.053 |
2.499 |
0.000 |
| y |
2.499 |
-6.540 |
0.000 |
| z |
0.000 |
0.000 |
3.487 |
|
| Polar |
|---|
| 3z2-r2 | 6.973 |
|---|
| x2-y2 | 6.395 |
|---|
| xy | 2.499 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.556 |
0.036 |
0.000 |
| y |
0.036 |
3.486 |
0.000 |
| z |
0.000 |
0.000 |
3.425 |
<r2> (average value of r
2) Å
2
| <r2> |
87.402 |
| (<r2>)1/2 |
9.349 |
Jump to
S1C1
Energy calculated at HSEh1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -278.022343 |
| Energy at 298.15K | |
| HF Energy | -278.022343 |
| Nuclear repulsion energy | 128.153776 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3112 |
2970 |
21.60 |
101.05 |
0.74 |
0.85 |
| 2 |
A |
3064 |
2924 |
38.77 |
175.06 |
0.03 |
0.05 |
| 3 |
A |
1512 |
1443 |
0.61 |
2.31 |
0.62 |
0.77 |
| 4 |
A |
1462 |
1395 |
19.99 |
4.79 |
0.68 |
0.81 |
| 5 |
A |
1316 |
1256 |
1.74 |
20.91 |
0.72 |
0.84 |
| 6 |
A |
1161 |
1108 |
70.21 |
3.18 |
0.52 |
0.69 |
| 7 |
A |
1144 |
1092 |
14.97 |
2.53 |
0.59 |
0.74 |
| 8 |
A |
897 |
856 |
20.14 |
5.75 |
0.28 |
0.44 |
| 9 |
A |
325 |
311 |
0.39 |
0.24 |
0.34 |
0.50 |
| 10 |
A |
149 |
142 |
2.98 |
0.04 |
0.75 |
0.86 |
| 11 |
B |
3125 |
2982 |
51.56 |
13.71 |
0.75 |
0.86 |
| 12 |
B |
3053 |
2914 |
15.38 |
38.88 |
0.75 |
0.86 |
| 13 |
B |
1509 |
1440 |
7.73 |
15.85 |
0.75 |
0.86 |
| 14 |
B |
1423 |
1358 |
12.05 |
1.83 |
0.75 |
0.86 |
| 15 |
B |
1273 |
1215 |
4.87 |
5.91 |
0.75 |
0.86 |
| 16 |
B |
1131 |
1079 |
45.48 |
3.48 |
0.75 |
0.86 |
| 17 |
B |
924 |
882 |
31.66 |
3.68 |
0.75 |
0.86 |
| 18 |
B |
497 |
475 |
14.57 |
0.51 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 13538.6 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 12919.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G**
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.262 |
0.704 |
0.511 |
| C2 |
-0.262 |
-0.704 |
0.511 |
| F3 |
-0.262 |
1.413 |
-0.549 |
| F4 |
0.262 |
-1.413 |
-0.549 |
| H5 |
-0.020 |
1.201 |
1.448 |
| H6 |
1.355 |
0.697 |
0.427 |
| H7 |
0.020 |
-1.201 |
1.448 |
| H8 |
-1.355 |
-0.697 |
0.427 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5021 | 1.3780 | 2.3672 | 1.0977 | 1.0962 | 2.1365 | 2.1411 |
C2 | 1.5021 | | 2.3672 | 1.3780 | 2.1365 | 2.1411 | 1.0977 | 1.0962 | F3 | 1.3780 | 2.3672 | | 2.8740 | 2.0224 | 2.0192 | 3.3011 | 2.5691 | F4 | 2.3672 | 1.3780 | 2.8740 | | 3.3011 | 2.5691 | 2.0224 | 2.0192 | H5 | 1.0977 | 2.1365 | 2.0224 | 3.3011 | | 1.7852 | 2.4016 | 2.5350 | H6 | 1.0962 | 2.1411 | 2.0192 | 2.5691 | 1.7852 | | 2.5350 | 3.0476 | H7 | 2.1365 | 1.0977 | 3.3011 | 2.0224 | 2.4016 | 2.5350 | | 1.7852 | H8 | 2.1411 | 1.0962 | 2.5691 | 2.0192 | 2.5350 | 3.0476 | 1.7852 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
110.476 |
|
C1 |
C2 |
H7 |
109.540 |
| C1 |
C2 |
H8 |
109.998 |
|
C2 |
C1 |
F3 |
110.476 |
| C2 |
C1 |
H5 |
109.540 |
|
C2 |
C1 |
H6 |
109.998 |
| F3 |
C1 |
H5 |
109.024 |
|
F3 |
C1 |
H6 |
108.856 |
| F4 |
C2 |
H7 |
109.024 |
|
F4 |
C2 |
H8 |
108.856 |
| H5 |
C1 |
H6 |
108.917 |
|
H7 |
C2 |
H8 |
108.917 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.064 |
|
|
|
| 2 |
C |
0.064 |
|
|
|
| 3 |
F |
-0.312 |
|
|
|
| 4 |
F |
-0.312 |
|
|
|
| 5 |
H |
0.115 |
|
|
|
| 6 |
H |
0.133 |
|
|
|
| 7 |
H |
0.115 |
|
|
|
| 8 |
H |
0.133 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
2.441 |
2.441 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-21.339 |
1.386 |
0.000 |
| y |
1.386 |
-25.327 |
0.000 |
| z |
0.000 |
0.000 |
-21.819 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.234 |
1.386 |
0.000 |
| y |
1.386 |
-3.748 |
0.000 |
| z |
0.000 |
0.000 |
1.514 |
|
| Polar |
|---|
| 3z2-r2 | 3.027 |
|---|
| x2-y2 | 3.988 |
|---|
| xy | 1.386 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.483 |
0.040 |
0.000 |
| y |
0.040 |
3.592 |
0.000 |
| z |
0.000 |
0.000 |
3.526 |
<r2> (average value of r
2) Å
2
| <r2> |
78.830 |
| (<r2>)1/2 |
8.879 |
