return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C8H6 (Calicene)

using model chemistry: MP2=FULL/6-31G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2=FULL/6-31G**
 hartrees
Energy at 0K-307.381224
Energy at 298.15K-307.385735
HF Energy-306.300678
Nuclear repulsion energy300.087956
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3381 3159 10.55      
2 A1 3315 3098 15.02      
3 A1 3292 3076 4.97      
4 A1 1872 1749 557.17      
5 A1 1594 1489 67.17      
6 A1 1533 1432 39.84      
7 A1 1452 1357 92.32      
8 A1 1245 1163 29.83      
9 A1 1110 1037 11.77      
10 A1 1059 989 11.45      
11 A1 979 915 6.82      
12 A1 900 841 41.12      
13 A1 493 461 0.34      
14 A2 882 824 0.00      
15 A2 825 771 0.00      
16 A2 663 619 0.00      
17 A2 545 510 0.00      
18 A2 212 198 0.00      
19 B1 845 790 1.19      
20 B1 718 671 108.85      
21 B1 656 613 16.23      
22 B1 554 518 1.31      
23 B1 381 356 8.45      
24 B1 110 102 4.18      
25 B2 3343 3123 6.25      
26 B2 3307 3090 23.57      
27 B2 3284 3068 1.16      
28 B2 1604 1499 0.75      
29 B2 1389 1298 0.00      
30 B2 1266 1183 0.56      
31 B2 1147 1071 11.28      
32 B2 1122 1048 7.70      
33 B2 965 902 6.69      
34 B2 839 784 1.52      
35 B2 501 468 1.31      
36 B2 146 136 1.46      

Unscaled Zero Point Vibrational Energy (zpe) 23763.8 cm-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 22204.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31G**
ABC
0.22636 0.04763 0.03935

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31G**

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 -0.049
C2 0.000 0.000 1.302
C3 0.000 0.667 2.566
C4 0.000 -0.667 2.566
C5 0.000 1.170 -0.907
C6 0.000 -1.170 -0.907
C7 0.000 0.725 -2.201
C8 0.000 -0.725 -2.201
H9 0.000 1.581 3.132
H10 0.000 -1.581 3.132
H11 0.000 2.192 -0.561
H12 0.000 -2.192 -0.561
H13 0.000 1.344 -3.085
H14 0.000 -1.344 -3.085

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 C8 H9 H10 H11 H12 H13 H14
C11.35012.69862.69861.45061.45062.27062.27063.55183.55182.25132.25133.32033.3203
C21.35011.42971.42972.49872.49873.57623.57622.41872.41872.87672.87674.58764.5876
C32.69861.42971.33343.50903.92854.76724.96571.07522.31793.47974.23735.69155.9981
C42.69861.42971.33343.92853.50904.96574.76722.31791.07524.23733.47975.99815.6915
C51.45062.49873.50903.92852.33951.36852.29414.05924.88631.07923.37962.18523.3262
C61.45062.49873.92853.50902.33952.29411.36854.88634.05923.37961.07923.32622.1852
C72.27063.57624.76724.96571.36852.29411.44905.40075.80952.20033.34581.07962.2496
C82.27063.57624.96574.76722.29411.36851.44905.80955.40073.34582.20032.24961.0796
H93.55182.41871.07522.31794.05924.88635.40075.80953.16233.74325.27986.22116.8704
H103.55182.41872.31791.07524.88634.05925.80955.40073.16235.27983.74326.87046.2211
H112.25132.87673.47974.23731.07923.37962.20033.34583.74325.27984.38432.66224.3443
H122.25132.87674.23733.47973.37961.07923.34582.20035.27983.74324.38434.34432.6622
H133.32034.58765.69155.99812.18523.32621.07962.24966.22116.87042.66224.34432.6881
H143.32034.58765.99815.69153.32622.18522.24961.07966.87046.22114.34432.66222.6881

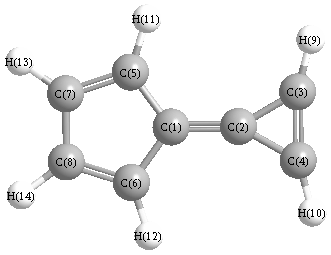
picture of Calicene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 152.205 C1 C2 C4 152.205
C1 C5 C7 107.270 C1 C5 H11 125.083
C1 C6 C8 107.270 C1 C6 H12 125.083
C2 C1 C5 126.257 C2 C1 C6 126.257
C2 C3 C4 62.205 C2 C3 H9 149.529
C2 C4 C3 62.205 C2 C4 H10 149.529
C3 C2 C4 55.591 C3 C4 H10 148.266
C4 C3 H9 148.266 C5 C1 C6 107.487
C5 C7 C8 108.987 C5 C7 H13 125.997
C6 C8 C7 108.987 C6 C8 H14 125.997
C7 C5 H11 127.647 C7 C8 H14 125.017
C8 C6 H12 127.647 C8 C7 H13 125.017
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability