return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H10 (2-Pentene, (Z)-)

using model chemistry: MP2=FULL/6-31G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2=FULL/6-31G**
 hartrees
Energy at 0K-195.895759
Energy at 298.15K-195.905826
HF Energy-195.157613
Nuclear repulsion energy171.915754
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3250 3037 49.95      
2 A 3237 3025 9.79      
3 A 3231 3019 35.56      
4 A 3225 3013 30.05      
5 A 3216 3005 3.91      
6 A 3200 2990 19.16      
7 A 3192 2983 7.38      
8 A 3128 2923 29.61      
9 A 3127 2922 18.07      
10 A 3124 2919 26.26      
11 A 1759 1644 1.47      
12 A 1568 1465 4.91      
13 A 1561 1459 5.13      
14 A 1558 1456 2.64      
15 A 1545 1443 4.21      
16 A 1541 1440 0.73      
17 A 1484 1386 4.93      
18 A 1462 1366 1.07      
19 A 1453 1357 1.99      
20 A 1378 1287 6.15      
21 A 1337 1249 0.06      
22 A 1315 1228 0.08      
23 A 1202 1123 0.50      
24 A 1128 1054 3.64      
25 A 1093 1021 0.97      
26 A 1081 1010 0.03      
27 A 1073 1002 1.88      
28 A 1008 942 0.23      
29 A 971 907 7.42      
30 A 892 834 1.03      
31 A 811 758 2.42      
32 A 740 691 28.96      
33 A 570 533 5.25      
34 A 477 446 1.07      
35 A 317 296 0.05      
36 A 259 242 0.01      
37 A 233 217 0.01      
38 A 115 107 0.28      
39 A 46 43 0.07      

Unscaled Zero Point Vibrational Energy (zpe) 30951.3 cm-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 28920.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31G**
ABC
0.35730 0.08823 0.08041

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31G**

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 2.129 -0.339 -0.425
H2 2.610 0.602 -0.685
H3 2.895 -1.013 -0.044
H4 1.720 -0.767 -1.338
C5 1.022 -0.118 0.604
H6 0.566 -1.072 0.862
H7 1.463 0.272 1.525
C8 -2.022 -0.699 -0.051
H9 -2.449 -0.994 -1.009
H10 -1.378 -1.504 0.289
H11 -2.850 -0.610 0.652
C12 -0.008 0.851 0.109
H13 0.354 1.862 -0.054
C14 -1.295 0.604 -0.179
H15 -1.891 1.432 -0.545

Atom - Atom Distances (Å)
  C1 H2 H3 H4 C5 H6 H7 C8 H9 H10 H11 C12 H13 C14 H15
C11.08881.08861.08811.52842.15342.14954.18394.66203.76475.10202.50442.85233.56004.3948
H21.08881.76151.75892.16883.06182.51194.85305.31504.61475.75092.74712.65993.93724.5787
H31.08861.76151.76482.17492.49912.48244.92685.43054.31415.80083.45333.83704.49245.3973
H41.08811.75891.76482.16412.50303.05653.95814.18873.57684.98752.77503.22923.50884.3018
C51.52842.16882.17492.16411.08861.09243.16753.92692.78993.90341.49812.19032.54943.4937
H62.15343.06182.49912.50301.08861.74632.76993.54982.07283.45382.14383.08082.71183.7799
H72.14952.51192.48243.05651.09241.74633.94594.83003.57134.48792.12212.50013.25834.1083
C84.18394.85304.92683.95813.16752.76993.94591.08991.08591.09012.54653.49291.49732.1914
H94.66205.31505.43054.18873.92693.54984.83001.08991.75901.75193.25804.11372.13882.5322
H103.76474.61474.31413.57682.78992.07283.57131.08591.75901.75992.73123.80112.16103.0954
H115.10205.75095.80084.98753.90343.45384.48791.09011.75191.75993.24174.10772.14122.5546
C122.50442.74713.45332.77501.49812.14382.12212.54653.25802.73123.24171.08551.34142.0761
H132.85232.65993.83703.22922.19033.08082.50013.49294.11373.80114.10771.08552.07722.3373
C143.56003.93724.49243.50882.54942.71183.25831.49732.13882.16102.14121.34142.07721.0844
H154.39484.57875.39734.30183.49373.77994.10832.19142.53223.09542.55462.07612.33731.0844

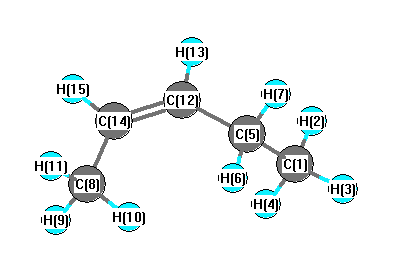
picture of 2-Pentene, (Z)- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C5 H6 109.596 C1 C5 H7 109.067
C1 C5 C12 111.671 H2 C1 H3 107.996
H2 C1 H4 107.800 H2 C1 C5 110.806
H3 C1 H4 108.346 H3 C1 C5 111.301
H4 C1 C5 110.470 C5 C12 H13 114.998
C5 C12 C14 127.655 H6 C5 H7 106.396
H6 C5 C12 110.952 H7 C5 C12 109.000
C8 C14 C12 127.460 C8 C14 H15 115.229
H9 C8 H10 107.886 H9 C8 H11 106.958
H9 C8 C14 110.527 H10 C8 H11 107.957
H10 C8 C14 112.585 H11 C8 C14 110.710
C12 C14 H15 117.311 H13 C12 C14 117.327
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability