Jump to
S1C2
Energy calculated at MP2=FULL/6-31G**
| | hartrees |
|---|
| Energy at 0K | -553.126423 |
| Energy at 298.15K | -553.131753 |
| HF Energy | -552.428470 |
| Nuclear repulsion energy | 199.249035 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3353 |
3133 |
6.11 |
|
|
|
| 2 |
A |
3268 |
3054 |
5.85 |
|
|
|
| 3 |
A |
3246 |
3033 |
0.72 |
|
|
|
| 4 |
A |
1694 |
1583 |
4.54 |
|
|
|
| 5 |
A |
1466 |
1370 |
8.31 |
|
|
|
| 6 |
A |
1339 |
1252 |
1.56 |
|
|
|
| 7 |
A |
1087 |
1016 |
0.21 |
|
|
|
| 8 |
A |
1024 |
956 |
10.81 |
|
|
|
| 9 |
A |
929 |
868 |
19.66 |
|
|
|
| 10 |
A |
767 |
717 |
0.66 |
|
|
|
| 11 |
A |
646 |
603 |
12.39 |
|
|
|
| 12 |
A |
385 |
359 |
0.31 |
|
|
|
| 13 |
A |
205 |
191 |
0.00 |
|
|
|
| 14 |
A |
82 |
77 |
0.17 |
|
|
|
| 15 |
A |
3353 |
3133 |
4.98 |
|
|
|
| 16 |
A |
3265 |
3051 |
12.53 |
|
|
|
| 17 |
A |
3245 |
3032 |
1.21 |
|
|
|
| 18 |
A |
1680 |
1570 |
42.30 |
|
|
|
| 19 |
A |
1458 |
1363 |
6.26 |
|
|
|
| 20 |
A |
1330 |
1242 |
17.16 |
|
|
|
| 21 |
A |
1071 |
1001 |
6.22 |
|
|
|
| 22 |
A |
1015 |
949 |
66.55 |
|
|
|
| 23 |
A |
928 |
867 |
34.57 |
|
|
|
| 24 |
A |
760 |
710 |
9.96 |
|
|
|
| 25 |
A |
619 |
578 |
12.95 |
|
|
|
| 26 |
A |
409 |
382 |
0.91 |
|
|
|
| 27 |
A |
71 |
66 |
3.26 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 19347.4 cm
-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 18078.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31G**
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.000 |
0.000 |
0.709 |
| C2 |
0.000 |
1.346 |
-0.430 |
| C3 |
0.000 |
-1.346 |
-0.430 |
| C4 |
0.731 |
2.451 |
-0.248 |
| C5 |
-0.731 |
-2.451 |
-0.248 |
| H6 |
-0.690 |
1.262 |
-1.260 |
| H7 |
0.690 |
-1.262 |
-1.260 |
| H8 |
0.635 |
3.288 |
-0.922 |
| H9 |
1.422 |
2.543 |
0.576 |
| H10 |
-0.635 |
-3.288 |
-0.922 |
| H11 |
-1.422 |
-2.543 |
0.576 |
Atom - Atom Distances (Å)
| |
S1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| S1 | | 1.7631 | 1.7631 | 2.7305 | 2.7305 | 2.4381 | 2.4381 | 3.7252 | 2.9171 | 3.7252 | 2.9171 |
C2 | 1.7631 | | 2.6911 | 1.3373 | 3.8701 | 1.0823 | 2.8216 | 2.1022 | 2.1144 | 4.7028 | 4.2615 | C3 | 1.7631 | 2.6911 | | 3.8701 | 1.3373 | 2.8216 | 1.0823 | 4.7028 | 4.2615 | 2.1022 | 2.1144 | C4 | 2.7305 | 1.3373 | 3.8701 | | 5.1145 | 2.1109 | 3.8478 | 1.0795 | 1.0798 | 5.9374 | 5.5005 | C5 | 2.7305 | 3.8701 | 1.3373 | 5.1145 | | 3.8478 | 2.1109 | 5.9374 | 5.5005 | 1.0795 | 1.0798 | H6 | 2.4381 | 1.0823 | 2.8216 | 2.1109 | 3.8478 | | 2.8757 | 2.4446 | 3.0782 | 4.5625 | 4.2879 | H7 | 2.4381 | 2.8216 | 1.0823 | 3.8478 | 2.1109 | 2.8757 | | 4.5625 | 4.2879 | 2.4446 | 3.0782 | H8 | 3.7252 | 2.1022 | 4.7028 | 1.0795 | 5.9374 | 2.4446 | 4.5625 | | 1.8493 | 6.6978 | 6.3629 | H9 | 2.9171 | 2.1144 | 4.2615 | 1.0798 | 5.5005 | 3.0782 | 4.2879 | 1.8493 | | 6.3629 | 5.8282 | H10 | 3.7252 | 4.7028 | 2.1022 | 5.9374 | 1.0795 | 4.5625 | 2.4446 | 6.6978 | 6.3629 | | 1.8493 | H11 | 2.9171 | 4.2615 | 2.1144 | 5.5005 | 1.0798 | 4.2879 | 3.0782 | 6.3629 | 5.8282 | 1.8493 | |
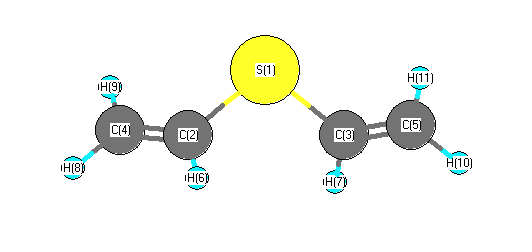 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
C2 |
C4 |
122.863 |
|
S1 |
C2 |
H6 |
115.861 |
| S1 |
C3 |
C5 |
122.863 |
|
S1 |
C3 |
H7 |
115.861 |
| C2 |
S1 |
C3 |
99.493 |
|
C2 |
C4 |
H8 |
120.499 |
| C2 |
C4 |
H9 |
121.665 |
|
C3 |
C5 |
H10 |
120.499 |
| C3 |
C5 |
H11 |
121.665 |
|
C4 |
C2 |
H6 |
121.120 |
| C5 |
C3 |
H7 |
121.120 |
|
H8 |
C4 |
H9 |
117.831 |
| H10 |
C5 |
H11 |
117.831 |
|
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/6-31G**
| | hartrees |
|---|
| Energy at 0K | -553.128010 |
| Energy at 298.15K | -553.133567 |
| HF Energy | -552.428244 |
| Nuclear repulsion energy | 202.799557 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3358 |
3137 |
5.54 |
|
|
|
| 2 |
A |
3354 |
3134 |
4.68 |
|
|
|
| 3 |
A |
3274 |
3060 |
6.51 |
|
|
|
| 4 |
A |
3262 |
3048 |
6.45 |
|
|
|
| 5 |
A |
3252 |
3038 |
0.76 |
|
|
|
| 6 |
A |
3245 |
3032 |
1.27 |
|
|
|
| 7 |
A |
1690 |
1579 |
21.24 |
|
|
|
| 8 |
A |
1680 |
1570 |
18.44 |
|
|
|
| 9 |
A |
1467 |
1371 |
3.45 |
|
|
|
| 10 |
A |
1463 |
1367 |
7.01 |
|
|
|
| 11 |
A |
1339 |
1251 |
2.57 |
|
|
|
| 12 |
A |
1332 |
1244 |
9.54 |
|
|
|
| 13 |
A |
1091 |
1019 |
6.27 |
|
|
|
| 14 |
A |
1064 |
994 |
14.54 |
|
|
|
| 15 |
A |
1028 |
961 |
22.33 |
|
|
|
| 16 |
A |
1013 |
946 |
36.17 |
|
|
|
| 17 |
A |
935 |
874 |
27.04 |
|
|
|
| 18 |
A |
895 |
836 |
33.12 |
|
|
|
| 19 |
A |
773 |
722 |
7.56 |
|
|
|
| 20 |
A |
726 |
678 |
4.11 |
|
|
|
| 21 |
A |
649 |
607 |
13.36 |
|
|
|
| 22 |
A |
619 |
579 |
10.84 |
|
|
|
| 23 |
A |
468 |
437 |
0.39 |
|
|
|
| 24 |
A |
379 |
354 |
0.59 |
|
|
|
| 25 |
A |
237 |
221 |
1.41 |
|
|
|
| 26 |
A |
148 |
138 |
4.07 |
|
|
|
| 27 |
A |
93 |
87 |
1.48 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 19415.4 cm
-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 18141.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31G**
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.005 |
-0.999 |
-0.006 |
| C2 |
1.065 |
0.345 |
0.396 |
| C3 |
-1.550 |
-0.181 |
-0.026 |
| C4 |
2.228 |
0.570 |
-0.226 |
| C5 |
-1.768 |
1.136 |
-0.127 |
| H6 |
0.751 |
0.945 |
1.241 |
| H7 |
-2.379 |
-0.871 |
0.065 |
| H8 |
2.894 |
1.344 |
0.122 |
| H9 |
2.538 |
-0.017 |
-1.078 |
| H10 |
-0.964 |
1.845 |
-0.247 |
| H11 |
-2.776 |
1.519 |
-0.099 |
Atom - Atom Distances (Å)
| |
S1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| S1 | | 1.7578 | 1.7579 | 2.7287 | 2.7776 | 2.4265 | 2.3891 | 3.7216 | 2.9197 | 3.0145 | 3.7528 |
C2 | 1.7578 | | 2.7010 | 1.3373 | 2.9871 | 1.0829 | 3.6673 | 2.1020 | 2.1143 | 2.6038 | 4.0468 | C3 | 1.7579 | 2.7010 | | 3.8570 | 1.3385 | 2.8579 | 1.0819 | 4.7013 | 4.2245 | 2.1213 | 2.0968 | C4 | 2.7287 | 1.3373 | 3.8570 | | 4.0362 | 2.1148 | 4.8353 | 1.0797 | 1.0795 | 3.4367 | 5.0942 | C5 | 2.7776 | 2.9871 | 1.3385 | 4.0362 | | 2.8723 | 2.1065 | 4.6730 | 4.5570 | 1.0790 | 1.0790 | H6 | 2.4265 | 1.0829 | 2.8579 | 2.1148 | 2.8723 | | 3.8045 | 2.4502 | 3.0809 | 2.4419 | 3.8162 | H7 | 2.3891 | 3.6673 | 1.0819 | 4.8353 | 2.1065 | 3.8045 | | 5.7197 | 5.1196 | 3.0787 | 2.4278 | H8 | 3.7216 | 2.1020 | 4.7013 | 1.0797 | 4.6730 | 2.4502 | 5.7197 | | 1.8492 | 3.9076 | 5.6771 | H9 | 2.9197 | 2.1143 | 4.2245 | 1.0795 | 4.5570 | 3.0809 | 5.1196 | 1.8492 | | 4.0518 | 5.6168 | H10 | 3.0145 | 2.6038 | 2.1213 | 3.4367 | 1.0790 | 2.4419 | 3.0787 | 3.9076 | 4.0518 | | 1.8474 | H11 | 3.7528 | 4.0468 | 2.0968 | 5.0942 | 1.0790 | 3.8162 | 2.4278 | 5.6771 | 5.6168 | 1.8474 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
C2 |
C4 |
123.102 |
|
S1 |
C2 |
H6 |
115.277 |
| S1 |
C3 |
C5 |
127.019 |
|
S1 |
C3 |
H7 |
112.360 |
| C2 |
S1 |
C3 |
100.397 |
|
C2 |
C4 |
H8 |
120.472 |
| C2 |
C4 |
H9 |
121.689 |
|
C3 |
C5 |
H10 |
122.317 |
| C3 |
C5 |
H11 |
119.918 |
|
C4 |
C2 |
H6 |
121.463 |
| C5 |
C3 |
H7 |
120.619 |
|
H8 |
C4 |
H9 |
117.835 |
| H10 |
C5 |
H11 |
117.763 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
