Jump to
S1C2
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
Geometric Data calculated at HSEh1PBE/6-31G**
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at HSEh1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -553.856874 |
| Energy at 298.15K | -553.862362 |
| HF Energy | -553.856874 |
| Nuclear repulsion energy | 202.144113 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3281 |
3131 |
6.39 |
|
|
|
| 2 |
A |
3280 |
3130 |
3.54 |
|
|
|
| 3 |
A |
3212 |
3065 |
3.91 |
|
|
|
| 4 |
A |
3197 |
3051 |
4.59 |
|
|
|
| 5 |
A |
3186 |
3040 |
1.04 |
|
|
|
| 6 |
A |
3182 |
3036 |
1.43 |
|
|
|
| 7 |
A |
1692 |
1615 |
38.94 |
|
|
|
| 8 |
A |
1682 |
1605 |
28.67 |
|
|
|
| 9 |
A |
1428 |
1363 |
3.47 |
|
|
|
| 10 |
A |
1425 |
1360 |
6.44 |
|
|
|
| 11 |
A |
1309 |
1249 |
2.08 |
|
|
|
| 12 |
A |
1295 |
1236 |
8.64 |
|
|
|
| 13 |
A |
1065 |
1016 |
10.57 |
|
|
|
| 14 |
A |
1035 |
988 |
13.90 |
|
|
|
| 15 |
A |
1009 |
963 |
24.77 |
|
|
|
| 16 |
A |
992 |
947 |
34.42 |
|
|
|
| 17 |
A |
934 |
891 |
34.48 |
|
|
|
| 18 |
A |
898 |
857 |
41.85 |
|
|
|
| 19 |
A |
758 |
723 |
12.08 |
|
|
|
| 20 |
A |
710 |
678 |
4.33 |
|
|
|
| 21 |
A |
634 |
605 |
13.97 |
|
|
|
| 22 |
A |
613 |
585 |
12.67 |
|
|
|
| 23 |
A |
468 |
446 |
0.63 |
|
|
|
| 24 |
A |
368 |
351 |
0.60 |
|
|
|
| 25 |
A |
224 |
213 |
1.18 |
|
|
|
| 26 |
A |
152 |
145 |
4.76 |
|
|
|
| 27 |
A |
91 |
87 |
1.34 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 19059.8 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 18188.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G**
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.021 |
-0.967 |
0.004 |
| C2 |
1.077 |
0.398 |
0.342 |
| C3 |
-1.566 |
-0.210 |
-0.015 |
| C4 |
2.284 |
0.538 |
-0.202 |
| C5 |
-1.848 |
1.088 |
-0.134 |
| H6 |
0.705 |
1.084 |
1.101 |
| H7 |
-2.358 |
-0.947 |
0.095 |
| H8 |
2.944 |
1.336 |
0.124 |
| H9 |
2.654 |
-0.138 |
-0.967 |
| H10 |
-1.080 |
1.841 |
-0.275 |
| H11 |
-2.879 |
1.424 |
-0.099 |
Atom - Atom Distances (Å)
| |
S1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| S1 | | 1.7586 | 1.7584 | 2.7261 | 2.7811 | 2.4246 | 2.3801 | 3.7236 | 2.9261 | 3.0298 | 3.7599 |
C2 | 1.7586 | | 2.7349 | 1.3320 | 3.0423 | 1.0886 | 3.6964 | 2.1013 | 2.1184 | 2.6679 | 4.1101 | C3 | 1.7584 | 2.7349 | | 3.9266 | 1.3332 | 2.8416 | 1.0873 | 4.7696 | 4.3263 | 2.1240 | 2.0976 | C4 | 2.7261 | 1.3320 | 3.9266 | | 4.1693 | 2.1192 | 4.8826 | 1.0855 | 1.0855 | 3.6091 | 5.2397 | C5 | 2.7811 | 3.0423 | 1.3332 | 4.1693 | | 2.8358 | 2.1101 | 4.8054 | 4.7394 | 1.0852 | 1.0849 | H6 | 2.4246 | 1.0886 | 2.8416 | 2.1192 | 2.8358 | | 3.8100 | 2.4564 | 3.0932 | 2.3782 | 3.7945 | H7 | 2.3801 | 3.6964 | 1.0873 | 4.8826 | 2.1101 | 3.8100 | | 5.7724 | 5.1858 | 3.0895 | 2.4354 | H8 | 3.7236 | 2.1013 | 4.7696 | 1.0855 | 4.8054 | 2.4564 | 5.7724 | | 1.8563 | 4.0756 | 5.8279 | H9 | 2.9261 | 2.1184 | 4.3263 | 1.0855 | 4.7394 | 3.0932 | 5.1858 | 1.8563 | | 4.2825 | 5.8140 | H10 | 3.0298 | 2.6679 | 2.1240 | 3.6091 | 1.0852 | 2.3782 | 3.0895 | 4.0756 | 4.2825 | | 1.8549 | H11 | 3.7599 | 4.1101 | 2.0976 | 5.2397 | 1.0849 | 3.7945 | 2.4354 | 5.8279 | 5.8140 | 1.8549 | |
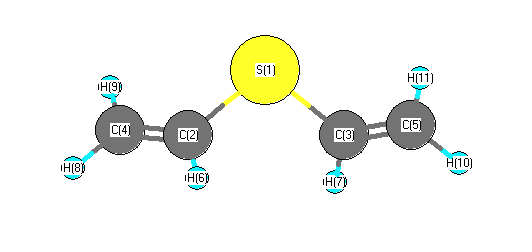 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
C2 |
C4 |
123.194 |
|
S1 |
C2 |
H6 |
114.719 |
| S1 |
C3 |
C5 |
127.674 |
|
S1 |
C3 |
H7 |
111.327 |
| C2 |
S1 |
C3 |
102.087 |
|
C2 |
C4 |
H8 |
120.399 |
| C2 |
C4 |
H9 |
122.061 |
|
C3 |
C5 |
H10 |
122.549 |
| C3 |
C5 |
H11 |
119.981 |
|
C4 |
C2 |
H6 |
121.886 |
| C5 |
C3 |
H7 |
120.997 |
|
H8 |
C4 |
H9 |
117.534 |
| H10 |
C5 |
H11 |
117.468 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
0.216 |
|
|
|
| 2 |
C |
-0.270 |
|
|
|
| 3 |
C |
-0.274 |
|
|
|
| 4 |
C |
-0.271 |
|
|
|
| 5 |
C |
-0.285 |
|
|
|
| 6 |
H |
0.162 |
|
|
|
| 7 |
H |
0.161 |
|
|
|
| 8 |
H |
0.141 |
|
|
|
| 9 |
H |
0.146 |
|
|
|
| 10 |
H |
0.136 |
|
|
|
| 11 |
H |
0.138 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.084 |
1.048 |
0.211 |
1.073 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-32.230 |
1.578 |
-0.774 |
| y |
1.578 |
-36.806 |
1.151 |
| z |
-0.774 |
1.151 |
-40.030 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
6.188 |
1.578 |
-0.774 |
| y |
1.578 |
-0.676 |
1.151 |
| z |
-0.774 |
1.151 |
-5.512 |
|
| Polar |
|---|
| 3z2-r2 | -11.024 |
|---|
| x2-y2 | 4.576 |
|---|
| xy | 1.578 |
|---|
| xz | -0.774 |
|---|
| yz | 1.151 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
12.813 |
-0.006 |
-0.770 |
| y |
-0.006 |
8.926 |
0.239 |
| z |
-0.770 |
0.239 |
4.839 |
<r2> (average value of r
2) Å
2
| <r2> |
165.027 |
| (<r2>)1/2 |
12.846 |
