Jump to
S1C2
Energy calculated at HSEh1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -189.575155 |
| Energy at 298.15K | -189.577892 |
| HF Energy | -189.575155 |
| Nuclear repulsion energy | 70.312618 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3792 |
3619 |
52.97 |
73.57 |
0.28 |
0.44 |
| 2 |
A' |
3092 |
2950 |
51.71 |
102.91 |
0.28 |
0.44 |
| 3 |
A' |
1885 |
1799 |
327.43 |
5.87 |
0.22 |
0.37 |
| 4 |
A' |
1420 |
1355 |
5.08 |
9.26 |
0.64 |
0.78 |
| 5 |
A' |
1335 |
1274 |
18.61 |
0.91 |
0.33 |
0.49 |
| 6 |
A' |
1166 |
1113 |
232.83 |
1.98 |
0.22 |
0.35 |
| 7 |
A' |
633 |
604 |
53.73 |
3.98 |
0.60 |
0.75 |
| 8 |
A" |
1067 |
1018 |
0.88 |
2.35 |
0.75 |
0.86 |
| 9 |
A" |
713 |
680 |
156.67 |
1.96 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 7551.5 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 7206.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.420 |
0.000 |
| O2 |
-1.031 |
-0.434 |
0.000 |
| O3 |
1.159 |
0.103 |
0.000 |
| H4 |
-0.377 |
1.453 |
0.000 |
| H5 |
-0.649 |
-1.326 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
O3 |
H4 |
H5 |
| C1 | | 1.3384 | 1.2014 | 1.1000 | 1.8621 |
O2 | 1.3384 | | 2.2548 | 1.9968 | 0.9702 | O3 | 1.2014 | 2.2548 | | 2.0450 | 2.3045 | H4 | 1.1000 | 1.9968 | 2.0450 | | 2.7919 | H5 | 1.8621 | 0.9702 | 2.3045 | 2.7919 | |
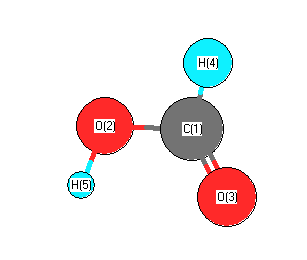 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H5 |
106.439 |
|
O2 |
C1 |
O3 |
125.106 |
| O2 |
C1 |
H4 |
109.563 |
|
O3 |
C1 |
H4 |
125.331 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.411 |
|
|
|
| 2 |
O |
-0.455 |
|
|
|
| 3 |
O |
-0.422 |
|
|
|
| 4 |
H |
0.130 |
|
|
|
| 5 |
H |
0.336 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.409 |
-0.416 |
0.000 |
1.469 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-21.512 |
-0.026 |
0.000 |
| y |
-0.026 |
-12.492 |
0.000 |
| z |
0.000 |
0.000 |
-16.570 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.981 |
-0.026 |
0.000 |
| y |
-0.026 |
6.549 |
0.000 |
| z |
0.000 |
0.000 |
0.432 |
|
| Polar |
|---|
| 3z2-r2 | 0.864 |
|---|
| x2-y2 | -9.019 |
|---|
| xy | -0.026 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.087 |
0.051 |
0.000 |
| y |
0.051 |
2.824 |
0.000 |
| z |
0.000 |
0.000 |
1.339 |
<r2> (average value of r
2) Å
2
| <r2> |
36.857 |
| (<r2>)1/2 |
6.071 |
Jump to
S1C1
Energy calculated at HSEh1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -189.566820 |
| Energy at 298.15K | |
| HF Energy | -189.566820 |
| Nuclear repulsion energy | 70.085121 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3857 |
3681 |
52.25 |
93.08 |
0.28 |
0.44 |
| 2 |
A' |
2988 |
2851 |
95.33 |
88.56 |
0.28 |
0.44 |
| 3 |
A' |
1932 |
1844 |
273.71 |
8.19 |
0.21 |
0.34 |
| 4 |
A' |
1448 |
1382 |
2.01 |
7.97 |
0.63 |
0.78 |
| 5 |
A' |
1302 |
1242 |
349.09 |
3.53 |
0.60 |
0.75 |
| 6 |
A' |
1153 |
1101 |
15.25 |
7.86 |
0.40 |
0.58 |
| 7 |
A' |
667 |
636 |
9.35 |
0.82 |
0.74 |
0.85 |
| 8 |
A" |
1053 |
1005 |
0.96 |
2.62 |
0.75 |
0.86 |
| 9 |
A" |
536 |
512 |
96.63 |
2.32 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 7467.4 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 7126.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.382 |
0.000 |
| O2 |
-0.898 |
-0.618 |
0.000 |
| O3 |
1.179 |
0.190 |
0.000 |
| H4 |
-0.460 |
1.390 |
0.000 |
| H5 |
-1.790 |
-0.251 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
O3 |
H4 |
H5 |
| C1 | | 1.3442 | 1.1947 | 1.1077 | 1.8985 |
O2 | 1.3442 | | 2.2288 | 2.0553 | 0.9648 | O3 | 1.1947 | 2.2288 | | 2.0312 | 3.0016 | H4 | 1.1077 | 2.0553 | 2.0312 | | 2.1120 | H5 | 1.8985 | 0.9648 | 3.0016 | 2.1120 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H5 |
109.519 |
|
O2 |
C1 |
O3 |
122.660 |
| O2 |
C1 |
H4 |
113.562 |
|
O3 |
C1 |
H4 |
123.779 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.417 |
|
|
|
| 2 |
O |
-0.449 |
|
|
|
| 3 |
O |
-0.395 |
|
|
|
| 4 |
H |
0.091 |
|
|
|
| 5 |
H |
0.336 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-3.616 |
1.505 |
0.000 |
3.917 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-15.153 |
-1.184 |
0.000 |
| y |
-1.184 |
-17.096 |
0.000 |
| z |
0.000 |
0.000 |
-16.576 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.683 |
-1.184 |
0.000 |
| y |
-1.184 |
-1.231 |
0.000 |
| z |
0.000 |
0.000 |
-0.452 |
|
| Polar |
|---|
| 3z2-r2 | -0.903 |
|---|
| x2-y2 | 1.943 |
|---|
| xy | -1.184 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.688 |
0.080 |
0.000 |
| y |
0.080 |
2.359 |
0.000 |
| z |
0.000 |
0.000 |
1.340 |
<r2> (average value of r
2) Å
2
| <r2> |
37.370 |
| (<r2>)1/2 |
6.113 |
