return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H10 (2-Pentene, (E)-)

using model chemistry: MP2=FULL/6-31G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-31G**
 hartrees
Energy at 0K-195.897285
Energy at 298.15K-195.907484
HF Energy-195.159045
Nuclear repulsion energy171.295342
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3253 3039 22.47      
2 A' 3230 3018 28.41      
3 A' 3223 3011 18.19      
4 A' 3214 3003 19.71      
5 A' 3133 2928 17.69      
6 A' 3118 2914 29.73      
7 A' 3108 2904 28.95      
8 A' 1769 1653 0.12      
9 A' 1572 1469 3.95      
10 A' 1558 1456 6.85      
11 A' 1541 1440 1.31      
12 A' 1472 1376 5.38      
13 A' 1464 1368 0.82      
14 A' 1421 1328 8.36      
15 A' 1366 1276 0.76      
16 A' 1359 1270 0.12      
17 A' 1184 1107 1.13      
18 A' 1133 1059 3.35      
19 A' 1065 995 2.18      
20 A' 986 921 5.07      
21 A' 892 834 0.05      
22 A' 591 552 0.64      
23 A' 373 349 0.15      
24 A' 215 201 0.21      
25 A" 3231 3019 28.03      
26 A" 3199 2989 21.90      
27 A" 3150 2943 19.52      
28 A" 1560 1458 5.15      
29 A" 1541 1440 4.81      
30 A" 1332 1244 0.13      
31 A" 1143 1068 1.41      
32 A" 1094 1022 0.23      
33 A" 1028 961 24.66      
34 A" 843 787 2.05      
35 A" 717 670 0.22      
36 A" 318 297 1.40      
37 A" 227 212 0.79      
38 A" 198 185 1.49      
39 A" 122 114 0.05      

Unscaled Zero Point Vibrational Energy (zpe) 30970.3 cm-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 28938.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31G**
ABC
0.44732 0.08290 0.07273

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31G**

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
H1 -0.711 -1.237 0.000
C2 -0.971 -0.185 0.000
H3 -0.289 1.783 0.000
C4 0.000 0.736 0.000
H5 -2.929 -0.279 0.876
H6 -2.929 -0.279 -0.876
H7 -2.594 1.215 0.000
C8 -2.432 0.138 0.000
H9 1.920 0.966 -0.869
H10 1.920 0.966 0.869
C11 1.478 0.471 0.000
H12 2.956 -1.102 0.000
H13 1.486 -1.507 0.880
H14 1.486 -1.507 -0.880
C15 1.873 -0.998 0.000

Atom - Atom Distances (Å)
  H1 C2 H3 C4 H5 H6 H7 C8 H9 H10 C11 H12 H13 H14 C15
H11.08343.04932.09682.57042.57043.09132.20363.53913.53912.77583.66902.38192.38192.5948
C21.08342.08331.33842.14702.14702.14321.49673.23073.23072.53514.03282.92572.92572.9581
H33.04932.08331.08643.46263.46262.37452.70212.51042.51042.20024.34223.84053.84053.5224
C42.09681.33841.08643.22113.22112.63792.50482.12002.12001.50103.48102.83082.83082.5523
H52.57042.14703.46263.22111.75271.76331.09015.30155.00604.55496.00654.58274.90784.9341
H62.57042.14703.46263.22111.75271.76331.09015.00605.30154.55496.00654.90784.58274.9341
H73.09132.14322.37452.63791.76331.76331.08824.60384.60384.13896.01434.98294.98294.9851
C82.20361.49672.70212.50481.09011.09011.08824.51484.51483.92425.52954.34024.34024.4531
H93.53913.23072.51042.12005.30155.00604.60384.51481.73801.09332.47083.05952.51022.1474
H103.53913.23072.51042.12005.00605.30154.60384.51481.73801.09332.47082.51023.05952.1474
C112.77582.53512.20021.50104.55494.55494.13893.92421.09331.09332.15922.16522.16521.5213
H123.66904.03284.34223.48106.00656.00656.01435.52952.47082.47082.15921.76041.76041.0878
H132.38192.92573.84052.83084.58274.90784.98294.34023.05952.51022.16521.76041.76061.0881
H142.38192.92573.84052.83084.90784.58274.98294.34022.51023.05952.16521.76041.76061.0881
C152.59482.95813.52242.55234.93414.93414.98514.45312.14742.14741.52131.08781.08811.0881

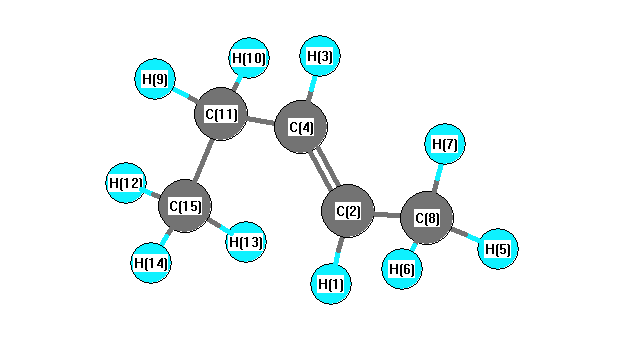
picture of 2-Pentene, (E)- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
H1 C2 C4 119.570 H1 C2 C8 116.395
C2 C4 H3 118.071 C2 C4 C11 126.360
C2 C8 H5 111.220 C2 C8 H6 111.220
C2 C8 H7 111.025 H3 C4 C11 115.569
C4 C2 C8 124.035 C4 C11 H9 108.583
C4 C11 H10 108.583 C4 C11 C15 115.232
H5 C8 H6 107.015 H5 C8 H7 108.096
H6 C8 H7 108.096 H9 C11 H10 105.282
H9 C11 C15 109.346 H10 C11 C15 109.346
C11 C15 H12 110.602 C11 C15 H13 111.057
C11 C15 H14 111.057 H12 C15 H13 108.002
H12 C15 H14 108.002 H13 C15 H14 107.995
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability