return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H6 (Cyclopropylacetylene)

using model chemistry: MP2=FULL/6-31G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A'
Energy calculated at MP2=FULL/6-31G**
 hartrees
Energy at 0K-193.439009
Energy at 298.15K-193.444164
HF Energy-192.739894
Nuclear repulsion energy147.448170
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3548 3316 60.54      
2 A 3350 3130 9.29      
3 A 3254 3041 5.91      
4 A 3244 3031 3.57      
5 A 2175 2032 3.72      
6 A 1568 1465 0.36      
7 A 1427 1334 4.83      
8 A 1263 1180 1.38      
9 A 1170 1093 0.52      
10 A 1115 1042 5.63      
11 A 1008 942 18.39      
12 A 839 784 1.05      
13 A 784 733 1.95      
14 A 601 561 47.55      
15 A 497 464 1.20      
16 A 198 185 0.55      
17 A 3338 3119 0.14      
18 A 3239 3027 10.84      
19 A 1525 1425 1.12      
20 A 1243 1161 0.47      
21 A 1164 1088 2.25      
22 A 1117 1043 4.72      
23 A 949 887 3.18      
24 A 859 803 7.29      
25 A 557 520 35.81      
26 A 485 453 13.36      
27 A 204 190 0.33      

Unscaled Zero Point Vibrational Energy (zpe) 20360.7 cm-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 19025.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31G**
ABC
0.53256 0.11135 0.10583

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31G**

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.641 0.000
C2 -1.076 -0.314 0.000
H3 -0.305 1.678 0.000
C4 1.276 0.326 0.747
C5 1.276 0.326 -0.747
C6 -1.988 -1.126 0.000
H7 -2.780 -1.832 0.000
H8 1.761 1.141 1.262
H9 1.319 -0.626 1.253
H10 1.761 1.141 -1.262
H11 1.319 -0.626 -1.253

Atom - Atom Distances (Å)
  C1 C2 H3 C4 C5 C6 H7 H8 H9 H10 H11
C11.43881.08161.51161.51162.65943.72082.22362.21662.22362.2166
C21.43882.13682.54972.54971.22062.28203.42942.72133.42942.7213
H31.08162.13682.21012.21013.27064.29562.47953.08482.47953.0848
C41.51162.54972.21011.49433.64974.65501.07921.07882.22202.2153
C51.51162.54972.21011.49433.64974.65502.22202.21531.07921.0788
C62.65941.22063.27063.64973.64971.06144.55923.57194.55923.5719
H73.72082.28204.29564.65504.65501.06145.57274.45335.57274.4533
H82.22363.42942.47951.07922.22204.55925.57271.82142.52473.1052
H92.21662.72133.08481.07882.21533.57194.45331.82143.10522.5052
H102.22363.42942.47952.22201.07924.55925.57272.52473.10521.8214
H112.21662.72133.08482.21531.07883.57194.45333.10522.50521.8214

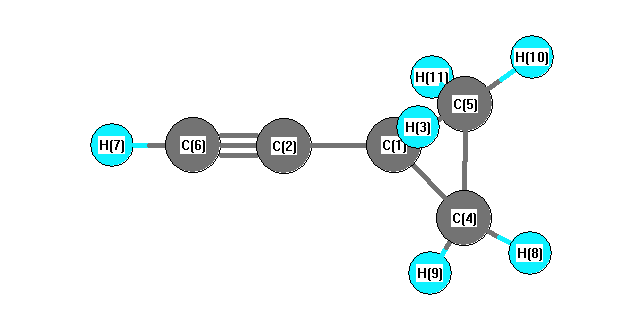
picture of Cyclopropylacetylene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C6 179.908 C1 C4 C5 60.377
C1 C4 H8 117.267 C1 C4 H9 116.688
C1 C5 C4 60.377 C1 C5 H10 117.267
C1 C5 H11 116.688 C2 C1 H3 115.214
C2 C1 C4 119.567 C2 C1 C5 119.567
C2 C6 H7 179.946 H3 C1 C4 115.930
H3 C1 C5 115.930 C4 C1 C5 59.247
C4 C5 H10 118.513 C4 C5 H11 117.936
C5 C4 H8 118.513 C5 C4 H9 117.936
H8 C4 H9 115.133 H10 C5 H11 115.133
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability