return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CHOHCH3 (Isopropyl alcohol)

using model chemistry: QCISD/6-31G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at QCISD/6-31G**
 hartrees
Energy at 0K-193.804752
Energy at 298.15K-193.813862
HF Energy-193.131942
Nuclear repulsion energy134.882848
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3894 3666 10.40      
2 A 3205 3017 26.98      
3 A 3198 3010 57.39      
4 A 3191 3004 1.42      
5 A 3175 2989 35.06      
6 A 3111 2929 11.85      
7 A 3097 2915 21.28      
8 A 3049 2871 61.10      
9 A 1568 1476 3.93      
10 A 1557 1466 2.69      
11 A 1547 1457 0.83      
12 A 1543 1452 0.64      
13 A 1491 1404 18.22      
14 A 1471 1385 15.25      
15 A 1443 1359 0.15      
16 A 1427 1344 16.45      
17 A 1319 1241 57.88      
18 A 1234 1162 31.06      
19 A 1200 1130 14.49      
20 A 1127 1061 25.29      
21 A 1010 950 37.75      
22 A 982 924 1.76      
23 A 956 900 0.10      
24 A 851 801 3.84      
25 A 492 463 9.54      
26 A 423 398 9.93      
27 A 368 347 1.73      
28 A 322 303 120.46      
29 A 282 265 0.83      
30 A 234 220 4.32      

Unscaled Zero Point Vibrational Energy (zpe) 24382.5 cm-1
Scaled (by 0.9414) Zero Point Vibrational Energy (zpe) 22953.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/6-31G**
ABC
0.29011 0.26876 0.15958

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/6-31G**

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.001 0.039 0.368
C2 -1.201 -0.772 -0.104
C3 1.321 -0.559 -0.088
O4 -0.037 1.365 -0.169
H5 -0.008 0.084 1.466
H6 -2.135 -0.310 0.223
H7 -1.206 -0.822 -1.194
H8 -1.170 -1.787 0.295
H9 2.151 0.060 0.250
H10 1.446 -1.567 0.311
H11 1.346 -0.606 -1.178
H12 -0.855 1.776 0.129

Atom - Atom Distances (Å)
  C1 C2 C3 O4 H5 H6 H7 H8 H9 H10 H11 H12
C11.52481.51911.43091.09882.16922.15392.17112.15292.16122.14851.9513
C21.52482.53032.43422.14921.09261.09081.09143.47112.79452.76892.5821
C31.51912.53032.35642.14353.47862.77092.80361.08941.09081.09093.1994
O41.43092.43422.35642.07722.71332.68383.38192.58123.32042.61120.9631
H51.09882.14922.14352.07722.49423.05452.49422.47802.48463.04962.3170
H62.16921.09263.47862.71332.49421.77011.76554.30163.79623.76442.4495
H72.15391.09082.77092.68383.05451.77011.77493.75923.13932.56192.9370
H82.17111.09142.80363.38192.49421.76551.77493.80062.62603.14613.5812
H92.15293.47111.08942.58122.47804.30163.75923.80061.77391.76893.4632
H102.16122.79451.09083.32042.48463.79623.13932.62601.77391.77434.0626
H112.14852.76891.09092.61123.04963.76442.56193.14611.76891.77433.4975
H121.95132.58213.19940.96312.31702.44952.93703.58123.46324.06263.4975

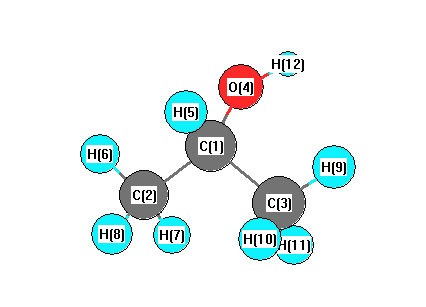
picture of Isopropyl alcohol state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 H6 110.863 C1 C2 H7 109.759
C1 C2 H8 111.083 C1 C3 H9 110.153
C1 C3 H10 110.730 C1 C3 H11 109.721
C1 O4 H12 107.580 C2 C1 C3 112.453
C2 C1 O4 110.847 C2 C1 H5 108.921
C3 C1 O4 105.988 C3 C1 H5 108.867
O4 C1 H5 109.703 H6 C2 H7 108.335
H6 C2 H8 107.878 H7 C2 H8 108.845
H9 C3 H10 108.908 H9 C3 H11 108.450
H10 C3 H11 108.830
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability