return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3SOCH3 (Dimethyl sulfoxide)

using model chemistry: MP2=FULL/6-31G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-31G**
 hartrees
Energy at 0K-552.185126
Energy at 298.15K-552.192147
HF Energy-551.546145
Nuclear repulsion energy183.686485
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3264 3050 2.88 95.17 0.71 0.83
2 A' 3255 3041 8.34 89.66 0.74 0.85
3 A' 3135 2929 5.32 186.93 0.00 0.00
4 A' 1537 1436 19.45 3.17 0.74 0.85
5 A' 1514 1414 2.70 23.85 0.73 0.84
6 A' 1413 1321 8.86 0.48 0.74 0.85
7 A' 1151 1076 128.98 5.63 0.45 0.62
8 A' 1069 999 21.34 6.38 0.60 0.75
9 A' 1005 939 11.05 7.39 0.73 0.85
10 A' 701 655 10.86 26.70 0.11 0.20
11 A' 377 352 8.99 1.30 0.26 0.42
12 A' 301 281 0.80 1.47 0.75 0.85
13 A' 249 232 0.59 0.07 0.53 0.69
14 A" 3263 3049 1.71 33.83 0.75 0.86
15 A" 3252 3038 0.01 5.82 0.75 0.86
16 A" 3134 2928 2.50 0.31 0.75 0.86
17 A" 1516 1417 0.82 29.01 0.75 0.86
18 A" 1499 1400 8.68 5.01 0.75 0.86
19 A" 1389 1298 2.26 1.09 0.75 0.86
20 A" 973 909 6.47 5.62 0.75 0.86
21 A" 934 872 2.17 0.64 0.75 0.86
22 A" 730 682 21.76 15.27 0.75 0.86
23 A" 305 285 8.35 3.43 0.75 0.86
24 A" 181 169 0.00 0.04 0.75 0.86

Unscaled Zero Point Vibrational Energy (zpe) 18071.4 cm-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 16885.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31G**
ABC
0.23271 0.22810 0.13910

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31G**

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
S1 0.258 0.424 0.000
O2 -1.094 1.095 0.000
C3 0.258 -0.789 1.337
C4 0.258 -0.789 -1.337
H5 1.166 -1.386 1.300
H6 1.166 -1.386 -1.300
H7 0.217 -0.239 2.273
H8 0.217 -0.239 -2.273
H9 -0.624 -1.419 1.250
H10 -0.624 -1.419 -1.250

Atom - Atom Distances (Å)
  S1 O2 C3 C4 H5 H6 H7 H8 H9 H10
S11.50941.80511.80512.40692.40692.36792.36792.39512.3951
O21.50942.67652.67653.59933.59932.94362.94362.84622.8462
C31.80512.67652.67381.08762.85251.08593.65141.08772.8047
C41.80512.67652.67382.85251.08763.65141.08592.80471.0877
H52.40693.59931.08762.85252.60061.77813.87071.79133.1159
H62.40693.59932.85251.08762.60063.87071.77813.11591.7913
H72.36792.94361.08593.65141.77813.87074.54541.77363.8088
H82.36792.94363.65141.08593.87071.77814.54543.80881.7736
H92.39512.84621.08772.80471.79133.11591.77363.80882.4995
H102.39512.84622.80471.08773.11591.79133.80881.77362.4995

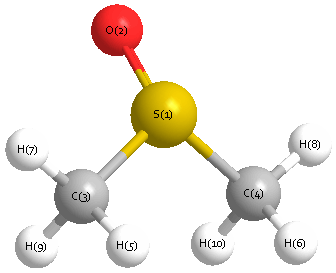
picture of Dimethyl sulfoxide state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
S1 C3 H5 110.135 S1 C3 H7 107.358
S1 C3 H9 109.260 S1 C4 H6 110.135
S1 C4 H8 107.358 S1 C4 H10 109.260
O2 S1 C3 107.370 O2 S1 C4 107.370
C3 S1 C4 95.571 H5 C3 H7 109.780
H5 C3 H9 110.870 H6 C4 H8 109.780
H6 C4 H10 110.870 H7 C3 H9 109.365
H8 C4 H10 109.365
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability