return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H7NO (dimethylformamide)

using model chemistry: QCISD/6-31G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at QCISD/6-31G**
 hartrees
Energy at 0K-247.816059
Energy at 298.15K-247.824406
HF Energy-247.000099
Nuclear repulsion energy181.348447
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3244 3054 1.39      
2 A' 3201 3014 20.97      
3 A' 3096 2915 53.70      
4 A' 3088 2907 48.05      
5 A' 3051 2873 88.59      
6 A' 1814 1707 405.24      
7 A' 1595 1502 13.90      
8 A' 1567 1475 12.43      
9 A' 1520 1431 10.12      
10 A' 1489 1402 57.53      
11 A' 1483 1396 9.55      
12 A' 1465 1379 70.03      
13 A' 1333 1255 40.07      
14 A' 1131 1064 114.13      
15 A' 1115 1049 2.50      
16 A' 900 847 2.66      
17 A' 669 630 7.36      
18 A' 398 375 1.30      
19 A' 330 311 12.30      
20 A" 3159 2973 22.88      
21 A" 3152 2967 52.28      
22 A" 1558 1466 11.02      
23 A" 1535 1445 2.90      
24 A" 1210 1139 5.36      
25 A" 1162 1094 0.02      
26 A" 1042 981 1.09      
27 A" 330 311 20.24      
28 A" 226 213 2.42      
29 A" 164 154 0.48      
30 A" 85 80 0.05      

Unscaled Zero Point Vibrational Energy (zpe) 23054.7 cm-1
Scaled (by 0.9414) Zero Point Vibrational Energy (zpe) 21703.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/6-31G**
ABC
0.29835 0.13905 0.09836

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/6-31G**

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.701 -0.821 0.000
O2 0.212 -1.944 0.000
N3 0.000 0.349 0.000
C4 -1.452 0.329 0.000
C5 0.660 1.638 0.000
H6 1.789 -0.650 0.000
H7 -1.773 -0.708 0.000
H8 -1.841 0.834 0.888
H9 -1.841 0.834 -0.888
H10 1.739 1.495 0.000
H11 0.385 2.216 -0.886
H12 0.385 2.216 0.886

Atom - Atom Distances (Å)
  C1 O2 N3 C4 C5 H6 H7 H8 H9 H10 H11 H12
C11.22421.36422.44122.46021.10132.47713.16053.16052.53873.17973.1797
O21.22422.30222.81683.60992.03952.33853.56603.56603.76294.25644.2564
N31.36422.30221.45191.44862.04932.06422.10022.10022.08342.10242.1024
C42.44122.81681.45192.48463.38581.08581.09251.09253.39772.77872.7787
C52.46023.60991.44862.48462.55233.38022.77262.77261.08911.09291.0929
H61.10132.03952.04933.38582.55233.56304.02074.02072.14633.31243.3124
H72.47712.33852.06421.08583.38023.56301.78031.78034.14663.74093.7409
H83.16053.56602.10021.09252.77264.02071.78031.77503.74733.16422.6201
H93.16053.56602.10021.09252.77264.02071.78031.77503.74732.62013.1642
H102.53873.76292.08343.39771.08912.14634.14663.74733.74731.77161.7716
H113.17974.25642.10242.77871.09293.31243.74093.16422.62011.77161.7729
H123.17974.25642.10242.77871.09293.31243.74092.62013.16421.77161.7729

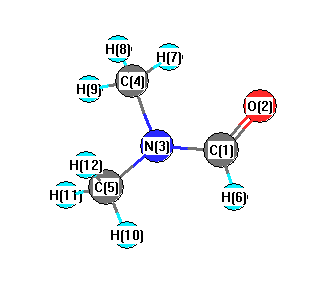
picture of dimethylformamide state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 N3 C4 120.158 C1 N3 C5 121.969
O2 C1 N3 125.515 O2 C1 H6 122.483
N3 C1 H6 112.002 N3 C4 H7 107.998
N3 C4 H8 110.461 N3 C4 H9 110.461
N3 C5 H10 109.547 N3 C5 H11 110.854
N3 C5 H12 110.854 C4 N3 C5 117.873
H7 C4 H8 109.630 H7 C4 H9 109.630
H8 C4 H9 108.651 H10 C5 H11 108.561
H10 C5 H12 108.561 H11 C5 H12 108.399
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability