Jump to
S1C2
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
Geometric Data calculated at HSEh1PBE/6-31G**
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at HSEh1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -194.145869 |
| Energy at 298.15K | |
| HF Energy | -194.145869 |
| Nuclear repulsion energy | 133.067494 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3895 |
3717 |
21.84 |
114.38 |
0.30 |
0.46 |
| 2 |
A |
3167 |
3022 |
21.30 |
40.86 |
0.75 |
0.86 |
| 3 |
A |
3142 |
2998 |
42.64 |
54.42 |
0.61 |
0.76 |
| 4 |
A |
3110 |
2967 |
19.65 |
87.67 |
0.73 |
0.85 |
| 5 |
A |
3066 |
2926 |
40.18 |
107.01 |
0.08 |
0.14 |
| 6 |
A |
3064 |
2924 |
26.16 |
106.86 |
0.03 |
0.05 |
| 7 |
A |
3024 |
2885 |
61.93 |
85.87 |
0.75 |
0.85 |
| 8 |
A |
2987 |
2851 |
67.73 |
102.74 |
0.12 |
0.21 |
| 9 |
A |
1536 |
1466 |
3.74 |
7.05 |
0.66 |
0.79 |
| 10 |
A |
1521 |
1451 |
7.93 |
9.42 |
0.75 |
0.86 |
| 11 |
A |
1504 |
1436 |
7.57 |
21.69 |
0.75 |
0.86 |
| 12 |
A |
1487 |
1419 |
3.06 |
17.42 |
0.75 |
0.86 |
| 13 |
A |
1467 |
1400 |
4.36 |
2.28 |
0.52 |
0.69 |
| 14 |
A |
1422 |
1357 |
4.81 |
3.86 |
0.75 |
0.85 |
| 15 |
A |
1382 |
1319 |
1.11 |
1.51 |
0.75 |
0.85 |
| 16 |
A |
1334 |
1273 |
25.77 |
15.89 |
0.74 |
0.85 |
| 17 |
A |
1279 |
1221 |
1.49 |
10.19 |
0.75 |
0.85 |
| 18 |
A |
1257 |
1199 |
42.80 |
5.12 |
0.68 |
0.81 |
| 19 |
A |
1171 |
1117 |
8.51 |
1.21 |
0.42 |
0.59 |
| 20 |
A |
1150 |
1097 |
19.83 |
3.87 |
0.55 |
0.71 |
| 21 |
A |
1087 |
1037 |
37.02 |
2.92 |
0.72 |
0.83 |
| 22 |
A |
1007 |
961 |
39.94 |
4.19 |
0.75 |
0.86 |
| 23 |
A |
932 |
889 |
2.87 |
0.31 |
0.67 |
0.80 |
| 24 |
A |
886 |
846 |
2.39 |
9.31 |
0.19 |
0.32 |
| 25 |
A |
778 |
742 |
0.67 |
0.51 |
0.60 |
0.75 |
| 26 |
A |
483 |
460 |
7.84 |
0.25 |
0.50 |
0.66 |
| 27 |
A |
331 |
316 |
10.44 |
0.45 |
0.57 |
0.73 |
| 28 |
A |
261 |
249 |
114.31 |
4.36 |
0.74 |
0.85 |
| 29 |
A |
226 |
216 |
2.41 |
0.08 |
0.61 |
0.76 |
| 30 |
A |
151 |
144 |
7.56 |
0.12 |
0.75 |
0.85 |
Unscaled Zero Point Vibrational Energy (zpe) 24052.3 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 22953.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G**
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.528 |
-0.515 |
0.128 |
| C2 |
-0.632 |
0.644 |
-0.285 |
| C3 |
0.768 |
0.541 |
0.289 |
| O4 |
1.368 |
-0.635 |
-0.219 |
| H5 |
-2.520 |
-0.434 |
-0.327 |
| H6 |
-1.661 |
-0.542 |
1.215 |
| H7 |
-1.087 |
-1.467 |
-0.178 |
| H8 |
-0.551 |
0.685 |
-1.378 |
| H9 |
-1.066 |
1.599 |
0.036 |
| H10 |
0.711 |
0.516 |
1.390 |
| H11 |
1.349 |
1.436 |
0.012 |
| H12 |
2.230 |
-0.734 |
0.195 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
| C1 | | 1.5227 | 2.5330 | 2.9200 | 1.0940 | 1.0959 | 1.0927 | 2.1598 | 2.1661 | 2.7696 | 3.4784 | 3.7653 |
C2 | 1.5227 | | 1.5168 | 2.3751 | 2.1750 | 2.1721 | 2.1618 | 1.0967 | 1.0971 | 2.1510 | 2.1539 | 3.2126 | C3 | 2.5330 | 1.5168 | | 1.4143 | 3.4848 | 2.8165 | 2.7729 | 2.1308 | 2.1327 | 1.1034 | 1.1021 | 1.9419 | O4 | 2.9200 | 2.3751 | 1.4143 | | 3.8952 | 3.3531 | 2.5925 | 2.6021 | 3.3138 | 2.0845 | 2.0838 | 0.9612 | H5 | 1.0940 | 2.1750 | 3.4848 | 3.8952 | | 1.7687 | 1.7727 | 2.4973 | 2.5259 | 3.7802 | 4.3108 | 4.7881 | H6 | 1.0959 | 2.1721 | 2.8165 | 3.3531 | 1.7687 | | 1.7684 | 3.0765 | 2.5157 | 2.6027 | 3.7972 | 4.0268 | H7 | 1.0927 | 2.1618 | 2.7729 | 2.5925 | 1.7727 | 1.7684 | | 2.5213 | 3.0731 | 3.1017 | 3.7939 | 3.4169 | H8 | 2.1598 | 1.0967 | 2.1308 | 2.6021 | 2.4973 | 3.0765 | 2.5213 | | 1.7606 | 3.0473 | 2.4714 | 3.4965 | H9 | 2.1661 | 1.0971 | 2.1327 | 3.3138 | 2.5259 | 2.5157 | 3.0731 | 1.7606 | | 2.4831 | 2.4206 | 4.0414 | H10 | 2.7696 | 2.1510 | 1.1034 | 2.0845 | 3.7802 | 2.6027 | 3.1017 | 3.0473 | 2.4831 | | 1.7755 | 2.3015 | H11 | 3.4784 | 2.1539 | 1.1021 | 2.0838 | 4.3108 | 3.7972 | 3.7939 | 2.4714 | 2.4206 | 1.7755 | | 2.3495 | H12 | 3.7653 | 3.2126 | 1.9419 | 0.9612 | 4.7881 | 4.0268 | 3.4169 | 3.4965 | 4.0414 | 2.3015 | 2.3495 | |
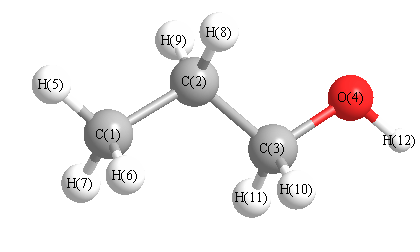 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
112.897 |
|
C1 |
C2 |
H8 |
110.023 |
| C1 |
C2 |
H9 |
110.497 |
|
C2 |
C1 |
H5 |
111.392 |
| C2 |
C1 |
H6 |
111.045 |
|
C2 |
C1 |
H7 |
110.413 |
| C2 |
C3 |
O4 |
108.204 |
|
C2 |
C3 |
H10 |
109.349 |
| C2 |
C3 |
H11 |
109.647 |
|
C3 |
C2 |
H8 |
108.162 |
| C3 |
C2 |
H9 |
108.288 |
|
C3 |
O4 |
H12 |
108.156 |
| O4 |
C3 |
H10 |
111.184 |
|
O4 |
C3 |
H11 |
111.206 |
| H5 |
C1 |
H6 |
107.736 |
|
H5 |
C1 |
H7 |
108.323 |
| H6 |
C1 |
H7 |
107.797 |
|
H8 |
C2 |
H9 |
106.753 |
| H10 |
C3 |
H11 |
107.237 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.389 |
|
|
|
| 2 |
C |
-0.251 |
|
|
|
| 3 |
C |
0.008 |
|
|
|
| 4 |
O |
-0.546 |
|
|
|
| 5 |
H |
0.125 |
|
|
|
| 6 |
H |
0.120 |
|
|
|
| 7 |
H |
0.154 |
|
|
|
| 8 |
H |
0.137 |
|
|
|
| 9 |
H |
0.120 |
|
|
|
| 10 |
H |
0.100 |
|
|
|
| 11 |
H |
0.103 |
|
|
|
| 12 |
H |
0.319 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.788 |
0.806 |
0.983 |
1.496 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-22.358 |
-0.318 |
2.001 |
| y |
-0.318 |
-26.682 |
-0.597 |
| z |
2.001 |
-0.597 |
-26.465 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.216 |
-0.318 |
2.001 |
| y |
-0.318 |
-2.270 |
-0.597 |
| z |
2.001 |
-0.597 |
-1.946 |
|
| Polar |
|---|
| 3z2-r2 | -3.892 |
|---|
| x2-y2 | 4.324 |
|---|
| xy | -0.318 |
|---|
| xz | 2.001 |
|---|
| yz | -0.597 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.060 |
0.001 |
0.129 |
| y |
0.001 |
5.376 |
0.020 |
| z |
0.129 |
0.020 |
5.103 |
<r2> (average value of r
2) Å
2
| <r2> |
94.228 |
| (<r2>)1/2 |
9.707 |
