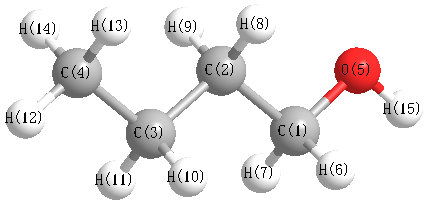
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3891 |
3713 |
19.11 |
|
|
|
| 2 |
A' |
3147 |
3003 |
33.77 |
|
|
|
| 3 |
A' |
3063 |
2923 |
53.59 |
|
|
|
| 4 |
A' |
3060 |
2920 |
15.52 |
|
|
|
| 5 |
A' |
3045 |
2905 |
21.38 |
|
|
|
| 6 |
A' |
2984 |
2848 |
59.49 |
|
|
|
| 7 |
A' |
1540 |
1470 |
3.54 |
|
|
|
| 8 |
A' |
1519 |
1450 |
5.29 |
|
|
|
| 9 |
A' |
1506 |
1438 |
1.16 |
|
|
|
| 10 |
A' |
1499 |
1431 |
0.09 |
|
|
|
| 11 |
A' |
1476 |
1408 |
7.84 |
|
|
|
| 12 |
A' |
1426 |
1360 |
2.84 |
|
|
|
| 13 |
A' |
1404 |
1340 |
4.19 |
|
|
|
| 14 |
A' |
1319 |
1259 |
29.31 |
|
|
|
| 15 |
A' |
1254 |
1196 |
41.47 |
|
|
|
| 16 |
A' |
1138 |
1086 |
18.57 |
|
|
|
| 17 |
A' |
1105 |
1055 |
50.71 |
|
|
|
| 18 |
A' |
1084 |
1035 |
22.55 |
|
|
|
| 19 |
A' |
1029 |
982 |
1.97 |
|
|
|
| 20 |
A' |
920 |
878 |
13.81 |
|
|
|
| 21 |
A' |
444 |
423 |
12.75 |
|
|
|
| 22 |
A' |
399 |
381 |
0.10 |
|
|
|
| 23 |
A' |
186 |
177 |
2.45 |
|
|
|
| 24 |
A" |
3141 |
2997 |
51.70 |
|
|
|
| 25 |
A" |
3114 |
2972 |
39.33 |
|
|
|
| 26 |
A" |
3080 |
2939 |
5.35 |
|
|
|
| 27 |
A" |
3017 |
2879 |
60.47 |
|
|
|
| 28 |
A" |
1510 |
1441 |
7.25 |
|
|
|
| 29 |
A" |
1332 |
1271 |
0.06 |
|
|
|
| 30 |
A" |
1324 |
1263 |
1.10 |
|
|
|
| 31 |
A" |
1255 |
1198 |
0.14 |
|
|
|
| 32 |
A" |
1196 |
1141 |
2.37 |
|
|
|
| 33 |
A" |
959 |
915 |
0.03 |
|
|
|
| 34 |
A" |
818 |
781 |
1.48 |
|
|
|
| 35 |
A" |
749 |
715 |
2.78 |
|
|
|
| 36 |
A" |
283 |
270 |
123.92 |
|
|
|
| 37 |
A" |
255 |
243 |
1.45 |
|
|
|
| 38 |
A" |
118 |
112 |
2.17 |
|
|
|
| 39 |
A" |
113 |
108 |
4.02 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30350.1 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 28963.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.013 |
|
|
|
| 2 |
C |
-0.239 |
|
|
|
| 3 |
C |
-0.246 |
|
|
|
| 4 |
C |
-0.388 |
|
|
|
| 5 |
O |
-0.553 |
|
|
|
| 6 |
H |
0.100 |
|
|
|
| 7 |
H |
0.100 |
|
|
|
| 8 |
H |
0.133 |
|
|
|
| 9 |
H |
0.133 |
|
|
|
| 10 |
H |
0.122 |
|
|
|
| 11 |
H |
0.122 |
|
|
|
| 12 |
H |
0.127 |
|
|
|
| 13 |
H |
0.130 |
|
|
|
| 14 |
H |
0.130 |
|
|
|
| 15 |
H |
0.318 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.322 |
-1.464 |
0.000 |
1.499 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-28.637 |
-3.391 |
0.000 |
| y |
-3.391 |
-34.430 |
0.000 |
| z |
0.000 |
0.000 |
-32.793 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.974 |
-3.391 |
0.000 |
| y |
-3.391 |
-3.715 |
0.000 |
| z |
0.000 |
0.000 |
-1.260 |
|
| Polar |
|---|
| 3z2-r2 | -2.519 |
|---|
| x2-y2 | 5.793 |
|---|
| xy | -3.391 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.388 |
0.070 |
0.000 |
| y |
0.070 |
6.667 |
0.000 |
| z |
0.000 |
0.000 |
6.308 |
<r2> (average value of r
2) Å
2
| <r2> |
185.704 |
| (<r2>)1/2 |
13.627 |