Jump to
S1C2
Energy calculated at HSEh1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -79.734223 |
| Energy at 298.15K | -79.740137 |
| HF Energy | -79.734223 |
| Nuclear repulsion energy | 42.246848 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1g |
3065 |
2925 |
0.00 |
221.32 |
0.01 |
0.02 |
| 2 |
A1g |
1440 |
1374 |
0.00 |
7.36 |
0.73 |
0.85 |
| 3 |
A1g |
1031 |
984 |
0.00 |
8.62 |
0.30 |
0.46 |
| 4 |
A1u |
315 |
301 |
0.00 |
0.00 |
0.00 |
0.00 |
| 5 |
A2u |
3066 |
2926 |
54.56 |
0.00 |
0.00 |
0.00 |
| 6 |
A2u |
1411 |
1346 |
1.36 |
0.00 |
0.00 |
0.00 |
| 7 |
Eg |
3132 |
2989 |
0.00 |
133.39 |
0.75 |
0.86 |
| 7 |
Eg |
3132 |
2989 |
0.00 |
133.39 |
0.75 |
0.86 |
| 8 |
Eg |
1514 |
1445 |
0.00 |
38.14 |
0.75 |
0.86 |
| 8 |
Eg |
1514 |
1445 |
0.00 |
38.14 |
0.75 |
0.86 |
| 9 |
Eg |
1224 |
1168 |
0.00 |
2.29 |
0.75 |
0.86 |
| 9 |
Eg |
1224 |
1168 |
0.00 |
2.29 |
0.75 |
0.86 |
| 10 |
Eu |
3156 |
3011 |
59.56 |
0.00 |
0.00 |
0.00 |
| 10 |
Eu |
3156 |
3011 |
59.56 |
0.00 |
0.00 |
0.00 |
| 11 |
Eu |
1517 |
1447 |
9.21 |
0.00 |
0.00 |
0.00 |
| 11 |
Eu |
1517 |
1447 |
9.21 |
0.00 |
0.00 |
0.00 |
| 12 |
Eu |
827 |
789 |
5.93 |
0.00 |
0.00 |
0.00 |
| 12 |
Eu |
827 |
789 |
5.93 |
0.00 |
0.00 |
0.00 |
Unscaled Zero Point Vibrational Energy (zpe) 16532.9 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 15777.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G**
Point Group is D3d
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.761 |
| C2 |
0.000 |
0.000 |
-0.761 |
| H3 |
0.000 |
1.019 |
1.161 |
| H4 |
-0.882 |
-0.509 |
1.161 |
| H5 |
0.882 |
-0.509 |
1.161 |
| H6 |
0.000 |
-1.019 |
-1.161 |
| H7 |
-0.882 |
0.509 |
-1.161 |
| H8 |
0.882 |
0.509 |
-1.161 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5220 | 1.0945 | 1.0945 | 1.0945 | 2.1751 | 2.1751 | 2.1751 |
C2 | 1.5220 | | 2.1751 | 2.1751 | 2.1751 | 1.0945 | 1.0945 | 1.0945 | H3 | 1.0945 | 2.1751 | | 1.7647 | 1.7647 | 3.0890 | 2.5353 | 2.5353 | H4 | 1.0945 | 2.1751 | 1.7647 | | 1.7647 | 2.5353 | 2.5353 | 3.0890 | H5 | 1.0945 | 2.1751 | 1.7647 | 1.7647 | | 2.5353 | 3.0890 | 2.5353 | H6 | 2.1751 | 1.0945 | 3.0890 | 2.5353 | 2.5353 | | 1.7647 | 1.7647 | H7 | 2.1751 | 1.0945 | 2.5353 | 2.5353 | 3.0890 | 1.7647 | | 1.7647 | H8 | 2.1751 | 1.0945 | 2.5353 | 3.0890 | 2.5353 | 1.7647 | 1.7647 | |
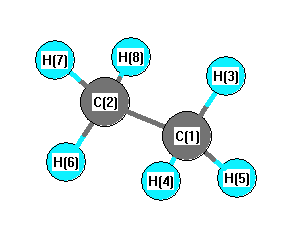 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H6 |
111.426 |
|
C1 |
C2 |
H7 |
111.426 |
| C1 |
C2 |
H8 |
111.426 |
|
C2 |
C1 |
H3 |
111.426 |
| C2 |
C1 |
H4 |
111.426 |
|
C2 |
C1 |
H5 |
111.426 |
| H3 |
C1 |
H4 |
107.447 |
|
H3 |
C1 |
H5 |
107.447 |
| H4 |
C1 |
H5 |
107.447 |
|
H6 |
C2 |
H7 |
107.447 |
| H6 |
C2 |
H8 |
107.447 |
|
H7 |
C2 |
H8 |
107.447 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.392 |
|
|
|
| 2 |
C |
-0.392 |
|
|
|
| 3 |
H |
0.131 |
|
|
|
| 4 |
H |
0.131 |
|
|
|
| 5 |
H |
0.131 |
|
|
|
| 6 |
H |
0.131 |
|
|
|
| 7 |
H |
0.131 |
|
|
|
| 8 |
H |
0.131 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-14.648 |
0.000 |
0.000 |
| y |
0.000 |
-14.648 |
0.000 |
| z |
0.000 |
0.000 |
-15.105 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.229 |
0.000 |
0.000 |
| y |
0.000 |
0.229 |
0.000 |
| z |
0.000 |
0.000 |
-0.457 |
|
| Polar |
|---|
| 3z2-r2 | -0.914 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.384 |
0.000 |
0.000 |
| y |
0.000 |
3.384 |
0.000 |
| z |
0.000 |
0.000 |
3.631 |
<r2> (average value of r
2) Å
2
| <r2> |
30.506 |
| (<r2>)1/2 |
5.523 |
Jump to
S1C1
Energy calculated at HSEh1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -79.729701 |
| Energy at 298.15K | |
| HF Energy | -79.729701 |
| Nuclear repulsion energy | 42.086016 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1' |
3082 |
2941 |
0.00 |
|
|
|
| 2 |
A1' |
1458 |
1392 |
0.00 |
|
|
|
| 3 |
A1' |
1030 |
983 |
0.00 |
|
|
|
| 4 |
A1" |
306i |
292i |
0.00 |
|
|
|
| 5 |
A2" |
3075 |
2934 |
56.62 |
|
|
|
| 6 |
A2" |
1409 |
1344 |
1.23 |
|
|
|
| 7 |
E' |
3164 |
3019 |
49.25 |
|
|
|
| 7 |
E' |
3164 |
3019 |
49.23 |
|
|
|
| 8 |
E' |
1523 |
1453 |
10.55 |
|
|
|
| 8 |
E' |
1523 |
1453 |
10.55 |
|
|
|
| 9 |
E' |
906 |
865 |
5.42 |
|
|
|
| 9 |
E' |
906 |
865 |
5.42 |
|
|
|
| 10 |
E" |
3144 |
3000 |
0.00 |
|
|
|
| 10 |
E" |
3144 |
3000 |
0.00 |
|
|
|
| 11 |
E" |
1513 |
1444 |
0.00 |
|
|
|
| 11 |
E" |
1513 |
1444 |
0.00 |
|
|
|
| 12 |
E" |
1173 |
1120 |
0.00 |
|
|
|
| 12 |
E" |
1173 |
1120 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16296.5 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 15551.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G**
Point Group is D3h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.768 |
| C2 |
0.000 |
0.000 |
-0.768 |
| H3 |
0.000 |
1.014 |
1.176 |
| H4 |
-0.879 |
-0.507 |
1.176 |
| H5 |
0.879 |
-0.507 |
1.176 |
| H6 |
0.000 |
1.014 |
-1.176 |
| H7 |
0.879 |
-0.507 |
-1.176 |
| H8 |
-0.879 |
-0.507 |
-1.176 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5353 | 1.0934 | 1.0934 | 1.0934 | 2.1921 | 2.1921 | 2.1921 |
C2 | 1.5353 | | 2.1921 | 2.1921 | 2.1921 | 1.0934 | 1.0934 | 1.0934 | H3 | 1.0934 | 2.1921 | | 1.7571 | 1.7571 | 2.3511 | 2.9352 | 2.9352 | H4 | 1.0934 | 2.1921 | 1.7571 | | 1.7571 | 2.9352 | 2.9352 | 2.3511 | H5 | 1.0934 | 2.1921 | 1.7571 | 1.7571 | | 2.9352 | 2.3511 | 2.9352 | H6 | 2.1921 | 1.0934 | 2.3511 | 2.9352 | 2.9352 | | 1.7571 | 1.7571 | H7 | 2.1921 | 1.0934 | 2.9352 | 2.9352 | 2.3511 | 1.7571 | | 1.7571 | H8 | 2.1921 | 1.0934 | 2.9352 | 2.3511 | 2.9352 | 1.7571 | 1.7571 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H6 |
111.905 |
|
C1 |
C2 |
H7 |
111.905 |
| C1 |
C2 |
H8 |
111.905 |
|
C2 |
C1 |
H3 |
111.905 |
| C2 |
C1 |
H4 |
111.905 |
|
C2 |
C1 |
H5 |
111.905 |
| H3 |
C1 |
H4 |
106.932 |
|
H3 |
C1 |
H5 |
106.932 |
| H4 |
C1 |
H5 |
106.931 |
|
H6 |
C2 |
H7 |
106.932 |
| H6 |
C2 |
H8 |
106.932 |
|
H7 |
C2 |
H8 |
106.931 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.401 |
|
|
|
| 2 |
C |
-0.401 |
|
|
|
| 3 |
H |
0.134 |
|
|
|
| 4 |
H |
0.134 |
|
|
|
| 5 |
H |
0.134 |
|
|
|
| 6 |
H |
0.134 |
|
|
|
| 7 |
H |
0.134 |
|
|
|
| 8 |
H |
0.134 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-14.631 |
0.000 |
0.000 |
| y |
0.000 |
-14.631 |
0.000 |
| z |
0.000 |
0.000 |
-15.042 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.206 |
0.000 |
0.000 |
| y |
0.000 |
0.206 |
0.000 |
| z |
0.000 |
0.000 |
-0.412 |
|
| Polar |
|---|
| 3z2-r2 | -0.824 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.316 |
0.000 |
0.000 |
| y |
0.000 |
3.316 |
0.000 |
| z |
0.000 |
0.000 |
3.554 |
<r2> (average value of r
2) Å
2
| <r2> |
30.761 |
| (<r2>)1/2 |
5.546 |
