Jump to
S1C2
Energy calculated at MP2=FULL/6-31G**
| | hartrees |
|---|
| Energy at 0K | -477.192466 |
| Energy at 298.15K | -477.198814 |
| HF Energy | -476.747192 |
| Nuclear repulsion energy | 107.850029 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3237 |
3025 |
17.84 |
|
|
|
| 2 |
A' |
3161 |
2953 |
22.41 |
|
|
|
| 3 |
A' |
3139 |
2933 |
16.12 |
|
|
|
| 4 |
A' |
2835 |
2649 |
12.82 |
|
|
|
| 5 |
A' |
1567 |
1464 |
1.45 |
|
|
|
| 6 |
A' |
1553 |
1451 |
4.13 |
|
|
|
| 7 |
A' |
1471 |
1375 |
3.30 |
|
|
|
| 8 |
A' |
1370 |
1280 |
39.12 |
|
|
|
| 9 |
A' |
1158 |
1082 |
2.71 |
|
|
|
| 10 |
A' |
1040 |
972 |
1.72 |
|
|
|
| 11 |
A' |
905 |
846 |
3.25 |
|
|
|
| 12 |
A' |
720 |
673 |
0.52 |
|
|
|
| 13 |
A' |
315 |
294 |
2.31 |
|
|
|
| 14 |
A" |
3245 |
3033 |
24.59 |
|
|
|
| 15 |
A" |
3221 |
3010 |
2.05 |
|
|
|
| 16 |
A" |
1556 |
1454 |
6.82 |
|
|
|
| 17 |
A" |
1321 |
1234 |
0.49 |
|
|
|
| 18 |
A" |
1099 |
1027 |
0.54 |
|
|
|
| 19 |
A" |
817 |
764 |
3.01 |
|
|
|
| 20 |
A" |
276 |
258 |
1.16 |
|
|
|
| 21 |
A" |
182 |
171 |
20.93 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17094.5 cm
-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 15973.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.513 |
0.683 |
0.000 |
| C2 |
0.000 |
0.825 |
0.000 |
| S3 |
-0.756 |
-0.833 |
0.000 |
| H4 |
1.979 |
1.666 |
0.000 |
| H5 |
1.853 |
0.145 |
0.881 |
| H6 |
1.853 |
0.145 |
-0.881 |
| H7 |
-0.325 |
1.371 |
0.882 |
| H8 |
-0.325 |
1.371 |
-0.882 |
| H9 |
-2.023 |
-0.425 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
S3 |
H4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5194 | 2.7284 | 1.0878 | 1.0874 | 1.0874 | 2.1513 | 2.1513 | 3.7052 |
C2 | 1.5194 | | 1.8212 | 2.1507 | 2.1615 | 2.1615 | 1.0874 | 1.0874 | 2.3776 | S3 | 2.7284 | 1.8212 | | 3.7044 | 2.9220 | 2.9220 | 2.4123 | 2.4123 | 1.3308 | H4 | 1.0878 | 2.1507 | 3.7044 | | 1.7626 | 1.7626 | 2.4847 | 2.4847 | 4.5153 | H5 | 1.0874 | 2.1615 | 2.9220 | 1.7626 | | 1.7630 | 2.4992 | 3.0588 | 4.0153 | H6 | 1.0874 | 2.1615 | 2.9220 | 1.7626 | 1.7630 | | 3.0588 | 2.4992 | 4.0153 | H7 | 2.1513 | 1.0874 | 2.4123 | 2.4847 | 2.4992 | 3.0588 | | 1.7644 | 2.6243 | H8 | 2.1513 | 1.0874 | 2.4123 | 2.4847 | 3.0588 | 2.4992 | 1.7644 | | 2.6243 | H9 | 3.7052 | 2.3776 | 1.3308 | 4.5153 | 4.0153 | 4.0153 | 2.6243 | 2.6243 | |
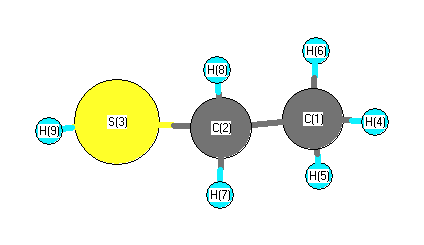 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
S3 |
109.185 |
|
C1 |
C2 |
H7 |
110.128 |
| C1 |
C2 |
H8 |
110.128 |
|
C2 |
C1 |
H4 |
110.053 |
| C2 |
C1 |
H5 |
110.942 |
|
C2 |
C1 |
H6 |
110.942 |
| C2 |
S3 |
H9 |
96.693 |
|
S3 |
C2 |
H7 |
109.467 |
| S3 |
C2 |
H8 |
109.467 |
|
H4 |
C1 |
H5 |
108.251 |
| H4 |
C1 |
H6 |
108.251 |
|
H5 |
C1 |
H6 |
108.311 |
| H7 |
C2 |
H8 |
108.450 |
|
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/6-31G**
| | hartrees |
|---|
| Energy at 0K | -477.193377 |
| Energy at 298.15K | -477.199814 |
| HF Energy | -476.747691 |
| Nuclear repulsion energy | 107.646331 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3247 |
3034 |
22.08 |
|
|
|
| 2 |
A |
3227 |
3016 |
20.10 |
|
|
|
| 3 |
A |
3218 |
3007 |
6.09 |
|
|
|
| 4 |
A |
3161 |
2954 |
16.11 |
|
|
|
| 5 |
A |
3133 |
2927 |
19.32 |
|
|
|
| 6 |
A |
2833 |
2647 |
11.99 |
|
|
|
| 7 |
A |
1562 |
1459 |
1.94 |
|
|
|
| 8 |
A |
1555 |
1453 |
7.84 |
|
|
|
| 9 |
A |
1537 |
1436 |
1.38 |
|
|
|
| 10 |
A |
1469 |
1373 |
2.49 |
|
|
|
| 11 |
A |
1374 |
1284 |
21.03 |
|
|
|
| 12 |
A |
1332 |
1245 |
2.68 |
|
|
|
| 13 |
A |
1174 |
1097 |
12.60 |
|
|
|
| 14 |
A |
1113 |
1040 |
0.22 |
|
|
|
| 15 |
A |
1033 |
965 |
4.72 |
|
|
|
| 16 |
A |
912 |
852 |
8.77 |
|
|
|
| 17 |
A |
768 |
717 |
1.66 |
|
|
|
| 18 |
A |
706 |
660 |
2.38 |
|
|
|
| 19 |
A |
342 |
319 |
2.52 |
|
|
|
| 20 |
A |
284 |
265 |
1.76 |
|
|
|
| 21 |
A |
228 |
213 |
18.68 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17103.7 cm
-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 15981.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31G**
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.629 |
-0.350 |
-0.054 |
| C2 |
0.492 |
0.645 |
0.089 |
| S3 |
-1.159 |
-0.099 |
-0.077 |
| H4 |
2.589 |
0.159 |
0.014 |
| H5 |
1.591 |
-1.100 |
0.734 |
| H6 |
1.574 |
-0.863 |
-1.010 |
| H7 |
0.555 |
1.175 |
1.037 |
| H8 |
0.540 |
1.393 |
-0.700 |
| H9 |
-1.033 |
-0.945 |
0.943 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
S3 |
H4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5180 | 2.7991 | 1.0890 | 1.0882 | 1.0867 | 2.1610 | 2.1546 | 2.9040 |
C2 | 1.5180 | | 1.8180 | 2.1541 | 2.1610 | 2.1577 | 1.0877 | 1.0883 | 2.3623 | S3 | 2.7991 | 1.8180 | | 3.7577 | 3.0366 | 2.9875 | 2.4088 | 2.3449 | 1.3310 | H4 | 1.0890 | 2.1541 | 3.7577 | | 1.7609 | 1.7675 | 2.4928 | 2.4961 | 3.8988 | H5 | 1.0882 | 2.1610 | 3.0366 | 1.7609 | | 1.7601 | 2.5187 | 3.0621 | 2.6370 | H6 | 1.0867 | 2.1577 | 2.9875 | 1.7675 | 1.7601 | | 3.0635 | 2.5013 | 3.2591 | H7 | 2.1610 | 1.0877 | 2.4088 | 2.4928 | 2.5187 | 3.0635 | | 1.7503 | 2.6506 | H8 | 2.1546 | 1.0883 | 2.3449 | 2.4961 | 3.0621 | 2.5013 | 1.7503 | | 3.2614 | H9 | 2.9040 | 2.3623 | 1.3310 | 3.8988 | 2.6370 | 3.2591 | 2.6506 | 3.2614 | |
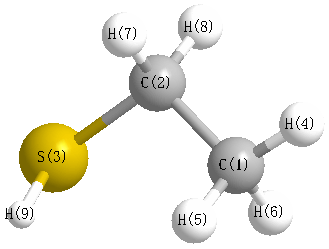 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
S3 |
113.779 |
|
C1 |
C2 |
H7 |
110.977 |
| C1 |
C2 |
H8 |
110.435 |
|
C2 |
C1 |
H4 |
110.350 |
| C2 |
C1 |
H5 |
110.955 |
|
C2 |
C1 |
H6 |
110.781 |
| C2 |
S3 |
H9 |
95.977 |
|
S3 |
C2 |
H7 |
109.410 |
| S3 |
C2 |
H8 |
104.773 |
|
H4 |
C1 |
H5 |
107.956 |
| H4 |
C1 |
H6 |
108.654 |
|
H5 |
C1 |
H6 |
108.051 |
| H7 |
C2 |
H8 |
107.093 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
