Jump to
S1C2
Energy calculated at PBE1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -477.751647 |
| Energy at 298.15K | |
| HF Energy | -477.751647 |
| Nuclear repulsion energy | 107.419122 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBE1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3160 |
3012 |
19.39 |
|
|
|
| 2 |
A' |
3099 |
2954 |
22.83 |
|
|
|
| 3 |
A' |
3080 |
2935 |
17.85 |
|
|
|
| 4 |
A' |
2727 |
2599 |
21.27 |
|
|
|
| 5 |
A' |
1532 |
1460 |
3.11 |
|
|
|
| 6 |
A' |
1517 |
1446 |
3.57 |
|
|
|
| 7 |
A' |
1441 |
1373 |
6.27 |
|
|
|
| 8 |
A' |
1328 |
1266 |
41.13 |
|
|
|
| 9 |
A' |
1136 |
1083 |
2.56 |
|
|
|
| 10 |
A' |
1021 |
973 |
3.68 |
|
|
|
| 11 |
A' |
876 |
835 |
2.79 |
|
|
|
| 12 |
A' |
698 |
665 |
1.14 |
|
|
|
| 13 |
A' |
304 |
290 |
2.82 |
|
|
|
| 14 |
A" |
3171 |
3023 |
26.07 |
|
|
|
| 15 |
A" |
3147 |
2999 |
0.43 |
|
|
|
| 16 |
A" |
1522 |
1451 |
9.58 |
|
|
|
| 17 |
A" |
1288 |
1228 |
0.78 |
|
|
|
| 18 |
A" |
1067 |
1017 |
0.33 |
|
|
|
| 19 |
A" |
802 |
764 |
5.19 |
|
|
|
| 20 |
A" |
263 |
251 |
0.93 |
|
|
|
| 21 |
A" |
163 |
155 |
23.39 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16670.7 cm
-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 15888.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBE1PBE/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.514 |
0.687 |
0.000 |
| C2 |
0.000 |
0.829 |
0.000 |
| S3 |
-0.755 |
-0.836 |
0.000 |
| H4 |
1.983 |
1.677 |
0.000 |
| H5 |
1.861 |
0.147 |
0.887 |
| H6 |
1.861 |
0.147 |
-0.887 |
| H7 |
-0.332 |
1.376 |
0.888 |
| H8 |
-0.332 |
1.376 |
-0.888 |
| H9 |
-2.041 |
-0.436 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
S3 |
H4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5202 | 2.7328 | 1.0952 | 1.0948 | 1.0948 | 2.1603 | 2.1603 | 3.7278 |
C2 | 1.5202 | | 1.8283 | 2.1569 | 2.1710 | 2.1710 | 1.0941 | 1.0941 | 2.4012 | S3 | 2.7328 | 1.8283 | | 3.7166 | 2.9318 | 2.9318 | 2.4210 | 2.4210 | 1.3465 | H4 | 1.0952 | 2.1569 | 3.7166 | | 1.7723 | 1.7723 | 2.4974 | 2.4974 | 4.5452 | H5 | 1.0948 | 2.1710 | 2.9318 | 1.7723 | | 1.7733 | 2.5134 | 3.0765 | 4.0433 | H6 | 1.0948 | 2.1710 | 2.9318 | 1.7723 | 1.7733 | | 3.0765 | 2.5134 | 4.0433 | H7 | 2.1603 | 1.0941 | 2.4210 | 2.4974 | 2.5134 | 3.0765 | | 1.7750 | 2.6446 | H8 | 2.1603 | 1.0941 | 2.4210 | 2.4974 | 3.0765 | 2.5134 | 1.7750 | | 2.6446 | H9 | 3.7278 | 2.4012 | 1.3465 | 4.5452 | 4.0433 | 4.0433 | 2.6446 | 2.6446 | |
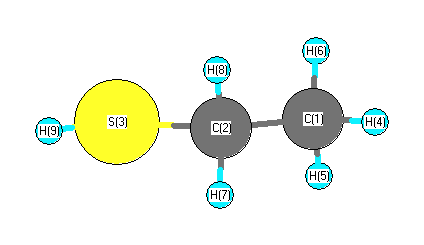 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
S3 |
109.055 |
|
C1 |
C2 |
H7 |
110.388 |
| C1 |
C2 |
H8 |
110.388 |
|
C2 |
C1 |
H4 |
110.059 |
| C2 |
C1 |
H5 |
111.200 |
|
C2 |
C1 |
H6 |
111.200 |
| C2 |
S3 |
H9 |
97.119 |
|
S3 |
C2 |
H7 |
109.287 |
| S3 |
C2 |
H8 |
109.287 |
|
H4 |
C1 |
H5 |
108.052 |
| H4 |
C1 |
H6 |
108.052 |
|
H5 |
C1 |
H6 |
108.162 |
| H7 |
C2 |
H8 |
108.416 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.534 |
|
|
|
| 2 |
C |
-0.472 |
|
|
|
| 3 |
S |
-0.088 |
|
|
|
| 4 |
H |
0.186 |
|
|
|
| 5 |
H |
0.195 |
|
|
|
| 6 |
H |
0.195 |
|
|
|
| 7 |
H |
0.206 |
|
|
|
| 8 |
H |
0.206 |
|
|
|
| 9 |
H |
0.107 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.068 |
1.831 |
0.000 |
1.832 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-24.323 |
-0.200 |
0.000 |
| y |
-0.200 |
-28.157 |
0.000 |
| z |
0.000 |
0.000 |
-28.775 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.143 |
-0.200 |
0.000 |
| y |
-0.200 |
-1.608 |
0.000 |
| z |
0.000 |
0.000 |
-2.535 |
|
| Polar |
|---|
| 3z2-r2 | -5.069 |
|---|
| x2-y2 | 3.834 |
|---|
| xy | -0.200 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.212 |
0.790 |
0.000 |
| y |
0.790 |
5.838 |
0.000 |
| z |
0.000 |
0.000 |
4.471 |
<r2> (average value of r
2) Å
2
| <r2> |
83.152 |
| (<r2>)1/2 |
9.119 |
Jump to
S1C1
Energy calculated at PBE1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -477.752574 |
| Energy at 298.15K | -477.758950 |
| HF Energy | -477.752574 |
| Nuclear repulsion energy | 107.137650 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBE1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3173 |
3024 |
23.27 |
|
|
|
| 2 |
A |
3152 |
3004 |
13.24 |
|
|
|
| 3 |
A |
3142 |
2995 |
13.79 |
|
|
|
| 4 |
A |
3101 |
2955 |
14.88 |
|
|
|
| 5 |
A |
3071 |
2927 |
22.91 |
|
|
|
| 6 |
A |
2728 |
2600 |
19.10 |
|
|
|
| 7 |
A |
1526 |
1455 |
3.51 |
|
|
|
| 8 |
A |
1519 |
1448 |
10.82 |
|
|
|
| 9 |
A |
1505 |
1435 |
2.18 |
|
|
|
| 10 |
A |
1437 |
1370 |
4.83 |
|
|
|
| 11 |
A |
1334 |
1271 |
19.86 |
|
|
|
| 12 |
A |
1299 |
1238 |
4.47 |
|
|
|
| 13 |
A |
1144 |
1091 |
12.23 |
|
|
|
| 14 |
A |
1090 |
1038 |
0.35 |
|
|
|
| 15 |
A |
1011 |
963 |
7.48 |
|
|
|
| 16 |
A |
888 |
846 |
10.00 |
|
|
|
| 17 |
A |
750 |
715 |
2.49 |
|
|
|
| 18 |
A |
682 |
650 |
3.86 |
|
|
|
| 19 |
A |
332 |
317 |
2.89 |
|
|
|
| 20 |
A |
271 |
259 |
3.78 |
|
|
|
| 21 |
A |
235 |
224 |
19.53 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16694.4 cm
-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 15911.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBE1PBE/6-31G**
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.636 |
-0.350 |
-0.054 |
| C2 |
0.496 |
0.643 |
0.092 |
| S3 |
-1.163 |
-0.099 |
-0.079 |
| H4 |
2.602 |
0.166 |
0.014 |
| H5 |
1.606 |
-1.107 |
0.737 |
| H6 |
1.586 |
-0.867 |
-1.017 |
| H7 |
0.554 |
1.179 |
1.044 |
| H8 |
0.536 |
1.397 |
-0.701 |
| H9 |
-1.061 |
-0.948 |
0.962 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
S3 |
H4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5189 | 2.8108 | 1.0967 | 1.0956 | 1.0940 | 2.1714 | 2.1632 | 2.9435 |
C2 | 1.5189 | | 1.8251 | 2.1608 | 2.1702 | 2.1671 | 1.0945 | 1.0949 | 2.3897 | S3 | 2.8108 | 1.8251 | | 3.7755 | 3.0578 | 3.0044 | 2.4169 | 2.3476 | 1.3472 | H4 | 1.0967 | 2.1608 | 3.7755 | | 1.7705 | 1.7781 | 2.5065 | 2.5088 | 3.9439 | H5 | 1.0956 | 2.1702 | 3.0578 | 1.7705 | | 1.7705 | 2.5350 | 3.0792 | 2.6808 | H6 | 1.0940 | 2.1671 | 3.0044 | 1.7781 | 1.7705 | | 3.0820 | 2.5152 | 3.3054 | H7 | 2.1714 | 1.0945 | 2.4169 | 2.5065 | 2.5350 | 3.0820 | | 1.7588 | 2.6718 | H8 | 2.1632 | 1.0949 | 2.3476 | 2.5088 | 3.0792 | 2.5152 | 1.7588 | | 3.2884 | H9 | 2.9435 | 2.3897 | 1.3472 | 3.9439 | 2.6808 | 3.3054 | 2.6718 | 3.2884 | |
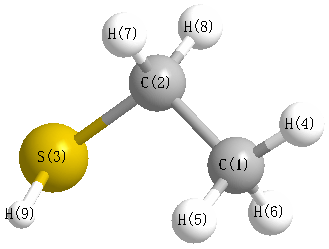 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
S3 |
114.086 |
|
C1 |
C2 |
H7 |
111.345 |
| C1 |
C2 |
H8 |
110.660 |
|
C2 |
C1 |
H4 |
110.365 |
| C2 |
C1 |
H5 |
111.181 |
|
C2 |
C1 |
H6 |
111.027 |
| C2 |
S3 |
H9 |
96.595 |
|
S3 |
C2 |
H7 |
109.183 |
| S3 |
C2 |
H8 |
104.212 |
|
H4 |
C1 |
H5 |
107.722 |
| H4 |
C1 |
H6 |
108.512 |
|
H5 |
C1 |
H6 |
107.916 |
| H7 |
C2 |
H8 |
106.895 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.512 |
|
|
|
| 2 |
C |
-0.486 |
|
|
|
| 3 |
S |
-0.084 |
|
|
|
| 4 |
H |
0.180 |
|
|
|
| 5 |
H |
0.177 |
|
|
|
| 6 |
H |
0.196 |
|
|
|
| 7 |
H |
0.209 |
|
|
|
| 8 |
H |
0.212 |
|
|
|
| 9 |
H |
0.108 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.674 |
0.050 |
0.832 |
1.870 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-29.042 |
1.415 |
-0.871 |
| y |
1.415 |
-26.744 |
-1.737 |
| z |
-0.871 |
-1.737 |
-26.812 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.264 |
1.415 |
-0.871 |
| y |
1.415 |
1.183 |
-1.737 |
| z |
-0.871 |
-1.737 |
1.081 |
|
| Polar |
|---|
| 3z2-r2 | 2.162 |
|---|
| x2-y2 | -2.298 |
|---|
| xy | 1.415 |
|---|
| xz | -0.871 |
|---|
| yz | -1.737 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.658 |
0.436 |
0.028 |
| y |
0.436 |
5.092 |
-0.327 |
| z |
0.028 |
-0.327 |
4.881 |
<r2> (average value of r
2) Å
2
| <r2> |
83.830 |
| (<r2>)1/2 |
9.156 |
