Jump to
S1C2
Energy calculated at HSEh1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -477.784081 |
| Energy at 298.15K | |
| HF Energy | -477.784081 |
| Nuclear repulsion energy | 107.411275 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3157 |
3013 |
18.99 |
|
|
|
| 2 |
A' |
3092 |
2951 |
22.81 |
|
|
|
| 3 |
A' |
3072 |
2932 |
17.77 |
|
|
|
| 4 |
A' |
2722 |
2598 |
14.32 |
|
|
|
| 5 |
A' |
1515 |
1445 |
2.81 |
|
|
|
| 6 |
A' |
1500 |
1431 |
3.06 |
|
|
|
| 7 |
A' |
1425 |
1360 |
5.59 |
|
|
|
| 8 |
A' |
1315 |
1255 |
39.90 |
|
|
|
| 9 |
A' |
1128 |
1077 |
2.35 |
|
|
|
| 10 |
A' |
1014 |
968 |
3.81 |
|
|
|
| 11 |
A' |
869 |
829 |
2.70 |
|
|
|
| 12 |
A' |
693 |
661 |
1.21 |
|
|
|
| 13 |
A' |
304 |
290 |
2.83 |
|
|
|
| 14 |
A" |
3167 |
3022 |
26.10 |
|
|
|
| 15 |
A" |
3143 |
2999 |
0.39 |
|
|
|
| 16 |
A" |
1505 |
1436 |
8.84 |
|
|
|
| 17 |
A" |
1278 |
1220 |
0.72 |
|
|
|
| 18 |
A" |
1057 |
1009 |
0.25 |
|
|
|
| 19 |
A" |
796 |
759 |
4.90 |
|
|
|
| 20 |
A" |
261 |
249 |
0.61 |
|
|
|
| 21 |
A" |
143 |
136 |
22.79 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16577.4 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 15819.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.513 |
0.690 |
0.000 |
| C2 |
0.000 |
0.829 |
0.000 |
| S3 |
-0.755 |
-0.838 |
0.000 |
| H4 |
1.981 |
1.679 |
0.000 |
| H5 |
1.861 |
0.151 |
0.886 |
| H6 |
1.861 |
0.151 |
-0.886 |
| H7 |
-0.334 |
1.374 |
0.887 |
| H8 |
-0.334 |
1.374 |
-0.887 |
| H9 |
-2.040 |
-0.440 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
S3 |
H4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5195 | 2.7344 | 1.0941 | 1.0938 | 1.0938 | 2.1601 | 2.1601 | 3.7284 |
C2 | 1.5195 | | 1.8298 | 2.1559 | 2.1698 | 2.1698 | 1.0933 | 1.0933 | 2.4027 | S3 | 2.7344 | 1.8298 | | 3.7174 | 2.9330 | 2.9330 | 2.4201 | 2.4201 | 1.3453 | H4 | 1.0941 | 2.1559 | 3.7174 | | 1.7704 | 1.7704 | 2.4980 | 2.4980 | 4.5455 | H5 | 1.0938 | 2.1698 | 2.9330 | 1.7704 | | 1.7713 | 2.5128 | 3.0752 | 4.0435 | H6 | 1.0938 | 2.1698 | 2.9330 | 1.7704 | 1.7713 | | 3.0752 | 2.5128 | 4.0435 | H7 | 2.1601 | 1.0933 | 2.4201 | 2.4980 | 2.5128 | 3.0752 | | 1.7741 | 2.6438 | H8 | 2.1601 | 1.0933 | 2.4201 | 2.4980 | 3.0752 | 2.5128 | 1.7741 | | 2.6438 | H9 | 3.7284 | 2.4027 | 1.3453 | 4.5455 | 4.0435 | 4.0435 | 2.6438 | 2.6438 | |
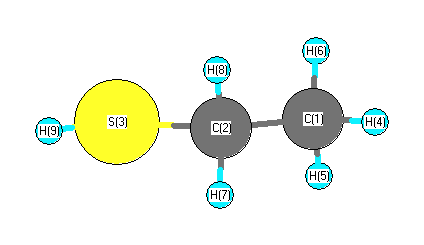 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
S3 |
109.103 |
|
C1 |
C2 |
H7 |
110.469 |
| C1 |
C2 |
H8 |
110.469 |
|
C2 |
C1 |
H4 |
110.090 |
| C2 |
C1 |
H5 |
111.215 |
|
C2 |
C1 |
H6 |
111.215 |
| C2 |
S3 |
H9 |
97.173 |
|
S3 |
C2 |
H7 |
109.160 |
| S3 |
C2 |
H8 |
109.160 |
|
H4 |
C1 |
H5 |
108.034 |
| H4 |
C1 |
H6 |
108.034 |
|
H5 |
C1 |
H6 |
108.133 |
| H7 |
C2 |
H8 |
108.453 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.403 |
|
|
|
| 2 |
C |
-0.393 |
|
|
|
| 3 |
S |
-0.053 |
|
|
|
| 4 |
H |
0.143 |
|
|
|
| 5 |
H |
0.152 |
|
|
|
| 6 |
H |
0.152 |
|
|
|
| 7 |
H |
0.165 |
|
|
|
| 8 |
H |
0.165 |
|
|
|
| 9 |
H |
0.071 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.053 |
1.810 |
0.000 |
1.810 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-24.292 |
-0.196 |
0.000 |
| y |
-0.196 |
-28.125 |
0.000 |
| z |
0.000 |
0.000 |
-28.738 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.140 |
-0.196 |
0.000 |
| y |
-0.196 |
-1.610 |
0.000 |
| z |
0.000 |
0.000 |
-2.530 |
|
| Polar |
|---|
| 3z2-r2 | -5.060 |
|---|
| x2-y2 | 3.833 |
|---|
| xy | -0.196 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.310 |
0.806 |
0.000 |
| y |
0.806 |
5.918 |
0.000 |
| z |
0.000 |
0.000 |
4.537 |
<r2> (average value of r
2) Å
2
| <r2> |
83.170 |
| (<r2>)1/2 |
9.120 |
Jump to
S1C1
Energy calculated at HSEh1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -477.785081 |
| Energy at 298.15K | -477.791448 |
| HF Energy | -477.785081 |
| Nuclear repulsion energy | 107.140972 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3169 |
3024 |
23.35 |
|
|
|
| 2 |
A |
3148 |
3004 |
13.03 |
|
|
|
| 3 |
A |
3139 |
2995 |
13.55 |
|
|
|
| 4 |
A |
3095 |
2954 |
15.02 |
|
|
|
| 5 |
A |
3064 |
2924 |
22.84 |
|
|
|
| 6 |
A |
2726 |
2601 |
13.21 |
|
|
|
| 7 |
A |
1509 |
1440 |
3.05 |
|
|
|
| 8 |
A |
1501 |
1433 |
10.04 |
|
|
|
| 9 |
A |
1488 |
1420 |
1.95 |
|
|
|
| 10 |
A |
1422 |
1357 |
4.20 |
|
|
|
| 11 |
A |
1320 |
1260 |
19.63 |
|
|
|
| 12 |
A |
1288 |
1229 |
4.34 |
|
|
|
| 13 |
A |
1135 |
1083 |
11.23 |
|
|
|
| 14 |
A |
1082 |
1033 |
0.47 |
|
|
|
| 15 |
A |
1004 |
958 |
7.44 |
|
|
|
| 16 |
A |
881 |
840 |
9.79 |
|
|
|
| 17 |
A |
745 |
711 |
2.34 |
|
|
|
| 18 |
A |
677 |
646 |
3.81 |
|
|
|
| 19 |
A |
332 |
317 |
3.03 |
|
|
|
| 20 |
A |
271 |
258 |
4.28 |
|
|
|
| 21 |
A |
239 |
228 |
17.89 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16617.0 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 15857.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G**
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.637 |
-0.350 |
-0.054 |
| C2 |
0.497 |
0.642 |
0.092 |
| S3 |
-1.164 |
-0.098 |
-0.079 |
| H4 |
2.602 |
0.165 |
0.011 |
| H5 |
1.608 |
-1.104 |
0.738 |
| H6 |
1.585 |
-0.869 |
-1.015 |
| H7 |
0.553 |
1.179 |
1.043 |
| H8 |
0.535 |
1.396 |
-0.701 |
| H9 |
-1.063 |
-0.948 |
0.960 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
S3 |
H4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5181 | 2.8119 | 1.0956 | 1.0945 | 1.0930 | 2.1711 | 2.1631 | 2.9454 |
C2 | 1.5181 | | 1.8263 | 2.1598 | 2.1686 | 2.1660 | 1.0936 | 1.0941 | 2.3910 | S3 | 2.8119 | 1.8263 | | 3.7757 | 3.0595 | 3.0042 | 2.4160 | 2.3462 | 1.3461 | H4 | 1.0956 | 2.1598 | 3.7757 | | 1.7687 | 1.7761 | 2.5077 | 2.5086 | 3.9458 | H5 | 1.0945 | 2.1686 | 3.0595 | 1.7687 | | 1.7684 | 2.5332 | 3.0777 | 2.6844 | H6 | 1.0930 | 2.1660 | 3.0042 | 1.7761 | 1.7684 | | 3.0806 | 2.5159 | 3.3042 | H7 | 2.1711 | 1.0936 | 2.4160 | 2.5077 | 2.5332 | 3.0806 | | 1.7575 | 2.6722 | H8 | 2.1631 | 1.0941 | 2.3462 | 2.5086 | 3.0777 | 2.5159 | 1.7575 | | 3.2872 | H9 | 2.9454 | 2.3910 | 1.3461 | 3.9458 | 2.6844 | 3.3042 | 2.6722 | 3.2872 | |
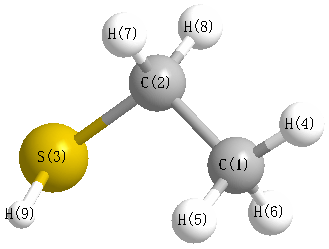 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
S3 |
114.128 |
|
C1 |
C2 |
H7 |
111.423 |
| C1 |
C2 |
H8 |
110.753 |
|
C2 |
C1 |
H4 |
110.400 |
| C2 |
C1 |
H5 |
111.174 |
|
C2 |
C1 |
H6 |
111.051 |
| C2 |
S3 |
H9 |
96.650 |
|
S3 |
C2 |
H7 |
109.079 |
| S3 |
C2 |
H8 |
104.081 |
|
H4 |
C1 |
H5 |
107.717 |
| H4 |
C1 |
H6 |
108.493 |
|
H5 |
C1 |
H6 |
107.886 |
| H7 |
C2 |
H8 |
106.901 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.383 |
|
|
|
| 2 |
C |
-0.405 |
|
|
|
| 3 |
S |
-0.051 |
|
|
|
| 4 |
H |
0.137 |
|
|
|
| 5 |
H |
0.136 |
|
|
|
| 6 |
H |
0.153 |
|
|
|
| 7 |
H |
0.168 |
|
|
|
| 8 |
H |
0.172 |
|
|
|
| 9 |
H |
0.073 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.660 |
0.052 |
0.814 |
1.850 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-28.959 |
1.366 |
-0.853 |
| y |
1.366 |
-26.710 |
-1.714 |
| z |
-0.853 |
-1.714 |
-26.801 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.203 |
1.366 |
-0.853 |
| y |
1.366 |
1.170 |
-1.714 |
| z |
-0.853 |
-1.714 |
1.034 |
|
| Polar |
|---|
| 3z2-r2 | 2.067 |
|---|
| x2-y2 | -2.249 |
|---|
| xy | 1.366 |
|---|
| xz | -0.853 |
|---|
| yz | -1.714 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.757 |
0.436 |
0.023 |
| y |
0.436 |
5.168 |
-0.325 |
| z |
0.023 |
-0.325 |
4.945 |
<r2> (average value of r
2) Å
2
| <r2> |
83.827 |
| (<r2>)1/2 |
9.156 |
