Jump to
S1C2
Energy calculated at wB97X-D/6-31G**
| | hartrees |
|---|
| Energy at 0K | -477.976724 |
| Energy at 298.15K | |
| HF Energy | -477.976724 |
| Nuclear repulsion energy | 107.233265 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at wB97X-D/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3161 |
3161 |
22.24 |
|
|
|
| 2 |
A' |
3098 |
3098 |
23.46 |
|
|
|
| 3 |
A' |
3075 |
3075 |
19.96 |
|
|
|
| 4 |
A' |
2758 |
2758 |
16.56 |
|
|
|
| 5 |
A' |
1524 |
1524 |
2.91 |
|
|
|
| 6 |
A' |
1510 |
1510 |
3.17 |
|
|
|
| 7 |
A' |
1438 |
1438 |
4.80 |
|
|
|
| 8 |
A' |
1323 |
1323 |
38.59 |
|
|
|
| 9 |
A' |
1130 |
1130 |
2.56 |
|
|
|
| 10 |
A' |
1014 |
1014 |
3.63 |
|
|
|
| 11 |
A' |
874 |
874 |
3.03 |
|
|
|
| 12 |
A' |
691 |
691 |
1.43 |
|
|
|
| 13 |
A' |
309 |
309 |
2.92 |
|
|
|
| 14 |
A" |
3169 |
3169 |
30.36 |
|
|
|
| 15 |
A" |
3148 |
3148 |
0.00 |
|
|
|
| 16 |
A" |
1509 |
1509 |
8.60 |
|
|
|
| 17 |
A" |
1282 |
1282 |
0.61 |
|
|
|
| 18 |
A" |
1063 |
1063 |
0.19 |
|
|
|
| 19 |
A" |
801 |
801 |
4.28 |
|
|
|
| 20 |
A" |
237 |
237 |
0.89 |
|
|
|
| 21 |
A" |
131 |
131 |
22.61 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16622.0 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 16622.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at wB97X-D/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.518 |
0.695 |
0.000 |
| C2 |
0.000 |
0.829 |
0.000 |
| S3 |
-0.757 |
-0.840 |
0.000 |
| H4 |
1.980 |
1.686 |
0.000 |
| H5 |
1.867 |
0.158 |
0.886 |
| H6 |
1.867 |
0.158 |
-0.886 |
| H7 |
-0.334 |
1.372 |
0.887 |
| H8 |
-0.334 |
1.372 |
-0.887 |
| H9 |
-2.039 |
-0.441 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
S3 |
H4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5241 | 2.7450 | 1.0935 | 1.0935 | 1.0935 | 2.1622 | 2.1622 | 3.7344 |
C2 | 1.5241 | | 1.8330 | 2.1576 | 2.1732 | 2.1732 | 1.0924 | 1.0924 | 2.4023 | S3 | 2.7450 | 1.8330 | | 3.7250 | 2.9446 | 2.9446 | 2.4213 | 2.4213 | 1.3425 | H4 | 1.0935 | 2.1576 | 3.7250 | | 1.7702 | 1.7702 | 2.4976 | 2.4976 | 4.5474 | H5 | 1.0935 | 2.1732 | 2.9446 | 1.7702 | | 1.7724 | 2.5140 | 3.0763 | 4.0503 | H6 | 1.0935 | 2.1732 | 2.9446 | 1.7702 | 1.7724 | | 3.0763 | 2.5140 | 4.0503 | H7 | 2.1622 | 1.0924 | 2.4213 | 2.4976 | 2.5140 | 3.0763 | | 1.7738 | 2.6429 | H8 | 2.1622 | 1.0924 | 2.4213 | 2.4976 | 3.0763 | 2.5140 | 1.7738 | | 2.6429 | H9 | 3.7344 | 2.4023 | 1.3425 | 4.5474 | 4.0503 | 4.0503 | 2.6429 | 2.6429 | |
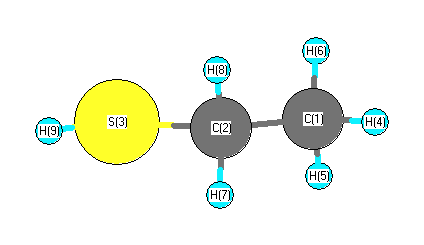 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
S3 |
109.361 |
|
C1 |
C2 |
H7 |
110.364 |
| C1 |
C2 |
H8 |
110.364 |
|
C2 |
C1 |
H4 |
109.939 |
| C2 |
C1 |
H5 |
111.178 |
|
C2 |
C1 |
H6 |
111.178 |
| C2 |
S3 |
H9 |
97.108 |
|
S3 |
C2 |
H7 |
109.086 |
| S3 |
C2 |
H8 |
109.086 |
|
H4 |
C1 |
H5 |
108.079 |
| H4 |
C1 |
H6 |
108.079 |
|
H5 |
C1 |
H6 |
108.278 |
| H7 |
C2 |
H8 |
108.552 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at wB97X-D/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.384 |
|
|
|
| 2 |
C |
-0.370 |
|
|
|
| 3 |
S |
-0.053 |
|
|
|
| 4 |
H |
0.136 |
|
|
|
| 5 |
H |
0.145 |
|
|
|
| 6 |
H |
0.145 |
|
|
|
| 7 |
H |
0.157 |
|
|
|
| 8 |
H |
0.157 |
|
|
|
| 9 |
H |
0.065 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.053 |
1.819 |
0.000 |
1.820 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-24.264 |
-0.198 |
0.000 |
| y |
-0.198 |
-28.102 |
0.000 |
| z |
0.000 |
0.000 |
-28.676 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.125 |
-0.198 |
0.000 |
| y |
-0.198 |
-1.632 |
0.000 |
| z |
0.000 |
0.000 |
-2.493 |
|
| Polar |
|---|
| 3z2-r2 | -4.986 |
|---|
| x2-y2 | 3.838 |
|---|
| xy | -0.198 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.266 |
0.791 |
0.000 |
| y |
0.791 |
5.909 |
0.000 |
| z |
0.000 |
0.000 |
4.551 |
<r2> (average value of r
2) Å
2
| <r2> |
83.471 |
| (<r2>)1/2 |
9.136 |
Jump to
S1C1
Energy calculated at wB97X-D/6-31G**
| | hartrees |
|---|
| Energy at 0K | -477.977774 |
| Energy at 298.15K | |
| HF Energy | -477.977774 |
| Nuclear repulsion energy | 107.012192 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at wB97X-D/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3168 |
3168 |
27.29 |
10.67 |
0.75 |
0.86 |
| 2 |
A |
3149 |
3149 |
10.22 |
88.88 |
0.73 |
0.85 |
| 3 |
A |
3139 |
3139 |
18.35 |
102.87 |
0.70 |
0.82 |
| 4 |
A |
3098 |
3098 |
16.13 |
103.04 |
0.13 |
0.23 |
| 5 |
A |
3063 |
3063 |
23.85 |
119.20 |
0.03 |
0.06 |
| 6 |
A |
2743 |
2743 |
15.30 |
113.77 |
0.36 |
0.53 |
| 7 |
A |
1517 |
1517 |
2.98 |
11.53 |
0.74 |
0.85 |
| 8 |
A |
1511 |
1511 |
9.86 |
18.26 |
0.75 |
0.86 |
| 9 |
A |
1499 |
1499 |
2.04 |
18.34 |
0.75 |
0.85 |
| 10 |
A |
1434 |
1434 |
4.20 |
5.16 |
0.72 |
0.84 |
| 11 |
A |
1331 |
1331 |
20.04 |
1.87 |
0.74 |
0.85 |
| 12 |
A |
1295 |
1295 |
3.13 |
9.87 |
0.74 |
0.85 |
| 13 |
A |
1140 |
1140 |
11.25 |
10.03 |
0.75 |
0.86 |
| 14 |
A |
1084 |
1084 |
0.23 |
5.16 |
0.43 |
0.60 |
| 15 |
A |
1005 |
1005 |
6.90 |
6.57 |
0.75 |
0.86 |
| 16 |
A |
890 |
890 |
9.91 |
4.93 |
0.74 |
0.85 |
| 17 |
A |
751 |
751 |
2.05 |
3.44 |
0.43 |
0.60 |
| 18 |
A |
676 |
676 |
4.12 |
15.28 |
0.29 |
0.45 |
| 19 |
A |
336 |
336 |
2.02 |
1.28 |
0.45 |
0.62 |
| 20 |
A |
266 |
266 |
3.01 |
0.64 |
0.70 |
0.83 |
| 21 |
A |
219 |
219 |
20.23 |
5.35 |
0.75 |
0.85 |
Unscaled Zero Point Vibrational Energy (zpe) 16657.8 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 16657.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at wB97X-D/6-31G**
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.640 |
-0.351 |
-0.054 |
| C2 |
0.497 |
0.645 |
0.091 |
| S3 |
-1.166 |
-0.099 |
-0.078 |
| H4 |
2.603 |
0.165 |
0.020 |
| H5 |
1.604 |
-1.109 |
0.734 |
| H6 |
1.593 |
-0.863 |
-1.018 |
| H7 |
0.554 |
1.176 |
1.044 |
| H8 |
0.539 |
1.398 |
-0.700 |
| H9 |
-1.060 |
-0.952 |
0.954 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
S3 |
H4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5228 | 2.8173 | 1.0949 | 1.0943 | 1.0927 | 2.1721 | 2.1655 | 2.9445 |
C2 | 1.5228 | | 1.8294 | 2.1613 | 2.1715 | 2.1694 | 1.0927 | 1.0932 | 2.3915 | S3 | 2.8173 | 1.8294 | | 3.7794 | 3.0587 | 3.0131 | 2.4172 | 2.3523 | 1.3435 | H4 | 1.0949 | 2.1613 | 3.7794 | | 1.7691 | 1.7761 | 2.5044 | 2.5102 | 3.9421 | H5 | 1.0943 | 2.1715 | 3.0587 | 1.7691 | | 1.7694 | 2.5340 | 3.0787 | 2.6786 | H6 | 1.0927 | 2.1694 | 3.0131 | 1.7761 | 1.7694 | | 3.0810 | 2.5156 | 3.3072 | H7 | 2.1721 | 1.0927 | 2.4172 | 2.5044 | 2.5340 | 3.0810 | | 1.7584 | 2.6722 | H8 | 2.1655 | 1.0932 | 2.3523 | 2.5102 | 3.0787 | 2.5156 | 1.7584 | | 3.2887 | H9 | 2.9445 | 2.3915 | 1.3435 | 3.9421 | 2.6786 | 3.3072 | 2.6722 | 3.2887 | |
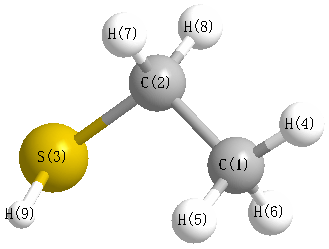 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
S3 |
114.061 |
|
C1 |
C2 |
H7 |
111.225 |
| C1 |
C2 |
H8 |
110.669 |
|
C2 |
C1 |
H4 |
110.236 |
| C2 |
C1 |
H5 |
111.091 |
|
C2 |
C1 |
H6 |
111.010 |
| C2 |
S3 |
H9 |
96.630 |
|
S3 |
C2 |
H7 |
109.005 |
| S3 |
C2 |
H8 |
104.351 |
|
H4 |
C1 |
H5 |
107.826 |
| H4 |
C1 |
H6 |
108.560 |
|
H5 |
C1 |
H6 |
108.009 |
| H7 |
C2 |
H8 |
107.101 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at wB97X-D/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.363 |
|
|
|
| 2 |
C |
-0.382 |
|
|
|
| 3 |
S |
-0.051 |
|
|
|
| 4 |
H |
0.131 |
|
|
|
| 5 |
H |
0.128 |
|
|
|
| 6 |
H |
0.146 |
|
|
|
| 7 |
H |
0.160 |
|
|
|
| 8 |
H |
0.164 |
|
|
|
| 9 |
H |
0.066 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.675 |
0.050 |
0.813 |
1.863 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-28.934 |
1.379 |
-0.853 |
| y |
1.379 |
-26.648 |
-1.707 |
| z |
-0.853 |
-1.707 |
-26.772 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.224 |
1.379 |
-0.853 |
| y |
1.379 |
1.205 |
-1.707 |
| z |
-0.853 |
-1.707 |
1.019 |
|
| Polar |
|---|
| 3z2-r2 | 2.038 |
|---|
| x2-y2 | -2.286 |
|---|
| xy | 1.379 |
|---|
| xz | -0.853 |
|---|
| yz | -1.707 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.681 |
0.449 |
0.029 |
| y |
0.449 |
5.165 |
-0.331 |
| z |
0.029 |
-0.331 |
4.951 |
<r2> (average value of r
2) Å
2
| <r2> |
83.992 |
| (<r2>)1/2 |
9.165 |
