Jump to
S1C2
Energy calculated at B2PLYP/6-31G**
| | hartrees |
|---|
| Energy at 0K | -169.734196 |
| Energy at 298.15K | -169.737714 |
| HF Energy | -169.573861 |
| Nuclear repulsion energy | 71.027410 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3790 |
3790 |
43.15 |
|
|
|
| 2 |
A' |
3642 |
3642 |
39.61 |
|
|
|
| 3 |
A' |
3008 |
3008 |
106.56 |
|
|
|
| 4 |
A' |
1831 |
1831 |
340.60 |
|
|
|
| 5 |
A' |
1640 |
1640 |
62.17 |
|
|
|
| 6 |
A' |
1444 |
1444 |
5.37 |
|
|
|
| 7 |
A' |
1289 |
1289 |
96.54 |
|
|
|
| 8 |
A' |
1062 |
1062 |
2.97 |
|
|
|
| 9 |
A' |
564 |
564 |
11.62 |
|
|
|
| 10 |
A" |
1055 |
1055 |
0.20 |
|
|
|
| 11 |
A" |
652 |
652 |
20.41 |
|
|
|
| 12 |
A" |
109 |
109 |
260.43 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 10042.7 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 10042.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.421 |
0.000 |
| O2 |
1.204 |
0.229 |
0.000 |
| N3 |
-0.944 |
-0.559 |
0.000 |
| H4 |
-0.450 |
1.429 |
0.000 |
| H5 |
-0.647 |
-1.521 |
0.000 |
| H6 |
-1.926 |
-0.346 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.2191 | 1.3610 | 1.1042 | 2.0473 | 2.0727 |
O2 | 1.2191 | | 2.2878 | 2.0439 | 2.5471 | 3.1818 | N3 | 1.3610 | 2.2878 | | 2.0490 | 1.0069 | 1.0047 | H4 | 1.1042 | 2.0439 | 2.0490 | | 2.9572 | 2.3081 | H5 | 2.0473 | 2.5471 | 1.0069 | 2.9572 | | 1.7371 | H6 | 2.0727 | 3.1818 | 1.0047 | 2.3081 | 1.7371 | |
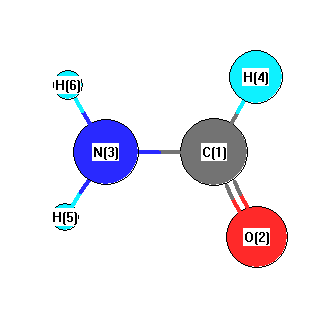 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
118.919 |
|
C1 |
N3 |
H6 |
121.637 |
| O2 |
C1 |
N3 |
124.837 |
|
O2 |
C1 |
H4 |
123.146 |
| N3 |
C1 |
H4 |
112.017 |
|
H5 |
N3 |
H6 |
119.443 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.411 |
|
|
|
| 2 |
O |
-0.454 |
|
|
|
| 3 |
N |
-0.597 |
|
|
|
| 4 |
H |
0.072 |
|
|
|
| 5 |
H |
0.289 |
|
|
|
| 6 |
H |
0.281 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-3.746 |
-0.715 |
0.000 |
3.814 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-17.801 |
-0.051 |
0.000 |
| y |
-0.051 |
-14.687 |
0.000 |
| z |
0.000 |
0.000 |
-18.452 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.232 |
-0.051 |
0.000 |
| y |
-0.051 |
3.440 |
0.000 |
| z |
0.000 |
0.000 |
-2.208 |
|
| Polar |
|---|
| 3z2-r2 | -4.416 |
|---|
| x2-y2 | -3.114 |
|---|
| xy | -0.051 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.188 |
0.277 |
0.000 |
| y |
0.277 |
3.243 |
0.000 |
| z |
0.000 |
0.000 |
1.456 |
<r2> (average value of r
2) Å
2
| <r2> |
40.915 |
| (<r2>)1/2 |
6.396 |
Jump to
S1C1
Energy calculated at B2PLYP/6-31G**
| | hartrees |
|---|
| Energy at 0K | -169.734197 |
| Energy at 298.15K | -169.737704 |
| HF Energy | -169.573857 |
| Nuclear repulsion energy | 71.028786 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3791 |
3791 |
43.26 |
|
|
|
| 2 |
A |
3642 |
3642 |
39.63 |
|
|
|
| 3 |
A |
3009 |
3009 |
106.33 |
|
|
|
| 4 |
A |
1830 |
1830 |
340.63 |
|
|
|
| 5 |
A |
1640 |
1640 |
61.97 |
|
|
|
| 6 |
A |
1444 |
1444 |
5.37 |
|
|
|
| 7 |
A |
1290 |
1290 |
96.32 |
|
|
|
| 8 |
A |
1062 |
1062 |
2.91 |
|
|
|
| 9 |
A |
1055 |
1055 |
0.21 |
|
|
|
| 10 |
A |
652 |
652 |
20.78 |
|
|
|
| 11 |
A |
564 |
564 |
11.66 |
|
|
|
| 12 |
A |
102 |
102 |
260.03 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 10039.9 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 10039.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/6-31G**
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.159 |
0.390 |
0.000 |
| O2 |
-1.200 |
-0.245 |
0.000 |
| N3 |
1.086 |
-0.160 |
-0.000 |
| H4 |
-0.126 |
1.493 |
0.000 |
| H5 |
1.174 |
-1.163 |
0.000 |
| H6 |
1.913 |
0.410 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.2193 | 1.3608 | 1.1041 | 2.0464 | 2.0727 |
O2 | 1.2193 | | 2.2876 | 2.0437 | 2.5456 | 3.1818 | N3 | 1.3608 | 2.2876 | | 2.0494 | 1.0069 | 1.0046 | H4 | 1.1041 | 2.0437 | 2.0494 | | 2.9571 | 2.3093 | H5 | 2.0464 | 2.5456 | 1.0069 | 2.9571 | | 1.7374 | H6 | 2.0727 | 3.1818 | 1.0046 | 2.3093 | 1.7374 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
118.856 |
|
C1 |
N3 |
H6 |
121.665 |
| O2 |
C1 |
N3 |
124.814 |
|
O2 |
C1 |
H4 |
123.107 |
| N3 |
C1 |
H4 |
112.079 |
|
H5 |
N3 |
H6 |
119.479 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.410 |
|
|
|
| 2 |
O |
-0.455 |
|
|
|
| 3 |
N |
-0.597 |
|
|
|
| 4 |
H |
0.072 |
|
|
|
| 5 |
H |
0.289 |
|
|
|
| 6 |
H |
0.281 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
3.739 |
0.760 |
0.001 |
3.815 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-17.390 |
-1.057 |
0.002 |
| y |
-1.057 |
-15.098 |
-0.000 |
| z |
0.002 |
-0.000 |
-18.452 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.615 |
-1.057 |
0.002 |
| y |
-1.057 |
2.823 |
-0.000 |
| z |
0.002 |
-0.000 |
-2.208 |
|
| Polar |
|---|
| 3z2-r2 | -4.416 |
|---|
| x2-y2 | -2.292 |
|---|
| xy | -1.057 |
|---|
| xz | 0.002 |
|---|
| yz | -0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.246 |
0.134 |
0.000 |
| y |
0.134 |
3.184 |
0.000 |
| z |
0.000 |
0.000 |
1.456 |
<r2> (average value of r
2) Å
2
| <r2> |
40.910 |
| (<r2>)1/2 |
6.396 |
