Jump to
S1C2
Energy calculated at B2PLYP=FULL/6-31G**
| | hartrees |
|---|
| Energy at 0K | -169.737558 |
| Energy at 298.15K | -169.741087 |
| HF Energy | -169.573889 |
| Nuclear repulsion energy | 71.044899 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULL/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3791 |
3791 |
43.22 |
|
|
|
| 2 |
A' |
3643 |
3643 |
39.69 |
|
|
|
| 3 |
A' |
3008 |
3008 |
106.59 |
|
|
|
| 4 |
A' |
1832 |
1832 |
340.86 |
|
|
|
| 5 |
A' |
1641 |
1641 |
62.29 |
|
|
|
| 6 |
A' |
1445 |
1445 |
5.34 |
|
|
|
| 7 |
A' |
1289 |
1289 |
96.46 |
|
|
|
| 8 |
A' |
1063 |
1063 |
2.95 |
|
|
|
| 9 |
A' |
564 |
564 |
11.61 |
|
|
|
| 10 |
A" |
1056 |
1056 |
0.19 |
|
|
|
| 11 |
A" |
653 |
653 |
20.53 |
|
|
|
| 12 |
A" |
115 |
115 |
260.37 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 10049.1 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 10049.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP=FULL/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.421 |
0.000 |
| O2 |
1.204 |
0.229 |
0.000 |
| N3 |
-0.944 |
-0.559 |
0.000 |
| H4 |
-0.450 |
1.429 |
0.000 |
| H5 |
-0.647 |
-1.521 |
0.000 |
| H6 |
-1.925 |
-0.346 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.2188 | 1.3606 | 1.1040 | 2.0468 | 2.0722 |
O2 | 1.2188 | | 2.2873 | 2.0434 | 2.5467 | 3.1811 | N3 | 1.3606 | 2.2873 | | 2.0484 | 1.0066 | 1.0044 | H4 | 1.1040 | 2.0434 | 2.0484 | | 2.9564 | 2.3074 | H5 | 2.0468 | 2.5467 | 1.0066 | 2.9564 | | 1.7367 | H6 | 2.0722 | 3.1811 | 1.0044 | 2.3074 | 1.7367 | |
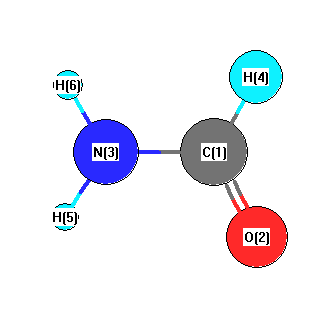 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
118.923 |
|
C1 |
N3 |
H6 |
121.640 |
| O2 |
C1 |
N3 |
124.849 |
|
O2 |
C1 |
H4 |
123.140 |
| N3 |
C1 |
H4 |
112.011 |
|
H5 |
N3 |
H6 |
119.437 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at B2PLYP=FULL/6-31G**
| | hartrees |
|---|
| Energy at 0K | -169.737559 |
| Energy at 298.15K | -169.741077 |
| HF Energy | -169.573885 |
| Nuclear repulsion energy | 71.046299 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULL/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3791 |
3791 |
43.33 |
|
|
|
| 2 |
A |
3642 |
3642 |
39.70 |
|
|
|
| 3 |
A |
3009 |
3009 |
106.37 |
|
|
|
| 4 |
A |
1831 |
1831 |
340.88 |
|
|
|
| 5 |
A |
1641 |
1641 |
62.11 |
|
|
|
| 6 |
A |
1444 |
1444 |
5.35 |
|
|
|
| 7 |
A |
1290 |
1290 |
96.24 |
|
|
|
| 8 |
A |
1063 |
1063 |
2.90 |
|
|
|
| 9 |
A |
1056 |
1056 |
0.20 |
|
|
|
| 10 |
A |
653 |
653 |
20.89 |
|
|
|
| 11 |
A |
564 |
564 |
11.66 |
|
|
|
| 12 |
A |
108 |
108 |
259.99 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 10046.4 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 10046.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP=FULL/6-31G**
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.159 |
0.389 |
0.000 |
| O2 |
1.200 |
-0.245 |
0.000 |
| N3 |
-1.085 |
-0.160 |
-0.000 |
| H4 |
0.126 |
1.493 |
-0.000 |
| H5 |
-1.174 |
-1.162 |
0.000 |
| H6 |
-1.913 |
0.410 |
0.001 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.2191 | 1.3604 | 1.1039 | 2.0459 | 2.0722 |
O2 | 1.2191 | | 2.2871 | 2.0431 | 2.5452 | 3.1811 | N3 | 1.3604 | 2.2871 | | 2.0488 | 1.0067 | 1.0044 | H4 | 1.1039 | 2.0431 | 2.0488 | | 2.9563 | 2.3085 | H5 | 2.0459 | 2.5452 | 1.0067 | 2.9563 | | 1.7370 | H6 | 2.0722 | 3.1811 | 1.0044 | 2.3085 | 1.7370 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
118.858 |
|
C1 |
N3 |
H6 |
121.665 |
| O2 |
C1 |
N3 |
124.828 |
|
O2 |
C1 |
H4 |
123.102 |
| N3 |
C1 |
H4 |
112.070 |
|
H5 |
N3 |
H6 |
119.477 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
