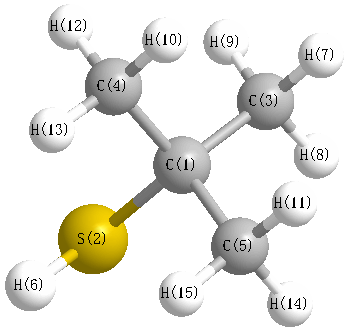
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3162 |
3018 |
22.40 |
|
|
|
| 2 |
A' |
3146 |
3002 |
22.47 |
|
|
|
| 3 |
A' |
3134 |
2991 |
40.92 |
|
|
|
| 4 |
A' |
3064 |
2924 |
19.08 |
|
|
|
| 5 |
A' |
3055 |
2915 |
31.62 |
|
|
|
| 6 |
A' |
2724 |
2600 |
15.26 |
|
|
|
| 7 |
A' |
1523 |
1453 |
7.54 |
|
|
|
| 8 |
A' |
1509 |
1440 |
8.89 |
|
|
|
| 9 |
A' |
1493 |
1425 |
0.59 |
|
|
|
| 10 |
A' |
1437 |
1371 |
1.41 |
|
|
|
| 11 |
A' |
1408 |
1344 |
12.46 |
|
|
|
| 12 |
A' |
1274 |
1215 |
0.49 |
|
|
|
| 13 |
A' |
1207 |
1152 |
45.93 |
|
|
|
| 14 |
A' |
1065 |
1016 |
2.35 |
|
|
|
| 15 |
A' |
953 |
909 |
0.83 |
|
|
|
| 16 |
A' |
890 |
849 |
5.07 |
|
|
|
| 17 |
A' |
844 |
805 |
1.41 |
|
|
|
| 18 |
A' |
601 |
573 |
5.36 |
|
|
|
| 19 |
A' |
391 |
374 |
1.11 |
|
|
|
| 20 |
A' |
368 |
351 |
0.44 |
|
|
|
| 21 |
A' |
299 |
285 |
0.24 |
|
|
|
| 22 |
A' |
283 |
270 |
0.90 |
|
|
|
| 23 |
A" |
3161 |
3016 |
21.22 |
|
|
|
| 24 |
A" |
3156 |
3012 |
6.00 |
|
|
|
| 25 |
A" |
3130 |
2987 |
0.61 |
|
|
|
| 26 |
A" |
3051 |
2912 |
17.05 |
|
|
|
| 27 |
A" |
1511 |
1442 |
7.43 |
|
|
|
| 28 |
A" |
1489 |
1421 |
0.50 |
|
|
|
| 29 |
A" |
1481 |
1414 |
0.01 |
|
|
|
| 30 |
A" |
1409 |
1344 |
11.48 |
|
|
|
| 31 |
A" |
1260 |
1203 |
4.05 |
|
|
|
| 32 |
A" |
1054 |
1006 |
0.13 |
|
|
|
| 33 |
A" |
968 |
924 |
0.02 |
|
|
|
| 34 |
A" |
946 |
903 |
0.57 |
|
|
|
| 35 |
A" |
399 |
381 |
0.55 |
|
|
|
| 36 |
A" |
303 |
289 |
3.51 |
|
|
|
| 37 |
A" |
285 |
272 |
0.01 |
|
|
|
| 38 |
A" |
256 |
244 |
7.95 |
|
|
|
| 39 |
A" |
213 |
203 |
10.76 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 28950.8 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 27627.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.178 |
|
|
|
| 2 |
S |
-0.049 |
|
|
|
| 3 |
C |
-0.371 |
|
|
|
| 4 |
C |
-0.349 |
|
|
|
| 5 |
C |
-0.349 |
|
|
|
| 6 |
H |
0.066 |
|
|
|
| 7 |
H |
0.135 |
|
|
|
| 8 |
H |
0.145 |
|
|
|
| 9 |
H |
0.145 |
|
|
|
| 10 |
H |
0.128 |
|
|
|
| 11 |
H |
0.128 |
|
|
|
| 12 |
H |
0.146 |
|
|
|
| 13 |
H |
0.127 |
|
|
|
| 14 |
H |
0.146 |
|
|
|
| 15 |
H |
0.127 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.645 |
-0.854 |
0.000 |
1.854 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-42.023 |
2.346 |
0.000 |
| y |
2.346 |
-38.708 |
0.000 |
| z |
0.000 |
0.000 |
-41.615 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.861 |
2.346 |
0.000 |
| y |
2.346 |
3.111 |
0.000 |
| z |
0.000 |
0.000 |
-1.249 |
|
| Polar |
|---|
| 3z2-r2 | -2.499 |
|---|
| x2-y2 | -3.314 |
|---|
| xy | 2.346 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.983 |
0.128 |
0.000 |
| y |
0.128 |
8.583 |
0.000 |
| z |
0.000 |
0.000 |
7.948 |
<r2> (average value of r
2) Å
2
| <r2> |
164.445 |
| (<r2>)1/2 |
12.824 |