return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CF3COOH (trifluoroacetic acid)

using model chemistry: QCISD/6-31G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at QCISD/6-31G**
 hartrees
Energy at 0K-525.512671
Energy at 298.15K-525.515932
HF Energy-524.375312
Nuclear repulsion energy336.052668
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3846 3621 85.34      
2 A' 1908 1797 227.63      
3 A' 1488 1401 47.89      
4 A' 1334 1256 144.77      
5 A' 1271 1197 196.62      
6 A' 1178 1109 318.78      
7 A' 812 764 4.10      
8 A' 669 630 77.55      
9 A' 595 560 10.26      
10 A' 429 403 0.12      
11 A' 398 374 2.42      
12 A' 242 228 1.16      
13 A" 1275 1200 277.52      
14 A" 792 745 39.11      
15 A" 612 576 117.51      
16 A" 506 476 0.93      
17 A" 243 229 0.14      
18 A" 34 32 1.25      

Unscaled Zero Point Vibrational Energy (zpe) 8815.2 cm-1
Scaled (by 0.9414) Zero Point Vibrational Energy (zpe) 8298.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/6-31G**
ABC
0.12710 0.08335 0.06894

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/6-31G**

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.087 0.590 0.000
C2 -0.295 -0.897 0.000
O3 0.817 -1.650 0.000
O4 -1.432 -1.300 0.000
F5 -1.008 1.352 0.000
F6 0.817 0.880 1.090
F7 0.817 0.880 -1.090
H8 0.533 -2.576 0.000

Atom - Atom Distances (Å)
  C1 C2 O3 O4 F5 F6 F7 H8
C11.53552.35572.42471.33451.34381.34383.1973
C21.53551.34321.20602.35962.36292.36291.8723
O32.35571.34322.27643.51352.75442.75440.9689
O42.42471.20602.27642.68573.31633.31632.3433
F51.33452.35963.51352.68572.17822.17824.2199
F61.34382.36292.75443.31632.17822.18083.6348
F71.34382.36292.75443.31632.17822.18083.6348
H83.19731.87230.96892.34334.21993.63483.6348

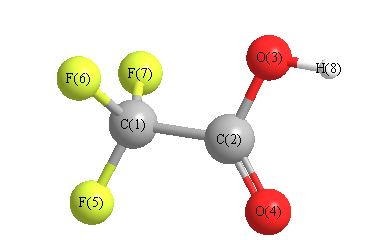
picture of trifluoroacetic acid state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 O3 109.660 C1 C2 O4 123.928
C2 C1 F5 110.410 C2 C1 F6 110.124
C2 C1 F7 110.124 C2 O3 H8 107.034
O3 C2 O4 126.412 F5 C1 F6 108.834
F5 C1 F7 108.834 F6 C1 F7 108.469
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability