Jump to
S1C2
Energy calculated at PBE1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -638.291868 |
| Energy at 298.15K | -638.296776 |
| HF Energy | -638.291868 |
| Nuclear repulsion energy | 157.313825 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBE1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3136 |
2988 |
15.61 |
|
|
|
| 2 |
A' |
3091 |
2946 |
21.95 |
|
|
|
| 3 |
A' |
1550 |
1477 |
1.08 |
|
|
|
| 4 |
A' |
1520 |
1448 |
6.04 |
|
|
|
| 5 |
A' |
1448 |
1380 |
5.61 |
|
|
|
| 6 |
A' |
1307 |
1245 |
12.97 |
|
|
|
| 7 |
A' |
1117 |
1065 |
107.85 |
|
|
|
| 8 |
A' |
1097 |
1046 |
3.20 |
|
|
|
| 9 |
A' |
801 |
763 |
45.83 |
|
|
|
| 10 |
A' |
388 |
370 |
2.02 |
|
|
|
| 11 |
A' |
242 |
230 |
11.62 |
|
|
|
| 12 |
A" |
3204 |
3054 |
8.76 |
|
|
|
| 13 |
A" |
3146 |
2999 |
22.76 |
|
|
|
| 14 |
A" |
1317 |
1256 |
0.01 |
|
|
|
| 15 |
A" |
1236 |
1178 |
0.97 |
|
|
|
| 16 |
A" |
1087 |
1036 |
3.07 |
|
|
|
| 17 |
A" |
804 |
766 |
1.03 |
|
|
|
| 18 |
A" |
135 |
128 |
9.09 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13312.0 cm
-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 12687.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBE1PBE/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.607 |
0.000 |
| C2 |
0.995 |
-0.535 |
0.000 |
| Cl3 |
-1.665 |
-0.038 |
0.000 |
| F4 |
2.261 |
0.009 |
0.000 |
| H5 |
0.122 |
1.227 |
0.890 |
| H6 |
0.122 |
1.227 |
-0.890 |
| H7 |
0.870 |
-1.158 |
0.892 |
| H8 |
0.870 |
-1.158 |
-0.892 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
F4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5144 | 1.7857 | 2.3391 | 1.0916 | 1.0916 | 2.1612 | 2.1612 |
C2 | 1.5144 | | 2.7054 | 1.3786 | 2.1584 | 2.1584 | 1.0955 | 1.0955 | Cl3 | 1.7857 | 2.7054 | | 3.9265 | 2.3641 | 2.3641 | 2.9118 | 2.9118 | F4 | 2.3391 | 1.3786 | 3.9265 | | 2.6175 | 2.6175 | 2.0231 | 2.0231 | H5 | 1.0916 | 2.1584 | 2.3641 | 2.6175 | | 1.7797 | 2.5003 | 3.0704 | H6 | 1.0916 | 2.1584 | 2.3641 | 2.6175 | 1.7797 | | 3.0704 | 2.5003 | H7 | 2.1612 | 1.0955 | 2.9118 | 2.0231 | 2.5003 | 3.0704 | | 1.7844 | H8 | 2.1612 | 1.0955 | 2.9118 | 2.0231 | 3.0704 | 2.5003 | 1.7844 | |
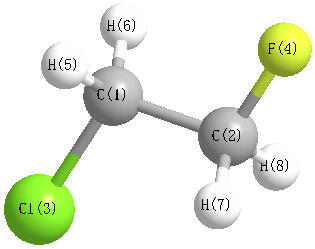 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
107.811 |
|
C1 |
C2 |
H7 |
110.779 |
| C1 |
C2 |
H8 |
110.779 |
|
C2 |
C1 |
Cl3 |
109.855 |
| C2 |
C1 |
H5 |
110.787 |
|
C2 |
C1 |
H6 |
110.787 |
| Cl3 |
C1 |
H5 |
108.058 |
|
Cl3 |
C1 |
H6 |
108.058 |
| F4 |
C2 |
H7 |
109.184 |
|
F4 |
C2 |
H8 |
109.184 |
| H5 |
C1 |
H6 |
109.212 |
|
H7 |
C2 |
H8 |
109.068 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.477 |
|
|
|
| 2 |
C |
0.015 |
|
|
|
| 3 |
Cl |
-0.060 |
|
|
|
| 4 |
F |
-0.308 |
|
|
|
| 5 |
H |
0.236 |
|
|
|
| 6 |
H |
0.236 |
|
|
|
| 7 |
H |
0.180 |
|
|
|
| 8 |
H |
0.180 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.360 |
0.104 |
0.000 |
0.375 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-36.844 |
-0.509 |
0.000 |
| y |
-0.509 |
-28.652 |
0.000 |
| z |
0.000 |
0.000 |
-29.301 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-7.868 |
-0.509 |
0.000 |
| y |
-0.509 |
4.421 |
0.000 |
| z |
0.000 |
0.000 |
3.447 |
|
| Polar |
|---|
| 3z2-r2 | 6.894 |
|---|
| x2-y2 | -8.192 |
|---|
| xy | -0.509 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.884 |
0.345 |
0.000 |
| y |
0.345 |
4.172 |
0.000 |
| z |
0.000 |
0.000 |
3.862 |
<r2> (average value of r
2) Å
2
| <r2> |
133.189 |
| (<r2>)1/2 |
11.541 |
Jump to
S1C1
Energy calculated at PBE1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -638.291341 |
| Energy at 298.15K | -638.296357 |
| HF Energy | -638.291341 |
| Nuclear repulsion energy | 161.438535 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBE1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3193 |
3043 |
5.96 |
|
|
|
| 2 |
A |
3134 |
2987 |
33.44 |
|
|
|
| 3 |
A |
3124 |
2977 |
6.14 |
|
|
|
| 4 |
A |
3066 |
2923 |
30.83 |
|
|
|
| 5 |
A |
1533 |
1461 |
2.73 |
|
|
|
| 6 |
A |
1492 |
1422 |
11.36 |
|
|
|
| 7 |
A |
1452 |
1384 |
17.82 |
|
|
|
| 8 |
A |
1360 |
1296 |
30.78 |
|
|
|
| 9 |
A |
1294 |
1234 |
0.39 |
|
|
|
| 10 |
A |
1245 |
1187 |
4.26 |
|
|
|
| 11 |
A |
1151 |
1097 |
68.73 |
|
|
|
| 12 |
A |
1091 |
1039 |
13.82 |
|
|
|
| 13 |
A |
993 |
946 |
5.97 |
|
|
|
| 14 |
A |
876 |
835 |
6.04 |
|
|
|
| 15 |
A |
704 |
671 |
30.18 |
|
|
|
| 16 |
A |
471 |
449 |
12.32 |
|
|
|
| 17 |
A |
288 |
275 |
0.90 |
|
|
|
| 18 |
A |
134 |
128 |
1.87 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13299.9 cm
-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 12676.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBE1PBE/6-31G**
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.086 |
0.845 |
-0.290 |
| C2 |
1.206 |
0.410 |
0.354 |
| Cl3 |
-1.426 |
-0.283 |
0.065 |
| F4 |
1.649 |
-0.765 |
-0.200 |
| H5 |
-0.378 |
1.831 |
0.079 |
| H6 |
0.025 |
0.885 |
-1.375 |
| H7 |
1.962 |
1.189 |
0.181 |
| H8 |
1.079 |
0.270 |
1.433 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
F4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5077 | 1.7873 | 2.3683 | 1.0929 | 1.0921 | 2.1295 | 2.1577 |
C2 | 1.5077 | | 2.7377 | 1.3727 | 2.1458 | 2.1472 | 1.0990 | 1.0958 | Cl3 | 1.7873 | 2.7377 | | 3.1237 | 2.3601 | 2.3549 | 3.6963 | 2.9076 | F4 | 2.3683 | 1.3727 | 3.1237 | | 3.3054 | 2.5959 | 2.0156 | 2.0159 | H5 | 1.0929 | 2.1458 | 2.3601 | 3.3054 | | 1.7816 | 2.4286 | 2.5284 | H6 | 1.0921 | 2.1472 | 2.3549 | 2.5959 | 1.7816 | | 2.5032 | 3.0620 | H7 | 2.1295 | 1.0990 | 3.6963 | 2.0156 | 2.4286 | 2.5032 | | 1.7869 | H8 | 2.1577 | 1.0958 | 2.9076 | 2.0159 | 2.5284 | 3.0620 | 1.7869 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
110.524 |
|
C1 |
C2 |
H7 |
108.547 |
| C1 |
C2 |
H8 |
110.956 |
|
C2 |
C1 |
Cl3 |
112.101 |
| C2 |
C1 |
H5 |
110.175 |
|
C2 |
C1 |
H6 |
110.341 |
| Cl3 |
C1 |
H5 |
107.602 |
|
Cl3 |
C1 |
H6 |
107.267 |
| F4 |
C2 |
H7 |
108.768 |
|
F4 |
C2 |
H8 |
108.994 |
| H5 |
C1 |
H6 |
109.250 |
|
H7 |
C2 |
H8 |
109.009 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.472 |
|
|
|
| 2 |
C |
0.014 |
|
|
|
| 3 |
Cl |
-0.054 |
|
|
|
| 4 |
F |
-0.298 |
|
|
|
| 5 |
H |
0.222 |
|
|
|
| 6 |
H |
0.237 |
|
|
|
| 7 |
H |
0.170 |
|
|
|
| 8 |
H |
0.182 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.009 |
2.585 |
0.231 |
2.784 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-32.815 |
2.093 |
0.895 |
| y |
2.093 |
-29.075 |
-0.418 |
| z |
0.895 |
-0.418 |
-29.170 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.692 |
2.093 |
0.895 |
| y |
2.093 |
1.918 |
-0.418 |
| z |
0.895 |
-0.418 |
1.775 |
|
| Polar |
|---|
| 3z2-r2 | 3.550 |
|---|
| x2-y2 | -3.740 |
|---|
| xy | 2.093 |
|---|
| xz | 0.895 |
|---|
| yz | -0.418 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.496 |
0.871 |
-0.103 |
| y |
0.871 |
4.559 |
-0.147 |
| z |
-0.103 |
-0.147 |
3.962 |
<r2> (average value of r
2) Å
2
| <r2> |
115.156 |
| (<r2>)1/2 |
10.731 |
